Jump to
S1C2
Vibrational Frequencies calculated at CID/6-31G*
Geometric Data calculated at CID/6-31G*
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CID/6-31G*
| | hartrees |
|---|
| Energy at 0K | -115.535175 |
| Energy at 298.15K | -115.536968 |
| HF Energy | -115.192208 |
| Nuclear repulsion energy | 58.126595 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3411 |
3151 |
0.26 |
|
|
|
| 2 |
A' |
3183 |
2941 |
58.75 |
|
|
|
| 3 |
A' |
1750 |
1616 |
0.28 |
|
|
|
| 4 |
A' |
1304 |
1205 |
10.66 |
|
|
|
| 5 |
A' |
1075 |
993 |
13.91 |
|
|
|
| 6 |
A' |
977 |
902 |
4.96 |
|
|
|
| 7 |
A' |
633 |
585 |
73.43 |
|
|
|
| 8 |
A" |
3510 |
3243 |
1675.52 |
|
|
|
| 9 |
A" |
2356 |
2176 |
4044.73 |
|
|
|
| 10 |
A" |
1061 |
980 |
14.36 |
|
|
|
| 11 |
A" |
1040 |
961 |
0.21 |
|
|
|
| 12 |
A" |
903 |
834 |
0.10 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 10601.1 cm
-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 9792.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CID/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.037 |
0.887 |
0.000 |
| H2 |
0.742 |
1.648 |
0.000 |
| C3 |
-0.037 |
-0.418 |
0.654 |
| C4 |
-0.037 |
-0.418 |
-0.654 |
| H5 |
-0.040 |
-0.977 |
1.573 |
| H6 |
-0.040 |
-0.977 |
-1.573 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
C3 |
C4 |
H5 |
H6 |
| C1 | | 1.0889 | 1.4594 | 1.4594 | 2.4385 | 2.4385 |
H2 | 1.0889 | | 2.3024 | 2.3024 | 3.1580 | 3.1580 | C3 | 1.4594 | 2.3024 | | 1.3077 | 1.0755 | 2.2957 | C4 | 1.4594 | 2.3024 | 1.3077 | | 2.2957 | 1.0755 | H5 | 2.4385 | 3.1580 | 1.0755 | 2.2957 | | 3.1457 | H6 | 2.4385 | 3.1580 | 2.2957 | 1.0755 | 3.1457 | |
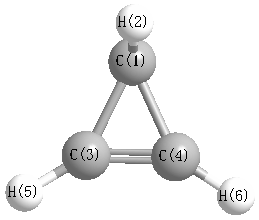 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C3 |
C4 |
63.384 |
|
C1 |
C3 |
H5 |
147.911 |
| C1 |
C4 |
C3 |
63.384 |
|
C1 |
C4 |
H6 |
147.911 |
| H2 |
C1 |
C3 |
128.656 |
|
H2 |
C1 |
C4 |
128.656 |
| C3 |
C1 |
C4 |
53.233 |
|
C3 |
C4 |
H6 |
148.705 |
| C4 |
C3 |
H5 |
148.705 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
