return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C10H8 (naphthalene)

using model chemistry: CID/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes D2H 1Ag
Energy calculated at CID/6-31G*
 hartrees
Energy at 0K-384.333206
Energy at 298.15K-384.341798
HF Energy-383.354906
Nuclear repulsion energy460.574190
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 Ag 3349 3093 0.00      
2 Ag 3321 3068 0.00      
3 Ag 1755 1621 0.00      
4 Ag 1591 1470 0.00      
5 Ag 1498 1383 0.00      
6 Ag 1264 1167 0.00      
7 Ag 1110 1025 0.00      
8 Ag 822 759 0.00      
9 Ag 543 502 0.00      
10 Au 1004 928 0.00      
11 Au 895 827 0.00      
12 Au 618 571 0.00      
13 Au 197 182 0.00      
14 B1g 1004 928 0.00      
15 B1g 777 717 0.00      
16 B1g 409 378 0.00      
17 B1u 3335 3081 59.98      
18 B1u 3315 3062 4.30      
19 B1u 1774 1639 6.91      
20 B1u 1505 1390 5.33      
21 B1u 1366 1262 7.84      
22 B1u 1222 1128 3.06      
23 B1u 845 780 0.79      
24 B1u 380 351 1.46      
25 B2g 1018 941 0.00      
26 B2g 915 845 0.00      
27 B2g 664 614 0.00      
28 B2g 490 453 0.00      
29 B2u 3347 3092 49.83      
30 B2u 3317 3064 0.77      
31 B2u 1659 1532 10.66      
32 B2u 1465 1353 2.63      
33 B2u 1296 1197 0.57      
34 B2u 1208 1115 2.55      
35 B2u 1085 1002 3.20      
36 B2u 660 610 4.67      
37 B3g 3333 3079 0.00      
38 B3g 3313 3060 0.00      
39 B3g 1816 1678 0.00      
40 B3g 1591 1470 0.00      
41 B3g 1342 1240 0.00      
42 B3g 1250 1155 0.00      
43 B3g 994 918 0.00      
44 B3g 542 501 0.00      
45 B3u 1014 937 1.85      
46 B3u 845 781 125.99      
47 B3u 507 468 16.58      
48 B3u 181 167 2.03      

Unscaled Zero Point Vibrational Energy (zpe) 33875.9 cm-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 31291.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CID/6-31G*
ABC
0.10555 0.04162 0.02985

See section I.F.4 to change rotational constant units
Geometric Data calculated at CID/6-31G*

Point Group is D2h

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 1.239 1.392
C2 0.000 2.415 0.706
C3 0.000 2.415 -0.706
C4 0.000 1.239 -1.392
C5 0.000 -1.239 -1.392
C6 0.000 -2.415 -0.706
C7 0.000 -2.415 0.706
C8 0.000 -1.239 1.392
C9 0.000 0.000 0.706
C10 0.000 0.000 -0.706
H11 0.000 1.237 2.472
H12 0.000 3.353 1.238
H13 0.000 3.353 -1.238
H14 0.000 1.237 -2.472
H15 0.000 -1.237 -2.472
H16 0.000 -3.353 -1.238
H17 0.000 -3.353 1.238
H18 0.000 -1.237 2.472

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 C8 C9 C10 H11 H12 H13 H14 H15 H16 H17 H18
C11.36082.40512.78403.72744.21353.71772.47841.41672.43621.07952.11933.37433.86354.58895.29214.59472.7013
C21.36081.41262.40514.21355.03174.82933.71772.41472.79712.12201.07862.15913.38904.84086.08665.79214.0559
C32.40511.41261.36083.71774.82935.03174.21352.79712.41473.38902.15911.07862.12204.05595.79216.08664.8408
C42.78402.40511.36082.47843.71774.21353.72742.43621.41673.86353.37432.11931.07952.70134.59475.29214.5889
C53.72744.21353.71772.47841.36082.40512.78402.43621.41674.58895.29214.59472.70131.07952.11933.37433.8635
C64.21355.03174.82933.71771.36081.41262.40512.79712.41474.84086.08665.79214.05592.12201.07862.15913.3890
C73.71774.82935.03174.21352.40511.41261.36082.41472.79714.05595.79216.08664.84083.38902.15911.07862.1220
C82.47843.71774.21353.72742.78402.40511.36081.41672.43622.70134.59475.29214.58893.86353.37432.11931.0795
C91.41672.41472.79712.43622.43622.79712.41471.41671.41102.15613.39503.87563.40933.40933.87563.39502.1561
C102.43622.79712.41471.41671.41672.41472.79712.43621.41103.40933.87563.39502.15612.15613.39503.87563.4093
H111.07952.12203.38903.86354.58894.84084.05592.70132.15613.40932.44914.27084.94305.52765.90174.75272.4740
H122.11931.07862.15913.37435.29216.08665.79214.59473.39503.87562.44912.47654.27085.90177.14866.70594.7527
H133.37432.15911.07862.11934.59475.79216.08665.29213.87563.39504.27082.47652.44914.75276.70597.14865.9017
H143.86353.38902.12201.07952.70134.05594.84084.58893.40932.15614.94304.27082.44912.47404.75275.90175.5276
H154.58894.84084.05592.70131.07952.12203.38903.86353.40932.15615.52765.90174.75272.47402.44914.27084.9430
H165.29216.08665.79214.59472.11931.07862.15913.37433.87563.39505.90177.14866.70594.75272.44912.47654.2708
H174.59475.79216.08665.29213.37432.15911.07862.11933.39503.87564.75276.70597.14865.90174.27082.47652.4491
H182.70134.05594.84084.58893.86353.38902.12201.07952.15613.40932.47404.75275.90175.52764.94304.27082.4491

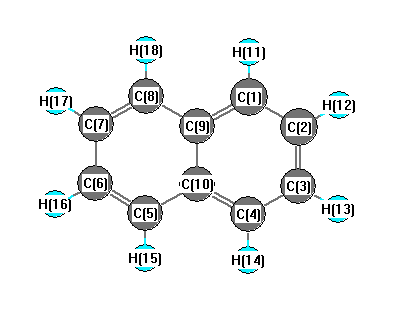
picture of naphthalene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 120.256 C1 C2 H12 120.194
C1 C9 C8 122.030 C1 C9 C10 118.985
C2 C1 C9 120.759 C2 C1 H11 120.375
C2 C3 C4 120.256 C2 C3 H13 119.549
C3 C2 H12 119.549 C3 C4 C10 120.759
C3 C4 H14 120.375 C4 C3 H13 120.194
C4 C10 C5 122.030 C4 C10 C9 118.985
C5 C6 C7 120.256 C5 C6 H16 120.194
C5 C10 C9 118.985 C6 C5 C10 120.759
C6 C5 H15 120.375 C6 C7 C8 120.256
C6 C7 H17 119.549 C7 C6 H16 119.549
C7 C8 C9 120.759 C7 C8 H18 120.375
C8 C7 H17 120.194 C8 C9 C10 118.985
C9 C1 H11 118.867 C9 C8 H18 118.867
C10 C4 H14 118.867 C10 C5 H15 118.867
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability