return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C10H8 (naphthalene)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes D2H 1Ag
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-385.454601
Energy at 298.15K-385.462837
HF Energy-385.033719
Nuclear repulsion energy457.413583
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 Ag 3234 3070 0.00      
2 Ag 3207 3044 0.00      
3 Ag 1640 1556 0.00      
4 Ag 1517 1440 0.00      
5 Ag 1437 1364 0.00      
6 Ag 1205 1143 0.00      
7 Ag 1062 1009 0.00      
8 Ag 780 740 0.00      
9 Ag 522 496 0.00      
10 Au 944 896 0.00      
11 Au 842 800 0.00      
12 Au 600 569 0.00      
13 Au 188 178 0.00      
14 B1g 928 881 0.00      
15 B1g 730 693 0.00      
16 B1g 390 370 0.00      
17 B1u 3221 3057 58.92      
18 B1u 3202 3040 11.16      
19 B1u 1663 1579 5.46      
20 B1u 1439 1366 3.87      
21 B1u 1306 1239 8.45      
22 B1u 1164 1105 2.78      
23 B1u 812 770 0.27      
24 B1u 364 345 1.16      
25 B2g 956 908 0.00      
26 B2g 864 820 0.00      
27 B2g 676 642 0.00      
28 B2g 470 446 0.00      
29 B2u 3233 3069 53.52      
30 B2u 3204 3041 0.39      
31 B2u 1577 1497 7.77      
32 B2u 1429 1357 1.42      
33 B2u 1256 1192 1.39      
34 B2u 1196 1136 0.73      
35 B2u 1051 997 4.18      
36 B2u 636 603 3.55      
37 B3g 3220 3057 0.00      
38 B3g 3200 3038 0.00      
39 B3g 1700 1614 0.00      
40 B3g 1513 1437 0.00      
41 B3g 1284 1219 0.00      
42 B3g 1189 1129 0.00      
43 B3g 951 903 0.00      
44 B3g 518 492 0.00      
45 B3u 938 890 1.75      
46 B3u 795 755 102.73      
47 B3u 481 456 12.17      
48 B3u 173 165 1.76      

Unscaled Zero Point Vibrational Energy (zpe) 32452.4 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 30803.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.10405 0.04108 0.02945

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is D2h

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 1.246 1.404
C2 0.000 2.434 0.710
C3 0.000 2.434 -0.710
C4 0.000 1.246 -1.404
C5 0.000 -1.246 -1.404
C6 0.000 -2.434 -0.710
C7 0.000 -2.434 0.710
C8 0.000 -1.246 1.404
C9 0.000 0.000 0.714
C10 0.000 0.000 -0.714
H11 0.000 1.239 2.491
H12 0.000 3.380 1.244
H13 0.000 3.380 -1.244
H14 0.000 1.239 -2.491
H15 0.000 -1.239 -2.491
H16 0.000 -3.380 -1.244
H17 0.000 -3.380 1.244
H18 0.000 -1.239 2.491

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 C8 C9 C10 H11 H12 H13 H14 H15 H16 H17 H18
C11.37592.42502.80703.75324.24383.74462.49151.42402.45641.08712.14003.40073.89414.61955.32974.62822.7125
C21.37591.42052.42504.24385.07144.86843.74462.43422.82002.14411.08602.17133.41664.87246.13375.83844.0822
C32.42501.42051.37593.74464.86845.07144.24382.82002.43423.41662.17131.08602.14414.08225.83846.13374.8724
C42.80702.42501.37592.49153.74464.24383.75322.45641.42403.89413.40072.14001.08712.71254.62825.32974.6195
C53.75324.24383.74462.49151.37592.42502.80702.45641.42404.61955.32974.62822.71251.08712.14003.40073.8941
C64.24385.07144.86843.74461.37591.42052.42502.82002.43424.87246.13375.83844.08222.14411.08602.17133.4166
C73.74464.86845.07144.24382.42501.42051.37592.43422.82004.08225.83846.13374.87243.41662.17131.08602.1441
C82.49153.74464.24383.75322.80702.42501.37591.42402.45642.71254.62825.32974.61953.89413.40072.14001.0871
C91.42402.43422.82002.45642.45642.82002.43421.42401.42722.16653.42123.90593.43553.43553.90593.42122.1665
C102.45642.82002.43421.42401.42402.43422.82002.45641.42723.43553.90593.42122.16652.16653.42123.90593.4355
H111.08712.14413.41663.89414.61954.87244.08222.71252.16653.43552.47684.30474.98125.56385.94024.78432.4787
H122.14001.08602.17133.40075.32976.13375.83844.62823.42123.90592.47682.48864.30475.94027.20316.75954.7843
H133.40072.17131.08602.14004.62825.83846.13375.32973.90593.42124.30472.48862.47684.78436.75957.20315.9402
H143.89413.41662.14411.08712.71254.08224.87244.61953.43552.16654.98124.30472.47682.47874.78435.94025.5638
H154.61954.87244.08222.71251.08712.14413.41663.89413.43552.16655.56385.94024.78432.47872.47684.30474.9812
H165.32976.13375.83844.62822.14001.08602.17133.40073.90593.42125.94027.20316.75954.78432.47682.48864.3047
H174.62825.83846.13375.32973.40072.17131.08602.14003.42123.90594.78436.75957.20315.94024.30472.48862.4768
H182.71254.08224.87244.61953.89413.41662.14411.08712.16653.43552.47874.78435.94025.56384.98124.30472.4768

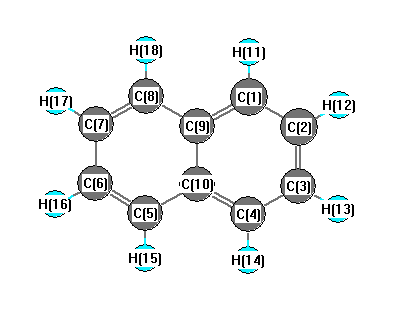
picture of naphthalene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 120.255 C1 C2 H12 120.289
C1 C9 C8 122.042 C1 C9 C10 118.979
C2 C1 C9 120.766 C2 C1 H11 120.591
C2 C3 C4 120.255 C2 C3 H13 119.455
C3 C2 H12 119.455 C3 C4 C10 120.766
C3 C4 H14 120.591 C4 C3 H13 120.289
C4 C10 C5 122.042 C4 C10 C9 118.979
C5 C6 C7 120.255 C5 C6 H16 120.289
C5 C10 C9 118.979 C6 C5 C10 120.766
C6 C5 H15 120.591 C6 C7 C8 120.255
C6 C7 H17 119.455 C7 C6 H16 119.455
C7 C8 C9 120.766 C7 C8 H18 120.591
C8 C7 H17 120.289 C8 C9 C10 118.979
C9 C1 H11 118.643 C9 C8 H18 118.643
C10 C4 H14 118.643 C10 C5 H15 118.643
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C -0.195      
2 C -0.167      
3 C -0.167      
4 C -0.195      
5 C -0.195      
6 C -0.167      
7 C -0.167      
8 C -0.195      
9 C 0.078      
10 C 0.078      
11 H 0.162      
12 H 0.161      
13 H 0.161      
14 H 0.162      
15 H 0.162      
16 H 0.161      
17 H 0.161      
18 H 0.162      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 4.797 0.000 0.000
y 0.000 22.402 0.000
z 0.000 0.000 16.169


<r2> (average value of r2) Å2
<r2> 361.464
(<r2>)1/2 19.012