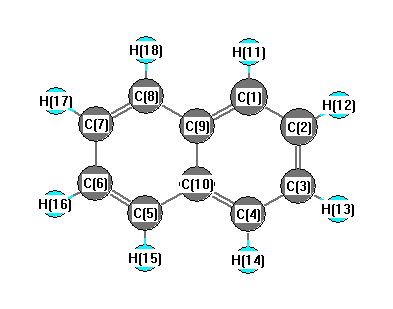
Vibrational Frequencies calculated at HSEh1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3236 |
3078 |
0.00 |
|
|
|
| 2 |
Ag |
3209 |
3052 |
0.00 |
|
|
|
| 3 |
Ag |
1656 |
1575 |
0.00 |
|
|
|
| 4 |
Ag |
1513 |
1439 |
0.00 |
|
|
|
| 5 |
Ag |
1451 |
1379 |
0.00 |
|
|
|
| 6 |
Ag |
1195 |
1136 |
0.00 |
|
|
|
| 7 |
Ag |
1066 |
1014 |
0.00 |
|
|
|
| 8 |
Ag |
786 |
747 |
0.00 |
|
|
|
| 9 |
Ag |
523 |
497 |
0.00 |
|
|
|
| 10 |
Au |
993 |
944 |
0.00 |
|
|
|
| 11 |
Au |
856 |
814 |
0.00 |
|
|
|
| 12 |
Au |
637 |
606 |
0.00 |
|
|
|
| 13 |
Au |
190 |
181 |
0.00 |
|
|
|
| 14 |
B1g |
954 |
908 |
0.00 |
|
|
|
| 15 |
B1g |
735 |
699 |
0.00 |
|
|
|
| 16 |
B1g |
397 |
378 |
0.00 |
|
|
|
| 17 |
B1u |
3223 |
3065 |
55.84 |
|
|
|
| 18 |
B1u |
3204 |
3047 |
9.29 |
|
|
|
| 19 |
B1u |
1679 |
1597 |
5.11 |
|
|
|
| 20 |
B1u |
1432 |
1362 |
4.64 |
|
|
|
| 21 |
B1u |
1301 |
1238 |
8.14 |
|
|
|
| 22 |
B1u |
1161 |
1104 |
4.48 |
|
|
|
| 23 |
B1u |
810 |
771 |
0.25 |
|
|
|
| 24 |
B1u |
362 |
345 |
1.23 |
|
|
|
| 25 |
B2g |
1002 |
953 |
0.00 |
|
|
|
| 26 |
B2g |
903 |
858 |
0.00 |
|
|
|
| 27 |
B2g |
785 |
747 |
0.00 |
|
|
|
| 28 |
B2g |
480 |
456 |
0.00 |
|
|
|
| 29 |
B2u |
3236 |
3077 |
48.09 |
|
|
|
| 30 |
B2u |
3207 |
3050 |
0.57 |
|
|
|
| 31 |
B2u |
1583 |
1505 |
8.39 |
|
|
|
| 32 |
B2u |
1432 |
1362 |
1.65 |
|
|
|
| 33 |
B2u |
1257 |
1195 |
1.76 |
|
|
|
| 34 |
B2u |
1187 |
1129 |
0.82 |
|
|
|
| 35 |
B2u |
1057 |
1005 |
5.67 |
|
|
|
| 36 |
B2u |
633 |
602 |
3.63 |
|
|
|
| 37 |
B3g |
3222 |
3064 |
0.00 |
|
|
|
| 38 |
B3g |
3203 |
3046 |
0.00 |
|
|
|
| 39 |
B3g |
1714 |
1630 |
0.00 |
|
|
|
| 40 |
B3g |
1515 |
1441 |
0.00 |
|
|
|
| 41 |
B3g |
1279 |
1217 |
0.00 |
|
|
|
| 42 |
B3g |
1186 |
1127 |
0.00 |
|
|
|
| 43 |
B3g |
950 |
903 |
0.00 |
|
|
|
| 44 |
B3g |
518 |
492 |
0.00 |
|
|
|
| 45 |
B3u |
972 |
924 |
3.96 |
|
|
|
| 46 |
B3u |
807 |
768 |
106.75 |
|
|
|
| 47 |
B3u |
491 |
467 |
15.44 |
|
|
|
| 48 |
B3u |
176 |
167 |
1.97 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 32681.1 cm
-1
Scaled (by 0.951) Zero Point Vibrational Energy (zpe) 31079.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.209 |
|
|
|
| 2 |
C |
-0.173 |
|
|
|
| 3 |
C |
-0.173 |
|
|
|
| 4 |
C |
-0.209 |
|
|
|
| 5 |
C |
-0.209 |
|
|
|
| 6 |
C |
-0.173 |
|
|
|
| 7 |
C |
-0.173 |
|
|
|
| 8 |
C |
-0.209 |
|
|
|
| 9 |
C |
0.099 |
|
|
|
| 10 |
C |
0.099 |
|
|
|
| 11 |
H |
0.166 |
|
|
|
| 12 |
H |
0.167 |
|
|
|
| 13 |
H |
0.167 |
|
|
|
| 14 |
H |
0.166 |
|
|
|
| 15 |
H |
0.166 |
|
|
|
| 16 |
H |
0.167 |
|
|
|
| 17 |
H |
0.167 |
|
|
|
| 18 |
H |
0.166 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-62.554 |
0.000 |
0.000 |
| y |
0.000 |
-49.806 |
0.000 |
| z |
0.000 |
0.000 |
-50.198 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-12.552 |
0.000 |
0.000 |
| y |
0.000 |
6.570 |
0.000 |
| z |
0.000 |
0.000 |
5.982 |
|
| Polar |
|---|
| 3z2-r2 | 11.964 |
|---|
| x2-y2 | -12.749 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.877 |
0.000 |
0.000 |
| y |
0.000 |
22.564 |
0.000 |
| z |
0.000 |
0.000 |
16.230 |
<r2> (average value of r
2) Å
2
| <r2> |
359.630 |
| (<r2>)1/2 |
18.964 |