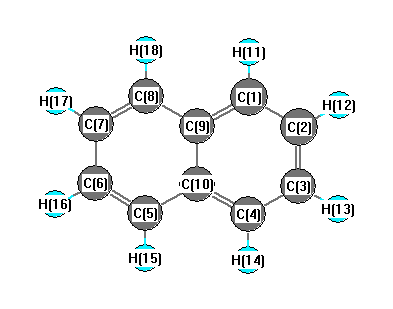
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3139 |
3087 |
0.00 |
|
|
|
| 2 |
Ag |
3112 |
3060 |
0.00 |
|
|
|
| 3 |
Ag |
1587 |
1561 |
0.00 |
|
|
|
| 4 |
Ag |
1463 |
1439 |
0.00 |
|
|
|
| 5 |
Ag |
1402 |
1379 |
0.00 |
|
|
|
| 6 |
Ag |
1157 |
1138 |
0.00 |
|
|
|
| 7 |
Ag |
1034 |
1017 |
0.00 |
|
|
|
| 8 |
Ag |
759 |
746 |
0.00 |
|
|
|
| 9 |
Ag |
507 |
499 |
0.00 |
|
|
|
| 10 |
Au |
943 |
927 |
0.00 |
|
|
|
| 11 |
Au |
820 |
806 |
0.00 |
|
|
|
| 12 |
Au |
614 |
604 |
0.00 |
|
|
|
| 13 |
Au |
182 |
179 |
0.00 |
|
|
|
| 14 |
B1g |
904 |
889 |
0.00 |
|
|
|
| 15 |
B1g |
705 |
693 |
0.00 |
|
|
|
| 16 |
B1g |
381 |
375 |
0.00 |
|
|
|
| 17 |
B1u |
3125 |
3074 |
59.34 |
|
|
|
| 18 |
B1u |
3107 |
3056 |
12.71 |
|
|
|
| 19 |
B1u |
1612 |
1586 |
3.98 |
|
|
|
| 20 |
B1u |
1385 |
1362 |
3.65 |
|
|
|
| 21 |
B1u |
1260 |
1239 |
8.39 |
|
|
|
| 22 |
B1u |
1123 |
1104 |
3.91 |
|
|
|
| 23 |
B1u |
786 |
773 |
0.11 |
|
|
|
| 24 |
B1u |
352 |
347 |
1.06 |
|
|
|
| 25 |
B2g |
951 |
935 |
0.00 |
|
|
|
| 26 |
B2g |
866 |
851 |
0.00 |
|
|
|
| 27 |
B2g |
758 |
746 |
0.00 |
|
|
|
| 28 |
B2g |
463 |
455 |
0.00 |
|
|
|
| 29 |
B2u |
3138 |
3086 |
52.27 |
|
|
|
| 30 |
B2u |
3109 |
3058 |
0.53 |
|
|
|
| 31 |
B2u |
1527 |
1502 |
6.68 |
|
|
|
| 32 |
B2u |
1401 |
1377 |
1.09 |
|
|
|
| 33 |
B2u |
1225 |
1205 |
1.71 |
|
|
|
| 34 |
B2u |
1158 |
1139 |
0.38 |
|
|
|
| 35 |
B2u |
1026 |
1009 |
5.60 |
|
|
|
| 36 |
B2u |
615 |
605 |
3.09 |
|
|
|
| 37 |
B3g |
3125 |
3073 |
0.00 |
|
|
|
| 38 |
B3g |
3106 |
3054 |
0.00 |
|
|
|
| 39 |
B3g |
1642 |
1615 |
0.00 |
|
|
|
| 40 |
B3g |
1462 |
1437 |
0.00 |
|
|
|
| 41 |
B3g |
1238 |
1218 |
0.00 |
|
|
|
| 42 |
B3g |
1147 |
1128 |
0.00 |
|
|
|
| 43 |
B3g |
919 |
903 |
0.00 |
|
|
|
| 44 |
B3g |
501 |
493 |
0.00 |
|
|
|
| 45 |
B3u |
920 |
905 |
3.52 |
|
|
|
| 46 |
B3u |
775 |
762 |
93.81 |
|
|
|
| 47 |
B3u |
472 |
465 |
13.16 |
|
|
|
| 48 |
B3u |
169 |
166 |
1.77 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31584.3 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 31063.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.199 |
|
|
|
| 2 |
C |
-0.144 |
|
|
|
| 3 |
C |
-0.144 |
|
|
|
| 4 |
C |
-0.199 |
|
|
|
| 5 |
C |
-0.199 |
|
|
|
| 6 |
C |
-0.144 |
|
|
|
| 7 |
C |
-0.144 |
|
|
|
| 8 |
C |
-0.199 |
|
|
|
| 9 |
C |
0.136 |
|
|
|
| 10 |
C |
0.136 |
|
|
|
| 11 |
H |
0.137 |
|
|
|
| 12 |
H |
0.138 |
|
|
|
| 13 |
H |
0.138 |
|
|
|
| 14 |
H |
0.137 |
|
|
|
| 15 |
H |
0.137 |
|
|
|
| 16 |
H |
0.138 |
|
|
|
| 17 |
H |
0.138 |
|
|
|
| 18 |
H |
0.137 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-62.299 |
0.000 |
0.000 |
| y |
0.000 |
-50.223 |
0.000 |
| z |
0.000 |
0.000 |
-50.667 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-11.854 |
0.000 |
0.000 |
| y |
0.000 |
6.260 |
0.000 |
| z |
0.000 |
0.000 |
5.594 |
|
| Polar |
|---|
| 3z2-r2 | 11.187 |
|---|
| x2-y2 | -12.076 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.929 |
0.000 |
0.000 |
| y |
0.000 |
23.497 |
0.000 |
| z |
0.000 |
0.000 |
16.790 |
<r2> (average value of r
2) Å
2
| <r2> |
364.401 |
| (<r2>)1/2 |
19.089 |