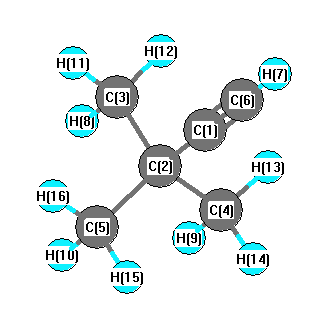
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3500 |
3349 |
55.30 |
|
|
|
| 2 |
A1 |
3140 |
3004 |
61.35 |
|
|
|
| 3 |
A1 |
3062 |
2929 |
25.52 |
|
|
|
| 4 |
A1 |
2234 |
2137 |
3.96 |
|
|
|
| 5 |
A1 |
1543 |
1476 |
7.87 |
|
|
|
| 6 |
A1 |
1448 |
1385 |
1.46 |
|
|
|
| 7 |
A1 |
1288 |
1233 |
25.04 |
|
|
|
| 8 |
A1 |
908 |
868 |
1.52 |
|
|
|
| 9 |
A1 |
703 |
673 |
0.67 |
|
|
|
| 10 |
A1 |
378 |
361 |
0.15 |
|
|
|
| 11 |
A2 |
3145 |
3009 |
0.00 |
|
|
|
| 12 |
A2 |
1500 |
1435 |
0.00 |
|
|
|
| 13 |
A2 |
972 |
930 |
0.00 |
|
|
|
| 14 |
A2 |
222 |
212 |
0.00 |
|
|
|
| 15 |
E |
3148 |
3012 |
36.29 |
|
|
|
| 15 |
E |
3148 |
3012 |
36.29 |
|
|
|
| 16 |
E |
3134 |
2999 |
5.62 |
|
|
|
| 16 |
E |
3134 |
2999 |
5.62 |
|
|
|
| 17 |
E |
3056 |
2923 |
23.01 |
|
|
|
| 17 |
E |
3056 |
2923 |
23.00 |
|
|
|
| 18 |
E |
1524 |
1458 |
6.60 |
|
|
|
| 18 |
E |
1524 |
1458 |
6.60 |
|
|
|
| 19 |
E |
1510 |
1445 |
0.50 |
|
|
|
| 19 |
E |
1510 |
1445 |
0.50 |
|
|
|
| 20 |
E |
1415 |
1354 |
5.78 |
|
|
|
| 20 |
E |
1415 |
1354 |
5.78 |
|
|
|
| 21 |
E |
1244 |
1190 |
5.91 |
|
|
|
| 21 |
E |
1244 |
1190 |
5.91 |
|
|
|
| 22 |
E |
1059 |
1013 |
0.06 |
|
|
|
| 22 |
E |
1059 |
1013 |
0.06 |
|
|
|
| 23 |
E |
946 |
905 |
0.76 |
|
|
|
| 23 |
E |
946 |
905 |
0.76 |
|
|
|
| 24 |
E |
619 |
592 |
51.07 |
|
|
|
| 24 |
E |
619 |
592 |
51.07 |
|
|
|
| 25 |
E |
555 |
531 |
1.68 |
|
|
|
| 25 |
E |
555 |
531 |
1.68 |
|
|
|
| 26 |
E |
353 |
338 |
0.41 |
|
|
|
| 26 |
E |
353 |
338 |
0.41 |
|
|
|
| 27 |
E |
278 |
266 |
0.00 |
|
|
|
| 27 |
E |
278 |
266 |
0.00 |
|
|
|
| 28 |
E |
178 |
170 |
0.28 |
|
|
|
| 28 |
E |
178 |
170 |
0.28 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31040.5 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 29696.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.349 |
|
|
-0.189 |
| 2 |
C |
-0.105 |
|
|
0.736 |
| 3 |
C |
-0.491 |
|
|
-0.531 |
| 4 |
C |
-0.491 |
|
|
-0.519 |
| 5 |
C |
-0.491 |
|
|
-0.512 |
| 6 |
C |
-0.548 |
|
|
-0.381 |
| 7 |
H |
0.221 |
|
|
0.280 |
| 8 |
H |
0.165 |
|
|
0.117 |
| 9 |
H |
0.165 |
|
|
0.115 |
| 10 |
H |
0.165 |
|
|
0.112 |
| 11 |
H |
0.177 |
|
|
0.132 |
| 12 |
H |
0.177 |
|
|
0.131 |
| 13 |
H |
0.177 |
|
|
0.128 |
| 14 |
H |
0.177 |
|
|
0.128 |
| 15 |
H |
0.177 |
|
|
0.127 |
| 16 |
H |
0.177 |
|
|
0.126 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-0.620 |
0.620 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
0.001 |
-0.001 |
-0.624 |
0.624 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.386 |
0.000 |
0.000 |
| y |
0.000 |
-38.386 |
0.000 |
| z |
0.000 |
0.000 |
-34.479 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.954 |
0.000 |
0.000 |
| y |
0.000 |
-1.954 |
0.000 |
| z |
0.000 |
0.000 |
3.907 |
|
| Polar |
|---|
| 3z2-r2 | 7.814 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
174.738 |
| (<r2>)1/2 |
13.219 |