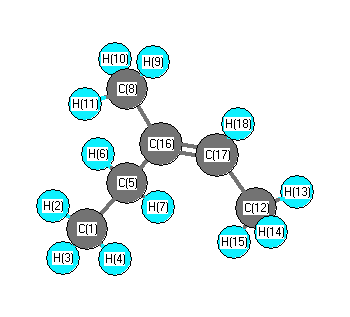
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3074 |
3023 |
56.99 |
|
|
|
| 2 |
A |
3065 |
3015 |
41.72 |
|
|
|
| 3 |
A |
3061 |
3011 |
7.58 |
|
|
|
| 4 |
A |
3059 |
3009 |
41.60 |
|
|
|
| 5 |
A |
3055 |
3005 |
2.95 |
|
|
|
| 6 |
A |
3022 |
2972 |
13.24 |
|
|
|
| 7 |
A |
3012 |
2962 |
18.19 |
|
|
|
| 8 |
A |
3010 |
2960 |
32.96 |
|
|
|
| 9 |
A |
2985 |
2936 |
28.79 |
|
|
|
| 10 |
A |
2966 |
2917 |
15.90 |
|
|
|
| 11 |
A |
2963 |
2914 |
66.35 |
|
|
|
| 12 |
A |
2959 |
2910 |
21.65 |
|
|
|
| 13 |
A |
1703 |
1675 |
0.24 |
|
|
|
| 14 |
A |
1493 |
1468 |
9.98 |
|
|
|
| 15 |
A |
1482 |
1458 |
5.44 |
|
|
|
| 16 |
A |
1480 |
1456 |
3.84 |
|
|
|
| 17 |
A |
1478 |
1453 |
3.76 |
|
|
|
| 18 |
A |
1463 |
1439 |
4.50 |
|
|
|
| 19 |
A |
1462 |
1437 |
7.20 |
|
|
|
| 20 |
A |
1455 |
1431 |
1.66 |
|
|
|
| 21 |
A |
1393 |
1370 |
1.57 |
|
|
|
| 22 |
A |
1391 |
1368 |
2.35 |
|
|
|
| 23 |
A |
1382 |
1359 |
3.57 |
|
|
|
| 24 |
A |
1357 |
1335 |
2.07 |
|
|
|
| 25 |
A |
1307 |
1285 |
5.20 |
|
|
|
| 26 |
A |
1273 |
1252 |
2.11 |
|
|
|
| 27 |
A |
1208 |
1188 |
1.69 |
|
|
|
| 28 |
A |
1121 |
1102 |
4.94 |
|
|
|
| 29 |
A |
1078 |
1060 |
2.63 |
|
|
|
| 30 |
A |
1056 |
1039 |
5.73 |
|
|
|
| 31 |
A |
1036 |
1019 |
1.13 |
|
|
|
| 32 |
A |
1025 |
1008 |
3.05 |
|
|
|
| 33 |
A |
998 |
981 |
8.16 |
|
|
|
| 34 |
A |
991 |
974 |
0.83 |
|
|
|
| 35 |
A |
910 |
895 |
6.17 |
|
|
|
| 36 |
A |
813 |
800 |
11.07 |
|
|
|
| 37 |
A |
776 |
763 |
2.26 |
|
|
|
| 38 |
A |
735 |
723 |
1.08 |
|
|
|
| 39 |
A |
549 |
540 |
2.53 |
|
|
|
| 40 |
A |
513 |
505 |
3.09 |
|
|
|
| 41 |
A |
387 |
381 |
0.43 |
|
|
|
| 42 |
A |
299 |
294 |
0.31 |
|
|
|
| 43 |
A |
262 |
258 |
1.06 |
|
|
|
| 44 |
A |
239 |
235 |
0.88 |
|
|
|
| 45 |
A |
212 |
209 |
0.06 |
|
|
|
| 46 |
A |
171 |
168 |
1.24 |
|
|
|
| 47 |
A |
131 |
129 |
0.27 |
|
|
|
| 48 |
A |
46 |
46 |
0.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 35451.6 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 34866.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.479 |
|
|
|
| 2 |
H |
0.159 |
|
|
|
| 3 |
H |
0.156 |
|
|
|
| 4 |
H |
0.164 |
|
|
|
| 5 |
C |
-0.361 |
|
|
|
| 6 |
H |
0.150 |
|
|
|
| 7 |
H |
0.154 |
|
|
|
| 8 |
C |
-0.558 |
|
|
|
| 9 |
H |
0.163 |
|
|
|
| 10 |
H |
0.163 |
|
|
|
| 11 |
H |
0.165 |
|
|
|
| 12 |
C |
-0.533 |
|
|
|
| 13 |
H |
0.167 |
|
|
|
| 14 |
H |
0.168 |
|
|
|
| 15 |
H |
0.164 |
|
|
|
| 16 |
C |
0.203 |
|
|
|
| 17 |
C |
-0.164 |
|
|
|
| 18 |
H |
0.120 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.108 |
-0.136 |
0.024 |
0.175 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.291 |
0.389 |
0.655 |
| y |
0.389 |
-38.914 |
-0.487 |
| z |
0.655 |
-0.487 |
-40.385 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.359 |
0.389 |
0.655 |
| y |
0.389 |
0.424 |
-0.487 |
| z |
0.655 |
-0.487 |
-1.782 |
|
| Polar |
|---|
| 3z2-r2 | -3.565 |
|---|
| x2-y2 | 0.624 |
|---|
| xy | 0.389 |
|---|
| xz | 0.655 |
|---|
| yz | -0.487 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
12.504 |
0.420 |
0.667 |
| y |
0.420 |
9.781 |
-0.231 |
| z |
0.667 |
-0.231 |
7.489 |
<r2> (average value of r
2) Å
2
| <r2> |
212.801 |
| (<r2>)1/2 |
14.588 |