return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H8O2 (Ethene, 1,1-dimethoxy-)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-307.305932
Energy at 298.15K-307.315310
HF Energy-307.018814
Nuclear repulsion energy248.787983
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3263 3097 0.41      
2 A1 3204 3042 23.57      
3 A1 3071 2915 6.42      
4 A1 1728 1640 217.80      
5 A1 1562 1483 3.50      
6 A1 1526 1448 8.05      
7 A1 1463 1389 14.27      
8 A1 1252 1189 3.91      
9 A1 1189 1129 0.11      
10 A1 900 854 39.32      
11 A1 506 480 0.30      
12 A1 273 259 4.66      
13 A2 3139 2980 0.00      
14 A2 1539 1461 0.00      
15 A2 1200 1139 0.00      
16 A2 684 649 0.00      
17 A2 251 238 0.00      
18 A2 126 120 0.00      
19 B1 3140 2980 74.94      
20 B1 1539 1461 11.78      
21 B1 1205 1144 2.07      
22 B1 704 668 91.58      
23 B1 674 640 3.13      
24 B1 271 258 0.02      
25 B1 98 93 9.50      
26 B2 3344 3174 5.23      
27 B2 3204 3042 16.34      
28 B2 3071 2915 61.56      
29 B2 1558 1479 16.75      
30 B2 1528 1450 31.82      
31 B2 1375 1305 491.12      
32 B2 1241 1178 18.28      
33 B2 1086 1030 108.93      
34 B2 961 913 0.07      
35 B2 566 537 0.13      
36 B2 388 369 8.14      

Unscaled Zero Point Vibrational Energy (zpe) 26414.5 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 25072.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.30268 0.07683 0.06274

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 1.420
C2 0.000 0.000 0.075
H3 0.000 0.921 1.981
H4 0.000 -0.921 1.981
O5 0.000 1.081 -0.739
O6 0.000 -1.081 -0.739
C7 0.000 2.354 -0.109
C8 0.000 -2.354 -0.109
H9 0.000 3.083 -0.918
H10 0.000 -3.083 -0.918
H11 -0.893 2.487 0.510
H12 0.893 2.487 0.510
H13 0.893 -2.487 0.510
H14 -0.893 -2.487 0.510

Atom - Atom Distances (Å)
  C1 C2 H3 H4 O5 O6 C7 C8 H9 H10 H11 H12 H13 H14
C11.34491.07851.07852.41452.41452.80692.80693.86893.86892.79472.79472.79472.7947
C21.34492.11692.11691.35331.35332.36122.36123.23883.23882.67822.67822.67822.6782
H31.07852.11691.84182.72483.37752.53413.88503.61624.94302.32662.32663.81773.8177
H41.07852.11691.84183.37752.72483.88502.53414.94303.61623.81773.81772.32662.3266
O52.41451.35332.72483.37752.16241.42033.49262.00974.16802.08212.08213.88473.8847
O62.41451.35333.37752.72482.16243.49261.42034.16802.00973.88473.88472.08212.0821
C72.80692.36122.53413.88501.42033.49264.70811.08885.49681.09471.09474.96164.9616
C82.80692.36123.88502.53413.49261.42034.70815.49681.08884.96164.96161.09471.0947
H93.86893.23883.61624.94302.00974.16801.08885.49686.16591.78651.78655.81915.8191
H103.86893.23884.94303.61624.16802.00975.49681.08886.16595.81915.81911.78651.7865
H112.79472.67822.32663.81772.08213.88471.09474.96161.78655.81911.78595.28514.9742
H122.79472.67822.32663.81772.08213.88471.09474.96161.78655.81911.78594.97425.2851
H132.79472.67823.81772.32663.88472.08214.96161.09475.81911.78655.28514.97421.7859
H142.79472.67823.81772.32663.88472.08214.96161.09475.81911.78654.97425.28511.7859

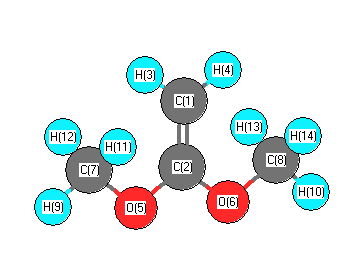
picture of Ethene, 1,1-dimethoxy- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 O5 126.974 C1 C2 O6 126.974
C2 C1 H3 121.359 C2 C1 H4 121.359
C2 O5 C7 116.686 C2 O6 C8 116.686
H3 C1 H4 117.282 O5 C2 O6 106.052
O5 C7 H9 105.685 O5 C7 H11 111.093
O5 C7 H12 111.093 O6 C8 H10 105.685
O6 C8 H13 111.093 O6 C8 H14 111.093
H9 C7 H11 109.804 H9 C7 H12 109.804
H10 C8 H13 109.804 H10 C8 H14 109.804
H11 C7 H12 109.305 H13 C8 H14 109.305
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C -0.550      
2 C 0.697      
3 H 0.160      
4 H 0.160      
5 O -0.531      
6 O -0.531      
7 C -0.208      
8 C -0.208      
9 H 0.181      
10 H 0.181      
11 H 0.162      
12 H 0.162      
13 H 0.162      
14 H 0.162      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 4.843 0.000 0.000
y 0.000 8.910 0.000
z 0.000 0.000 7.566


<r2> (average value of r2) Å2
<r2> 186.933
(<r2>)1/2 13.672