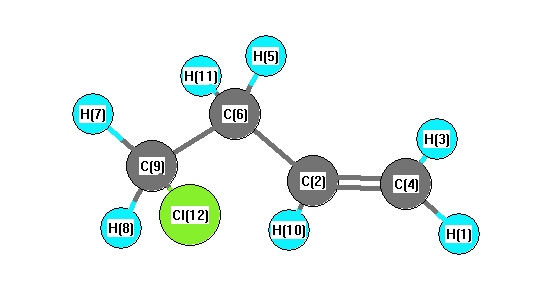
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3250 |
3109 |
16.41 |
|
|
|
| 2 |
A |
3182 |
3045 |
3.83 |
|
|
|
| 3 |
A |
3176 |
3038 |
10.46 |
|
|
|
| 4 |
A |
3163 |
3026 |
11.39 |
|
|
|
| 5 |
A |
3113 |
2979 |
26.73 |
|
|
|
| 6 |
A |
3090 |
2956 |
6.72 |
|
|
|
| 7 |
A |
3046 |
2914 |
17.15 |
|
|
|
| 8 |
A |
1740 |
1665 |
5.43 |
|
|
|
| 9 |
A |
1507 |
1442 |
2.85 |
|
|
|
| 10 |
A |
1495 |
1430 |
12.46 |
|
|
|
| 11 |
A |
1471 |
1407 |
2.95 |
|
|
|
| 12 |
A |
1375 |
1315 |
1.84 |
|
|
|
| 13 |
A |
1349 |
1291 |
36.60 |
|
|
|
| 14 |
A |
1327 |
1270 |
7.13 |
|
|
|
| 15 |
A |
1277 |
1222 |
0.11 |
|
|
|
| 16 |
A |
1202 |
1150 |
0.07 |
|
|
|
| 17 |
A |
1145 |
1096 |
3.74 |
|
|
|
| 18 |
A |
1049 |
1004 |
6.38 |
|
|
|
| 19 |
A |
1037 |
992 |
12.46 |
|
|
|
| 20 |
A |
1010 |
966 |
5.67 |
|
|
|
| 21 |
A |
947 |
906 |
42.42 |
|
|
|
| 22 |
A |
911 |
871 |
17.45 |
|
|
|
| 23 |
A |
826 |
791 |
5.10 |
|
|
|
| 24 |
A |
679 |
650 |
27.32 |
|
|
|
| 25 |
A |
668 |
639 |
13.59 |
|
|
|
| 26 |
A |
455 |
435 |
2.21 |
|
|
|
| 27 |
A |
357 |
341 |
0.40 |
|
|
|
| 28 |
A |
247 |
236 |
1.94 |
|
|
|
| 29 |
A |
124 |
118 |
2.36 |
|
|
|
| 30 |
A |
77 |
74 |
0.18 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 22147.0 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 21188.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.170 |
|
|
0.182 |
| 2 |
C |
-0.071 |
|
|
-0.097 |
| 3 |
H |
0.160 |
|
|
0.179 |
| 4 |
C |
-0.388 |
|
|
-0.446 |
| 5 |
H |
0.197 |
|
|
0.059 |
| 6 |
C |
-0.370 |
|
|
0.031 |
| 7 |
H |
0.218 |
|
|
0.133 |
| 8 |
H |
0.223 |
|
|
0.150 |
| 9 |
C |
-0.417 |
|
|
-0.208 |
| 10 |
H |
0.181 |
|
|
0.131 |
| 11 |
H |
0.181 |
|
|
0.061 |
| 12 |
Cl |
-0.086 |
|
|
-0.175 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.369 |
1.524 |
-0.284 |
2.068 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
1.415 |
1.585 |
-0.312 |
2.147 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-40.369 |
-1.275 |
-0.629 |
| y |
-1.275 |
-35.247 |
-0.564 |
| z |
-0.629 |
-0.564 |
-38.501 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.495 |
-1.275 |
-0.629 |
| y |
-1.275 |
4.189 |
-0.564 |
| z |
-0.629 |
-0.564 |
-0.693 |
|
| Polar |
|---|
| 3z2-r2 | -1.387 |
|---|
| x2-y2 | -5.123 |
|---|
| xy | -1.275 |
|---|
| xz | -0.629 |
|---|
| yz | -0.564 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
199.838 |
| (<r2>)1/2 |
14.136 |