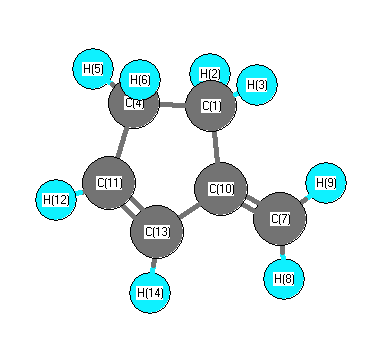
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3252 |
3111 |
15.11 |
|
|
|
| 2 |
A |
3235 |
3095 |
20.59 |
|
|
|
| 3 |
A |
3212 |
3073 |
4.61 |
|
|
|
| 4 |
A |
3171 |
3034 |
7.38 |
|
|
|
| 5 |
A |
3119 |
2984 |
21.47 |
|
|
|
| 6 |
A |
3079 |
2946 |
21.13 |
|
|
|
| 7 |
A |
3071 |
2938 |
15.85 |
|
|
|
| 8 |
A |
3044 |
2913 |
42.21 |
|
|
|
| 9 |
A |
1729 |
1654 |
24.64 |
|
|
|
| 10 |
A |
1672 |
1599 |
0.48 |
|
|
|
| 11 |
A |
1520 |
1454 |
3.56 |
|
|
|
| 12 |
A |
1493 |
1428 |
1.20 |
|
|
|
| 13 |
A |
1474 |
1410 |
0.79 |
|
|
|
| 14 |
A |
1386 |
1326 |
0.90 |
|
|
|
| 15 |
A |
1346 |
1288 |
2.35 |
|
|
|
| 16 |
A |
1291 |
1235 |
0.30 |
|
|
|
| 17 |
A |
1281 |
1226 |
1.98 |
|
|
|
| 18 |
A |
1254 |
1200 |
0.23 |
|
|
|
| 19 |
A |
1180 |
1129 |
0.49 |
|
|
|
| 20 |
A |
1131 |
1082 |
1.69 |
|
|
|
| 21 |
A |
1035 |
990 |
4.02 |
|
|
|
| 22 |
A |
1029 |
984 |
3.37 |
|
|
|
| 23 |
A |
979 |
937 |
2.35 |
|
|
|
| 24 |
A |
972 |
929 |
2.07 |
|
|
|
| 25 |
A |
930 |
890 |
7.46 |
|
|
|
| 26 |
A |
880 |
842 |
41.48 |
|
|
|
| 27 |
A |
863 |
826 |
0.52 |
|
|
|
| 28 |
A |
853 |
816 |
14.25 |
|
|
|
| 29 |
A |
796 |
761 |
9.34 |
|
|
|
| 30 |
A |
778 |
744 |
0.80 |
|
|
|
| 31 |
A |
678 |
648 |
10.22 |
|
|
|
| 32 |
A |
630 |
602 |
0.39 |
|
|
|
| 33 |
A |
515 |
493 |
13.90 |
|
|
|
| 34 |
A |
357 |
341 |
0.70 |
|
|
|
| 35 |
A |
293 |
280 |
0.64 |
|
|
|
| 36 |
A |
54 |
52 |
0.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 26791.1 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 25631.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.395 |
|
|
-0.137 |
| 2 |
H |
0.185 |
|
|
0.068 |
| 3 |
H |
0.185 |
|
|
0.068 |
| 4 |
C |
-0.366 |
|
|
-0.108 |
| 5 |
H |
0.177 |
|
|
0.064 |
| 6 |
H |
0.177 |
|
|
0.064 |
| 7 |
C |
-0.477 |
|
|
-0.684 |
| 8 |
H |
0.159 |
|
|
0.224 |
| 9 |
H |
0.158 |
|
|
0.209 |
| 10 |
C |
0.170 |
|
|
0.357 |
| 11 |
C |
-0.165 |
|
|
-0.043 |
| 12 |
H |
0.162 |
|
|
0.097 |
| 13 |
C |
-0.129 |
|
|
-0.354 |
| 14 |
H |
0.159 |
|
|
0.174 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.774 |
-0.213 |
0.000 |
0.802 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
0.773 |
-0.210 |
0.001 |
0.801 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-34.823 |
-0.049 |
0.001 |
| y |
-0.049 |
-33.739 |
0.002 |
| z |
0.001 |
0.002 |
-38.441 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.267 |
-0.049 |
0.001 |
| y |
-0.049 |
2.893 |
0.002 |
| z |
0.001 |
0.002 |
-4.160 |
|
| Polar |
|---|
| 3z2-r2 | -8.320 |
|---|
| x2-y2 | -1.084 |
|---|
| xy | -0.049 |
|---|
| xz | 0.001 |
|---|
| yz | 0.002 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
152.750 |
| (<r2>)1/2 |
12.359 |