return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H8 (3-Methylenecyclopentene)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-233.129315
Energy at 298.15K-233.137559
HF Energy-232.875537
Nuclear repulsion energy216.469723
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3270 3104 14.69      
2 A 3250 3085 19.67      
3 A 3228 3064 3.47      
4 A 3189 3027 6.89      
5 A 3139 2979 25.09      
6 A 3099 2941 22.75      
7 A 3096 2939 15.40      
8 A 3069 2913 39.88      
9 A 1727 1639 17.54      
10 A 1664 1580 0.34      
11 A 1548 1469 2.05      
12 A 1522 1445 0.75      
13 A 1490 1414 0.60      
14 A 1398 1327 0.74      
15 A 1361 1292 2.06      
16 A 1307 1241 0.48      
17 A 1291 1226 1.62      
18 A 1271 1206 0.15      
19 A 1193 1132 0.67      
20 A 1143 1085 1.43      
21 A 1053 1000 3.01      
22 A 1032 979 2.67      
23 A 977 927 1.59      
24 A 967 918 1.50      
25 A 932 884 5.89      
26 A 877 833 31.07      
27 A 864 820 0.33      
28 A 861 818 18.38      
29 A 798 758 11.56      
30 A 783 743 0.80      
31 A 682 648 10.23      
32 A 632 600 0.29      
33 A 513 487 11.11      
34 A 358 339 0.70      
35 A 288 273 0.53      
36 A 28 26 0.01      

Unscaled Zero Point Vibrational Energy (zpe) 26949.0 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 25580.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.24382 0.11514 0.08052

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.250 -1.144 -0.360
H2 -0.046 -2.184 -0.091
H3 -0.502 -1.093 -1.427
C4 -1.445 -0.500 0.364
H5 -1.250 -0.632 1.431
H6 -2.429 -0.892 0.102
C7 2.092 -0.072 0.227
H8 2.533 0.858 0.377
H9 2.677 -0.920 0.349
C10 0.779 -0.092 -0.095
C11 -1.212 0.910 0.011
H12 -1.936 1.647 -0.027
C13 0.102 1.100 -0.201
H14 0.553 2.009 -0.394

Atom - Atom Distances (Å)
  C1 H2 H3 C4 H5 H6 C7 H8 H9 C10 C11 H12 C13 H14
C11.09351.09721.53872.11522.24222.64163.50583.01931.49442.29853.27792.27633.2536
H21.09351.78442.23562.48502.71783.02154.01453.03342.24783.30804.27233.28834.2462
H31.09721.78442.10872.99042.46813.24214.03383.64552.10182.56623.39462.58383.4357
C41.53872.23562.10871.09331.09063.56584.20324.14302.30691.47272.23732.29583.2959
H52.11522.48502.99041.09331.79543.59654.20004.08332.59542.09682.79152.73663.6820
H62.24222.71782.46811.09061.79544.59665.26825.11163.31162.17662.58943.23434.1894
C72.64163.02153.24213.56583.59654.59661.03971.03641.35263.45374.38752.34932.6626
H83.50584.01454.03384.20324.20005.26821.03971.78322.04983.76264.55622.51062.4171
H93.01933.03343.64554.14304.08335.11161.03641.78322.11734.31065.29223.31813.6935
C101.49442.24782.10182.30692.59543.31161.35262.04982.11732.23123.22511.37502.1349
C112.29853.30802.56621.47272.09682.17663.45373.76264.31062.23121.03411.34372.1176
H123.27794.27233.39462.23732.79152.58944.38754.55625.29223.22511.03412.11722.5416
C132.27633.28832.58382.29582.73663.23432.34932.51063.31811.37501.34372.11721.0336
H143.25364.24623.43573.29593.68204.18942.66262.41713.69352.13492.11762.54161.0336

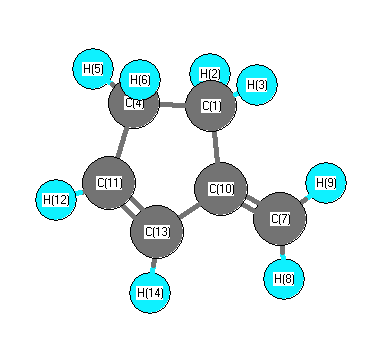
picture of 3-Methylenecyclopentene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C4 H5 105.713 C1 C4 H6 115.983
C1 C4 C11 99.486 C1 C10 C7 136.140
C1 C10 C13 104.912 H2 C1 H3 109.076
H2 C1 C4 115.237 H2 C1 C10 119.788
H3 C1 C4 105.019 H3 C1 C10 107.391
C4 C1 C10 99.015 C4 C11 H12 125.477
C4 C11 C13 109.122 H5 C4 H6 110.589
H5 C4 C11 108.703 H6 C4 C11 115.428
C7 C10 C13 118.925 H8 C7 H9 118.386
H8 C7 C10 117.324 H9 C7 C10 124.281
C10 C13 C11 110.306 C10 C13 H14 124.235
C11 C13 H14 125.444 H12 C11 C13 125.354
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C -0.352      
2 H 0.170      
3 H 0.170      
4 C -0.328      
5 H 0.161      
6 H 0.161      
7 C -0.447      
8 H 0.150      
9 H 0.150      
10 C 0.147      
11 C -0.162      
12 H 0.159      
13 C -0.136      
14 H 0.158      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 13.435 -0.002 0.000
y -0.002 8.514 0.000
z 0.000 0.000 4.851


<r2> (average value of r2) Å2
<r2> 153.305
(<r2>)1/2 12.382