return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H6O (3-Methylfuran)

using model chemistry: CID/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at CID/6-31G*
 hartrees
Energy at 0K-268.360202
Energy at 298.15K-268.367017
HF Energy-267.663864
Nuclear repulsion energy222.355836
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3419 3158 1.33      
2 A' 3407 3148 1.46      
3 A' 3376 3119 1.70      
4 A' 3249 3001 19.66      
5 A' 3166 2924 28.91      
6 A' 1772 1637 1.14      
7 A' 1655 1529 21.42      
8 A' 1599 1477 5.86      
9 A' 1525 1409 2.72      
10 A' 1502 1387 3.29      
11 A' 1359 1255 4.68      
12 A' 1347 1244 1.29      
13 A' 1301 1202 25.29      
14 A' 1164 1075 10.80      
15 A' 1124 1039 39.08      
16 A' 1077 994 2.64      
17 A' 1024 946 1.35      
18 A' 932 861 25.74      
19 A' 672 621 3.19      
20 A' 331 306 0.94      
21 A" 3232 2986 21.51      
22 A" 1582 1461 5.94      
23 A" 1129 1043 1.38      
24 A" 897 828 0.54      
25 A" 838 774 54.83      
26 A" 793 732 6.74      
27 A" 644 595 12.02      
28 A" 637 589 11.92      
29 A" 276 255 2.54      
30 A" 138 128 0.00      

Unscaled Zero Point Vibrational Energy (zpe) 22583.6 cm-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 20860.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CID/6-31G*
ABC
0.30114 0.11350 0.08370

See section I.F.4 to change rotational constant units
Geometric Data calculated at CID/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
O1 -0.698 -1.446 0.000
C2 -1.072 -0.144 0.000
C3 0.652 -1.459 0.000
C4 0.000 0.673 0.000
C5 1.140 -0.204 0.000
C6 0.010 2.167 0.000
H7 -2.130 0.040 0.000
H8 1.114 -2.428 0.000
H9 2.177 0.080 0.000
H10 -1.003 2.563 0.000
H11 0.522 2.557 0.879
H12 0.522 2.557 -0.879

Atom - Atom Distances (Å)
  O1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12
O11.35531.35062.23152.21863.68252.06412.06053.25564.02124.27704.2770
C21.35532.16871.34792.21322.55201.07383.16143.25742.70783.25743.2574
C31.35062.16872.22931.34603.68233.16051.07332.16644.34914.11324.1132
C42.23151.34792.22931.43851.49422.22223.29472.25672.13972.14372.1437
C52.21862.21321.34601.43852.62673.27932.22381.07563.50002.96322.9632
C63.68252.55203.68231.49422.62673.01774.72563.00871.08781.08911.0891
H72.06411.07383.16052.22223.27933.01774.07584.30782.76353.76083.7608
H82.06053.16141.07333.29472.22384.72564.07582.72425.42125.09665.0966
H93.25563.25742.16642.25671.07563.00874.30782.72424.03483.10643.1064
H104.02122.70784.34912.13973.50001.08782.76355.42124.03481.75991.7599
H114.27703.25744.11322.14372.96321.08913.76085.09663.10641.75991.7572
H124.27703.25744.11322.14372.96321.08913.76085.09663.10641.75991.7572

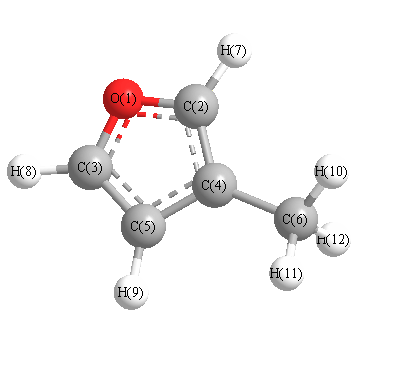
picture of 3-Methylfuran state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
O1 C2 C4 111.276 O1 C2 H7 115.881
O1 C3 C5 110.728 O1 C3 H8 115.968
C2 O1 C3 106.540 C2 C4 C5 105.125
C2 C4 C6 127.697 C3 C5 C4 106.331
C3 C5 H9 126.568 C4 C2 H7 132.843
C4 C5 H9 127.101 C4 C6 H10 110.949
C4 C6 H11 111.198 C4 C6 H12 111.198
C5 C3 H8 133.304 C5 C4 C6 127.178
H10 C6 H11 107.889 H10 C6 H12 107.889
H11 C6 H12 107.558
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability