return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H6O (3-Methylfuran)

using model chemistry: QCISD/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at QCISD/6-31G*
 hartrees
Energy at 0K-268.519157
Energy at 298.15K-268.525733
HF Energy-267.661427
Nuclear repulsion energy220.470028
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3310 3152 1.71      
2 A' 3299 3141 1.80      
3 A' 3269 3113 1.96      
4 A' 3144 2994 19.50      
5 A' 3063 2917 28.41      
6 A' 1703 1621 0.67      
7 A' 1592 1516 16.99      
8 A' 1551 1477 5.49      
9 A' 1473 1403 2.20      
10 A' 1448 1378 3.94      
11 A' 1309 1246 2.51      
12 A' 1295 1233 1.16      
13 A' 1239 1179 25.95      
14 A' 1119 1066 7.28      
15 A' 1070 1019 34.75      
16 A' 1040 990 1.31      
17 A' 990 942 1.39      
18 A' 894 852 23.62      
19 A' 649 618 2.88      
20 A' 321 305 0.87      
21 A" 3128 2979 21.45      
22 A" 1533 1460 5.69      
23 A" 1088 1036 1.05      
24 A" 808 769 0.90      
25 A" 768 731 61.37      
26 A" 721 686 0.37      
27 A" 610 581 17.67      
28 A" 592 563 1.52      
29 A" 262 249 2.32      
30 A" 129 122 0.00      

Unscaled Zero Point Vibrational Energy (zpe) 21705.7 cm-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 20670.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/6-31G*
ABC
0.29571 0.11184 0.08240

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
O1 -0.705 -1.463 0.000
C2 -1.082 -0.144 0.000
C3 0.663 -1.467 0.000
C4 0.000 0.680 0.000
C5 1.148 -0.199 0.000
C6 0.009 2.180 0.000
H7 -2.149 0.037 0.000
H8 1.128 -2.444 0.000
H9 2.193 0.092 0.000
H10 -1.013 2.577 0.000
H11 0.524 2.574 0.886
H12 0.524 2.574 -0.886

Atom - Atom Distances (Å)
  O1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12
O11.37241.36852.25622.24353.71292.08222.07903.28884.05214.31264.3126
C21.37242.19011.35942.23072.56711.08233.18963.28272.72163.27873.2787
C31.36852.19012.24701.35733.70553.18911.08172.18394.37774.13964.1396
C42.25621.35942.24701.44631.50042.24273.32112.27002.15072.15602.1560
C52.24352.23071.35731.44632.63823.30552.24461.08413.51852.97792.9779
C63.71292.56713.70551.50042.63823.04124.75753.02121.09651.09761.0976
H72.08221.08233.18912.24273.30553.04124.11014.34162.78223.79013.7901
H82.07903.18961.08173.32112.24464.75754.11012.75035.45835.13145.1314
H93.28883.28272.18392.27001.08413.02124.34162.75034.05603.11933.1193
H104.05212.72164.37772.15073.51851.09652.78225.45834.05601.77391.7739
H114.31263.27874.13962.15602.97791.09763.79015.13143.11931.77391.7712
H124.31263.27874.13962.15602.97791.09763.79015.13143.11931.77391.7712

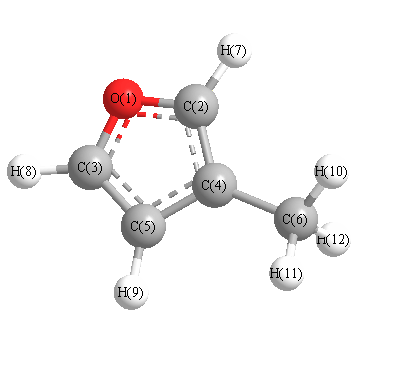
picture of 3-Methylfuran state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
O1 C2 C4 111.361 O1 C2 H7 115.541
O1 C3 C5 110.784 O1 C3 H8 115.594
C2 O1 C3 106.074 C2 C4 C5 105.281
C2 C4 C6 127.631 C3 C5 C4 106.499
C3 C5 H9 126.530 C4 C2 H7 133.098
C4 C5 H9 126.971 C4 C6 H10 110.870
C4 C6 H11 111.220 C4 C6 H12 111.220
C5 C3 H8 133.622 C5 C4 C6 127.088
H10 C6 H11 107.898 H10 C6 H12 107.898
H11 C6 H12 107.578
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability