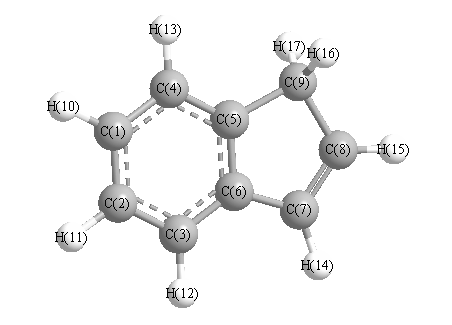
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3169 |
3116 |
16.92 |
|
|
|
| 2 |
A' |
3144 |
3092 |
7.04 |
|
|
|
| 3 |
A' |
3137 |
3085 |
27.01 |
|
|
|
| 4 |
A' |
3124 |
3073 |
32.11 |
|
|
|
| 5 |
A' |
3113 |
3062 |
2.79 |
|
|
|
| 6 |
A' |
3107 |
3055 |
5.80 |
|
|
|
| 7 |
A' |
2966 |
2917 |
12.68 |
|
|
|
| 8 |
A' |
1623 |
1596 |
2.04 |
|
|
|
| 9 |
A' |
1609 |
1582 |
0.60 |
|
|
|
| 10 |
A' |
1574 |
1548 |
0.42 |
|
|
|
| 11 |
A' |
1465 |
1441 |
7.04 |
|
|
|
| 12 |
A' |
1462 |
1438 |
5.22 |
|
|
|
| 13 |
A' |
1412 |
1389 |
9.63 |
|
|
|
| 14 |
A' |
1393 |
1370 |
1.51 |
|
|
|
| 15 |
A' |
1335 |
1313 |
3.77 |
|
|
|
| 16 |
A' |
1286 |
1265 |
0.18 |
|
|
|
| 17 |
A' |
1227 |
1206 |
1.62 |
|
|
|
| 18 |
A' |
1208 |
1188 |
3.60 |
|
|
|
| 19 |
A' |
1160 |
1141 |
0.96 |
|
|
|
| 20 |
A' |
1153 |
1134 |
0.07 |
|
|
|
| 21 |
A' |
1110 |
1092 |
0.55 |
|
|
|
| 22 |
A' |
1061 |
1044 |
1.09 |
|
|
|
| 23 |
A' |
1028 |
1011 |
3.17 |
|
|
|
| 24 |
A' |
950 |
934 |
9.03 |
|
|
|
| 25 |
A' |
849 |
835 |
2.67 |
|
|
|
| 26 |
A' |
822 |
809 |
1.50 |
|
|
|
| 27 |
A' |
725 |
714 |
2.48 |
|
|
|
| 28 |
A' |
585 |
575 |
1.55 |
|
|
|
| 29 |
A' |
529 |
520 |
0.15 |
|
|
|
| 30 |
A' |
377 |
370 |
1.28 |
|
|
|
| 31 |
A" |
2994 |
2945 |
8.72 |
|
|
|
| 32 |
A" |
1111 |
1092 |
1.57 |
|
|
|
| 33 |
A" |
938 |
923 |
0.25 |
|
|
|
| 34 |
A" |
923 |
908 |
1.32 |
|
|
|
| 35 |
A" |
914 |
899 |
1.93 |
|
|
|
| 36 |
A" |
890 |
876 |
2.27 |
|
|
|
| 37 |
A" |
840 |
826 |
0.64 |
|
|
|
| 38 |
A" |
761 |
748 |
41.39 |
|
|
|
| 39 |
A" |
711 |
699 |
17.23 |
|
|
|
| 40 |
A" |
684 |
672 |
14.36 |
|
|
|
| 41 |
A" |
545 |
536 |
3.93 |
|
|
|
| 42 |
A" |
413 |
406 |
6.17 |
|
|
|
| 43 |
A" |
383 |
377 |
4.92 |
|
|
|
| 44 |
A" |
207 |
204 |
3.21 |
|
|
|
| 45 |
A" |
192 |
189 |
0.30 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30103.5 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 29606.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.146 |
|
|
|
| 2 |
C |
-0.143 |
|
|
|
| 3 |
C |
-0.207 |
|
|
|
| 4 |
C |
-0.196 |
|
|
|
| 5 |
C |
0.106 |
|
|
|
| 6 |
C |
0.123 |
|
|
|
| 7 |
C |
-0.143 |
|
|
|
| 8 |
C |
-0.150 |
|
|
|
| 9 |
C |
-0.424 |
|
|
|
| 10 |
H |
0.133 |
|
|
|
| 11 |
H |
0.134 |
|
|
|
| 12 |
H |
0.133 |
|
|
|
| 13 |
H |
0.130 |
|
|
|
| 14 |
H |
0.137 |
|
|
|
| 15 |
H |
0.141 |
|
|
|
| 16 |
H |
0.187 |
|
|
|
| 17 |
H |
0.187 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.428 |
0.581 |
0.000 |
0.722 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-47.207 |
0.651 |
0.000 |
| y |
0.651 |
-45.997 |
0.000 |
| z |
0.000 |
0.000 |
-55.694 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.638 |
0.651 |
0.000 |
| y |
0.651 |
5.453 |
0.000 |
| z |
0.000 |
0.000 |
-9.092 |
|
| Polar |
|---|
| 3z2-r2 | -18.183 |
|---|
| x2-y2 | -1.210 |
|---|
| xy | 0.651 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
18.577 |
1.717 |
0.000 |
| y |
1.717 |
15.598 |
0.000 |
| z |
0.000 |
0.000 |
5.396 |
<r2> (average value of r
2) Å
2
| <r2> |
295.827 |
| (<r2>)1/2 |
17.200 |