return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H4Cl2 (1,2-dichlorobenzene)

using model chemistry: QCISD/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at QCISD/6-31G*
 hartrees
Energy at 0K-1149.567373
Energy at 298.15K-1149.571476
HF Energy-1148.495428
Nuclear repulsion energy468.059207
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3236 3082 6.54      
2 A1 3221 3067 7.25      
3 A1 1663 1584 3.50      
4 A1 1534 1461 58.65      
5 A1 1289 1227 0.79      
6 A1 1208 1151 2.09      
7 A1 1186 1130 30.73      
8 A1 1080 1029 6.08      
9 A1 682 649 13.94      
10 A1 491 467 5.39      
11 A1 207 197 0.00      
12 A2 916 872 0.00      
13 A2 847 807 0.00      
14 A2 566 539 0.00      
15 A2 503 479 0.00      
16 A2 138 131 0.00      
17 B1 914 870 1.42      
18 B1 763 727 65.45      
19 B1 440 419 3.82      
20 B1 236 225 0.77      
21 B2 3232 3077 5.27      
22 B2 3208 3055 1.22      
23 B2 1676 1596 6.02      
24 B2 1495 1424 19.43      
25 B2 1306 1243 2.71      
26 B2 1180 1124 2.48      
27 B2 1062 1011 34.27      
28 B2 765 729 14.00      
29 B2 437 416 0.38      
30 B2 344 327 0.10      

Unscaled Zero Point Vibrational Energy (zpe) 17911.4 cm-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 17057.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/6-31G*
ABC
0.06318 0.04750 0.02711

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/6-31G*

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.700 -0.032
C2 0.000 -0.700 -0.032
C3 0.000 1.394 1.182
C4 0.000 -1.394 1.182
C5 0.000 0.698 2.389
C6 0.000 -0.698 2.389
Cl7 0.000 1.609 -1.514
Cl8 0.000 -1.609 -1.514
H9 0.000 2.481 1.165
H10 0.000 -2.481 1.165
H11 0.000 1.250 3.328
H12 0.000 -1.250 3.328

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 Cl7 Cl8 H9 H10 H11 H12
C11.39951.39812.42002.42092.79571.73902.74392.14633.39883.40393.8839
C21.39952.42001.39812.79572.42092.74391.73903.39882.14633.88393.4039
C31.39812.42002.78811.39352.41592.70414.03571.08743.87552.15053.4050
C42.42001.39812.78812.41591.39354.03572.70413.87551.08743.40502.1505
C52.42092.79571.39352.41591.39704.00794.53432.16273.40731.08822.1624
C62.79572.42092.41591.39351.39704.53434.00793.40732.16272.16241.0882
Cl71.73902.74392.70414.03574.00794.53433.21882.81734.88994.85465.6225
Cl82.74391.73904.03572.70414.53434.00793.21884.88992.81735.62254.8546
H92.14633.39881.08743.87552.16273.40732.81734.88994.96282.48844.3125
H103.39882.14633.87551.08743.40732.16274.88992.81734.96284.31252.4884
H113.40393.88392.15053.40501.08822.16244.85465.62252.48844.31252.4996
H123.88393.40393.40502.15052.16241.08825.62254.85464.31252.48842.4996

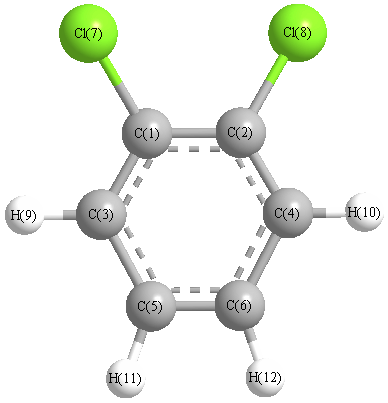
picture of 1,2-dichlorobenzene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 119.777 C1 C2 Cl8 121.540
C1 C3 C5 120.278 C1 C3 H9 118.901
C2 C1 C3 119.777 C2 C1 Cl7 121.540
C2 C4 C6 120.278 C2 C4 H10 118.901
C3 C1 Cl7 118.683 C3 C5 C6 119.945
C3 C5 H11 119.615 C4 C2 Cl8 118.683
C4 C6 C5 119.945 C4 C6 H12 119.615
C5 C3 H9 120.821 C5 C6 H12 120.440
C6 C4 H10 120.821 C6 C5 H11 120.440
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability