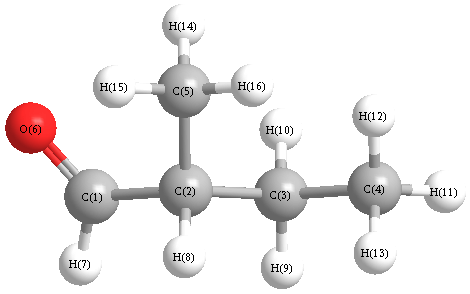
Vibrational Frequencies calculated at HSEh1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3173 |
3018 |
15.74 |
|
|
|
| 2 |
A |
3163 |
3008 |
21.18 |
|
|
|
| 3 |
A |
3155 |
3001 |
30.74 |
|
|
|
| 4 |
A |
3147 |
2993 |
33.86 |
|
|
|
| 5 |
A |
3094 |
2942 |
19.91 |
|
|
|
| 6 |
A |
3085 |
2934 |
13.64 |
|
|
|
| 7 |
A |
3071 |
2920 |
23.06 |
|
|
|
| 8 |
A |
3051 |
2902 |
29.54 |
|
|
|
| 9 |
A |
3009 |
2861 |
17.64 |
|
|
|
| 10 |
A |
2907 |
2765 |
125.40 |
|
|
|
| 11 |
A |
1865 |
1773 |
167.43 |
|
|
|
| 12 |
A |
1536 |
1461 |
4.20 |
|
|
|
| 13 |
A |
1529 |
1454 |
10.46 |
|
|
|
| 14 |
A |
1523 |
1449 |
6.31 |
|
|
|
| 15 |
A |
1519 |
1445 |
9.34 |
|
|
|
| 16 |
A |
1509 |
1435 |
0.70 |
|
|
|
| 17 |
A |
1451 |
1380 |
9.56 |
|
|
|
| 18 |
A |
1441 |
1371 |
3.39 |
|
|
|
| 19 |
A |
1428 |
1358 |
4.38 |
|
|
|
| 20 |
A |
1395 |
1326 |
4.78 |
|
|
|
| 21 |
A |
1377 |
1310 |
1.64 |
|
|
|
| 22 |
A |
1325 |
1260 |
2.01 |
|
|
|
| 23 |
A |
1287 |
1224 |
0.36 |
|
|
|
| 24 |
A |
1208 |
1149 |
2.04 |
|
|
|
| 25 |
A |
1184 |
1126 |
3.70 |
|
|
|
| 26 |
A |
1135 |
1079 |
3.00 |
|
|
|
| 27 |
A |
1067 |
1015 |
0.03 |
|
|
|
| 28 |
A |
1032 |
981 |
13.23 |
|
|
|
| 29 |
A |
987 |
939 |
8.53 |
|
|
|
| 30 |
A |
949 |
902 |
9.01 |
|
|
|
| 31 |
A |
923 |
878 |
13.33 |
|
|
|
| 32 |
A |
805 |
766 |
6.49 |
|
|
|
| 33 |
A |
788 |
750 |
3.01 |
|
|
|
| 34 |
A |
663 |
631 |
7.57 |
|
|
|
| 35 |
A |
406 |
387 |
0.93 |
|
|
|
| 36 |
A |
391 |
372 |
0.69 |
|
|
|
| 37 |
A |
290 |
276 |
5.42 |
|
|
|
| 38 |
A |
267 |
254 |
0.39 |
|
|
|
| 39 |
A |
231 |
220 |
2.72 |
|
|
|
| 40 |
A |
195 |
185 |
0.51 |
|
|
|
| 41 |
A |
95 |
90 |
2.36 |
|
|
|
| 42 |
A |
75 |
71 |
3.31 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31363.6 cm
-1
Scaled (by 0.951) Zero Point Vibrational Energy (zpe) 29826.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.246 |
|
|
|
| 2 |
C |
-0.216 |
|
|
|
| 3 |
C |
-0.325 |
|
|
|
| 4 |
C |
-0.516 |
|
|
|
| 5 |
C |
-0.514 |
|
|
|
| 6 |
O |
-0.387 |
|
|
|
| 7 |
H |
0.123 |
|
|
|
| 8 |
H |
0.179 |
|
|
|
| 9 |
H |
0.167 |
|
|
|
| 10 |
H |
0.176 |
|
|
|
| 11 |
H |
0.177 |
|
|
|
| 12 |
H |
0.178 |
|
|
|
| 13 |
H |
0.170 |
|
|
|
| 14 |
H |
0.181 |
|
|
|
| 15 |
H |
0.190 |
|
|
|
| 16 |
H |
0.170 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.561 |
-0.456 |
0.768 |
2.712 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-45.383 |
-0.031 |
-1.707 |
| y |
-0.031 |
-36.771 |
-0.273 |
| z |
-1.707 |
-0.273 |
-36.990 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-8.503 |
-0.031 |
-1.707 |
| y |
-0.031 |
4.416 |
-0.273 |
| z |
-1.707 |
-0.273 |
4.087 |
|
| Polar |
|---|
| 3z2-r2 | 8.173 |
|---|
| x2-y2 | -8.612 |
|---|
| xy | -0.031 |
|---|
| xz | -1.707 |
|---|
| yz | -0.273 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.450 |
-0.236 |
0.272 |
| y |
-0.236 |
8.094 |
-0.245 |
| z |
0.272 |
-0.245 |
6.826 |
<r2> (average value of r
2) Å
2
| <r2> |
203.074 |
| (<r2>)1/2 |
14.250 |