return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H10O (3-Pentanone)

using model chemistry: QCISD/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at QCISD/6-31G*
 hartrees
Energy at 0K-270.919033
Energy at 298.15K 
HF Energy-270.030932
Nuclear repulsion energy238.511667
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3155 3005 32.26      
2 A1 3081 2934 1.32      
3 A1 3056 2910 27.02      
4 A1 1815 1728 87.49      
5 A1 1548 1474 8.37      
6 A1 1526 1453 11.01      
7 A1 1470 1400 9.84      
8 A1 1402 1335 3.39      
9 A1 1149 1094 1.62      
10 A1 1050 1000 2.72      
11 A1 805 767 0.97      
12 A1 418 398 0.39      
13 A1 201 191 0.52      
14 A2 3166 3015 0.00      
15 A2 3080 2933 0.00      
16 A2 1544 1471 0.00      
17 A2 1298 1236 0.00      
18 A2 1034 985 0.00      
19 A2 741 706 0.00      
20 A2 225 214 0.00      
21 A2 32 30 0.00      
22 B1 3166 3015 44.82      
23 B1 3090 2943 23.00      
24 B1 1545 1471 11.35      
25 B1 1336 1272 1.18      
26 B1 1177 1121 0.09      
27 B1 842 802 9.11      
28 B1 469 447 0.05      
29 B1 208 198 0.27      
30 B1 64 61 0.20      
31 B2 3155 3005 14.91      
32 B2 3080 2933 40.79      
33 B2 3047 2902 8.18      
34 B2 1548 1474 10.24      
35 B2 1515 1442 1.16      
36 B2 1470 1400 0.12      
37 B2 1443 1374 34.78      
38 B2 1178 1121 57.23      
39 B2 1051 1000 6.61      
40 B2 998 951 13.88      
41 B2 637 607 6.32      
42 B2 321 306 13.30      

Unscaled Zero Point Vibrational Energy (zpe) 31567.0 cm-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 30061.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/6-31G*
ABC
0.29556 0.06514 0.05557

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/6-31G*

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
O1 0.000 0.000 1.285
C2 0.000 0.000 0.061
C3 0.000 1.296 -0.739
C4 0.000 -1.296 -0.739
C5 0.000 2.543 0.139
C6 0.000 -2.543 0.139
H7 0.876 1.282 -1.406
H8 -0.876 1.282 -1.406
H9 -0.876 -1.282 -1.406
H10 0.876 -1.282 -1.406
H11 0.000 3.449 -0.479
H12 -0.883 2.563 0.786
H13 0.883 2.563 0.786
H14 0.000 -3.449 -0.479
H15 0.883 -2.563 0.786
H16 -0.883 -2.563 0.786

Atom - Atom Distances (Å)
  O1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16
O11.22392.40282.40282.78922.78923.10663.10663.10663.10663.87372.75592.75593.87372.75592.7559
C21.22391.52251.52252.54422.54422.13582.13582.13582.13583.49082.80592.80593.49082.80592.8059
C32.40281.52252.59101.52533.93761.10111.10112.80242.80242.16882.17062.17064.75144.24164.2416
C42.40281.52252.59103.93761.52532.80242.80241.10111.10114.75144.24164.24162.16882.17062.1706
C52.78922.54421.52533.93765.08602.17842.17844.21684.21681.09661.09511.09516.02355.22175.2217
C62.78922.54423.93761.52535.08604.21684.21682.17842.17846.02355.22175.22171.09661.09511.0951
H73.10662.13581.10112.80242.17844.21681.75123.10432.56322.51453.08892.53964.89924.42564.7622
H83.10662.13581.10112.80242.17844.21681.75122.56323.10432.51452.53963.08894.89924.76224.4256
H93.10662.13582.80241.10114.21682.17843.10432.56321.75124.89924.42564.76222.51453.08892.5396
H103.10662.13582.80241.10114.21682.17842.56323.10431.75124.89924.76224.42562.51452.53963.0889
H113.87373.49082.16884.75141.09666.02352.51452.51454.89924.89921.77951.77956.89756.20636.2063
H122.75592.80592.17064.24161.09515.22173.08892.53964.42564.76221.77951.76586.20635.42095.1252
H132.75592.80592.17064.24161.09515.22172.53963.08894.76224.42561.77951.76586.20635.12525.4209
H143.87373.49084.75142.16886.02351.09664.89924.89922.51452.51456.89756.20636.20631.77951.7795
H152.75592.80594.24162.17065.22171.09514.42564.76223.08892.53966.20635.42095.12521.77951.7658
H162.75592.80594.24162.17065.22171.09514.76224.42562.53963.08896.20635.12525.42091.77951.7658

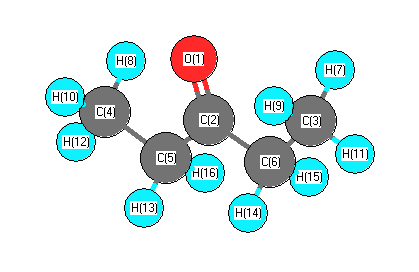
picture of 3-Pentanone state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
O1 C2 C3 121.687 O1 C2 C4 121.687
C2 C3 C5 113.182 C2 C3 H7 107.918
C2 C3 H8 107.918 C2 C4 C6 113.182
C2 C4 H9 107.918 C2 C4 H10 107.918
C3 C2 C4 116.625 C3 C5 H11 110.558
C3 C5 H12 110.791 C3 C5 H13 110.791
C4 C6 H14 110.558 C4 C6 H15 110.791
C4 C6 H16 110.791 C5 C3 H7 111.050
C5 C3 H8 111.050 C6 C4 H9 111.050
C6 C4 H10 111.050 H7 C3 H8 105.351
H9 C4 H10 105.351 H11 C5 H12 108.574
H11 C5 H13 108.574 H12 C5 H13 107.456
H14 C6 H15 108.574 H14 C6 H16 108.574
H15 C6 H16 107.456
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability