return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H10O (3-Pentanone)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-271.460427
Energy at 298.15K 
HF Energy-271.186927
Nuclear repulsion energy238.695750
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3172 3011 31.66      
2 A1 3098 2940 2.02      
3 A1 3067 2912 26.62      
4 A1 1792 1701 89.38      
5 A1 1549 1470 6.86      
6 A1 1515 1438 12.74      
7 A1 1462 1388 9.31      
8 A1 1390 1319 5.85      
9 A1 1140 1082 1.87      
10 A1 1040 987 2.04      
11 A1 795 755 1.43      
12 A1 413 392 0.40      
13 A1 198 188 0.52      
14 A2 3181 3019 0.00      
15 A2 3089 2933 0.00      
16 A2 1544 1466 0.00      
17 A2 1294 1228 0.00      
18 A2 1027 975 0.00      
19 A2 733 696 0.00      
20 A2 214 203 0.00      
21 A2 33 31 0.00      
22 B1 3181 3020 43.59      
23 B1 3100 2943 23.06      
24 B1 1544 1466 11.39      
25 B1 1334 1266 1.01      
26 B1 1168 1109 0.09      
27 B1 838 795 10.11      
28 B1 469 445 0.00      
29 B1 193 183 0.28      
30 B1 66 63 0.16      
31 B2 3172 3010 14.75      
32 B2 3097 2940 42.15      
33 B2 3058 2903 9.55      
34 B2 1548 1470 9.35      
35 B2 1504 1428 2.14      
36 B2 1461 1387 0.35      
37 B2 1424 1352 28.66      
38 B2 1164 1105 65.81      
39 B2 1037 984 9.60      
40 B2 987 937 16.64      
41 B2 630 598 4.56      
42 B2 315 299 13.20      

Unscaled Zero Point Vibrational Energy (zpe) 31518.4 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 29917.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.29655 0.06511 0.05556

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
O1 0.000 0.000 1.284
C2 0.000 0.000 0.062
C3 0.000 1.296 -0.738
C4 0.000 -1.296 -0.738
C5 0.000 2.544 0.137
C6 0.000 -2.544 0.137
H7 0.872 1.281 -1.404
H8 -0.872 1.281 -1.404
H9 -0.872 -1.281 -1.404
H10 0.872 -1.281 -1.404
H11 0.000 3.447 -0.479
H12 -0.880 2.565 0.784
H13 0.880 2.565 0.784
H14 0.000 -3.447 -0.479
H15 0.880 -2.565 0.784
H16 -0.880 -2.565 0.784

Atom - Atom Distances (Å)
  O1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16
O11.22152.40082.40082.79052.79053.10273.10273.10273.10273.87132.75722.75723.87132.75722.7572
C21.22151.52261.52262.54532.54532.13362.13362.13362.13363.48902.80592.80593.48902.80592.8059
C32.40081.52262.59151.52453.93841.09791.09792.80102.80102.16662.16772.16774.74964.24184.2418
C42.40081.52262.59153.93841.52452.80102.80101.09791.09794.74964.24184.24182.16662.16772.1677
C52.79052.54531.52453.93845.08842.17552.17554.21554.21551.09291.09181.09186.02275.22445.2244
C62.79052.54533.93841.52455.08844.21554.21552.17552.17556.02275.22445.22441.09291.09181.0918
H73.10272.13361.09792.80102.17554.21551.74413.09952.56222.51143.08292.53704.89604.42504.7590
H83.10272.13361.09792.80102.17554.21551.74412.56223.09952.51142.53703.08294.89604.75904.4250
H93.10272.13362.80101.09794.21552.17553.09952.56221.74414.89604.42504.75902.51143.08292.5370
H103.10272.13362.80101.09794.21552.17552.56223.09951.74414.89604.75904.42502.51142.53703.0829
H113.87133.48902.16664.74961.09296.02272.51142.51144.89604.89601.77371.77376.89376.20566.2056
H122.75722.80592.16774.24181.09185.22443.08292.53704.42504.75901.77371.75916.20565.42305.1298
H132.75722.80592.16774.24181.09185.22442.53703.08294.75904.42501.77371.75916.20565.12985.4230
H143.87133.48904.74962.16666.02271.09294.89604.89602.51142.51146.89376.20566.20561.77371.7737
H152.75722.80594.24182.16775.22441.09184.42504.75903.08292.53706.20565.42305.12981.77371.7591
H162.75722.80594.24182.16775.22441.09184.75904.42502.53703.08296.20565.12985.42301.77371.7591

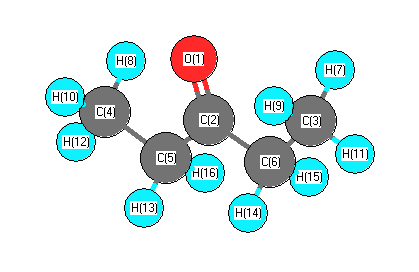
picture of 3-Pentanone state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
O1 C2 C3 121.680 O1 C2 C4 121.680
C2 C3 C5 113.299 C2 C3 H7 107.918
C2 C3 H8 107.918 C2 C4 C6 113.299
C2 C4 H9 107.918 C2 C4 H10 107.918
C3 C2 C4 116.641 C3 C5 H11 110.655
C3 C5 H12 110.814 C3 C5 H13 110.814
C4 C6 H14 110.655 C4 C6 H15 110.814
C4 C6 H16 110.814 C5 C3 H7 111.069
C5 C3 H8 111.069 C6 C4 H9 111.069
C6 C4 H10 111.069 H7 C3 H8 105.176
H9 C4 H10 105.176 H11 C5 H12 108.556
H11 C5 H13 108.556 H12 C5 H13 107.339
H14 C6 H15 108.556 H14 C6 H16 108.556
H15 C6 H16 107.339
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 O -0.489      
2 C 0.487      
3 C -0.375      
4 C -0.375      
5 C -0.450      
6 C -0.450      
7 H 0.169      
8 H 0.169      
9 H 0.169      
10 H 0.169      
11 H 0.149      
12 H 0.170      
13 H 0.170      
14 H 0.149      
15 H 0.170      
16 H 0.170      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 6.588 0.000 0.000
y 0.000 9.195 0.000
z 0.000 0.000 8.155


<r2> (average value of r2) Å2
<r2> 215.277
(<r2>)1/2 14.672