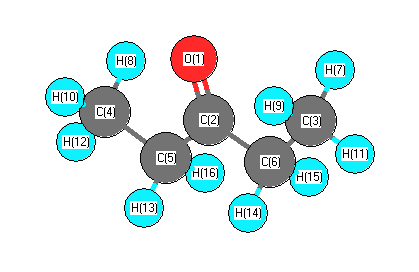
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3078 |
3027 |
27.75 |
|
|
|
| 2 |
A1 |
3002 |
2953 |
3.40 |
|
|
|
| 3 |
A1 |
2964 |
2915 |
26.70 |
|
|
|
| 4 |
A1 |
1750 |
1721 |
95.47 |
|
|
|
| 5 |
A1 |
1481 |
1456 |
6.67 |
|
|
|
| 6 |
A1 |
1440 |
1416 |
14.27 |
|
|
|
| 7 |
A1 |
1390 |
1367 |
11.35 |
|
|
|
| 8 |
A1 |
1317 |
1295 |
8.73 |
|
|
|
| 9 |
A1 |
1096 |
1078 |
1.39 |
|
|
|
| 10 |
A1 |
1007 |
991 |
2.23 |
|
|
|
| 11 |
A1 |
766 |
753 |
2.59 |
|
|
|
| 12 |
A1 |
400 |
394 |
0.36 |
|
|
|
| 13 |
A1 |
189 |
186 |
0.54 |
|
|
|
| 14 |
A2 |
3082 |
3031 |
0.00 |
|
|
|
| 15 |
A2 |
2985 |
2936 |
0.00 |
|
|
|
| 16 |
A2 |
1476 |
1451 |
0.00 |
|
|
|
| 17 |
A2 |
1232 |
1211 |
0.00 |
|
|
|
| 18 |
A2 |
974 |
958 |
0.00 |
|
|
|
| 19 |
A2 |
693 |
682 |
0.00 |
|
|
|
| 20 |
A2 |
203 |
200 |
0.00 |
|
|
|
| 21 |
A2 |
21 |
20 |
0.00 |
|
|
|
| 22 |
B1 |
3082 |
3032 |
38.76 |
|
|
|
| 23 |
B1 |
2995 |
2946 |
20.44 |
|
|
|
| 24 |
B1 |
1476 |
1451 |
13.81 |
|
|
|
| 25 |
B1 |
1272 |
1251 |
0.68 |
|
|
|
| 26 |
B1 |
1109 |
1091 |
0.25 |
|
|
|
| 27 |
B1 |
801 |
788 |
12.82 |
|
|
|
| 28 |
B1 |
452 |
444 |
0.45 |
|
|
|
| 29 |
B1 |
180 |
177 |
0.28 |
|
|
|
| 30 |
B1 |
63 |
62 |
0.11 |
|
|
|
| 31 |
B2 |
3078 |
3027 |
13.67 |
|
|
|
| 32 |
B2 |
3002 |
2952 |
41.18 |
|
|
|
| 33 |
B2 |
2954 |
2906 |
12.53 |
|
|
|
| 34 |
B2 |
1480 |
1456 |
11.40 |
|
|
|
| 35 |
B2 |
1426 |
1402 |
3.79 |
|
|
|
| 36 |
B2 |
1389 |
1366 |
1.72 |
|
|
|
| 37 |
B2 |
1350 |
1328 |
32.28 |
|
|
|
| 38 |
B2 |
1115 |
1097 |
64.04 |
|
|
|
| 39 |
B2 |
985 |
968 |
15.21 |
|
|
|
| 40 |
B2 |
960 |
945 |
19.56 |
|
|
|
| 41 |
B2 |
605 |
595 |
1.83 |
|
|
|
| 42 |
B2 |
300 |
295 |
12.93 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30310.5 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 29810.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.409 |
|
|
|
| 2 |
C |
0.433 |
|
|
|
| 3 |
C |
-0.381 |
|
|
|
| 4 |
C |
-0.381 |
|
|
|
| 5 |
C |
-0.476 |
|
|
|
| 6 |
C |
-0.476 |
|
|
|
| 7 |
H |
0.172 |
|
|
|
| 8 |
H |
0.172 |
|
|
|
| 9 |
H |
0.172 |
|
|
|
| 10 |
H |
0.172 |
|
|
|
| 11 |
H |
0.153 |
|
|
|
| 12 |
H |
0.174 |
|
|
|
| 13 |
H |
0.174 |
|
|
|
| 14 |
H |
0.153 |
|
|
|
| 15 |
H |
0.174 |
|
|
|
| 16 |
H |
0.174 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-2.473 |
2.473 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-36.658 |
0.000 |
0.000 |
| y |
0.000 |
-36.586 |
0.000 |
| z |
0.000 |
0.000 |
-40.372 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.821 |
0.000 |
0.000 |
| y |
0.000 |
1.929 |
0.000 |
| z |
0.000 |
0.000 |
-3.749 |
|
| Polar |
|---|
| 3z2-r2 | -7.499 |
|---|
| x2-y2 | -0.072 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.780 |
0.000 |
0.000 |
| y |
0.000 |
9.922 |
0.000 |
| z |
0.000 |
0.000 |
8.471 |
<r2> (average value of r
2) Å
2
| <r2> |
216.970 |
| (<r2>)1/2 |
14.730 |