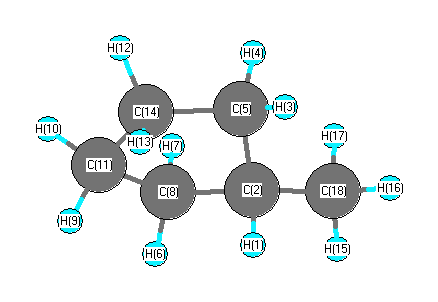
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3128 |
2992 |
99.63 |
|
|
|
| 2 |
A |
3123 |
2988 |
32.75 |
|
|
|
| 3 |
A |
3118 |
2983 |
26.85 |
|
|
|
| 4 |
A |
3109 |
2975 |
12.78 |
|
|
|
| 5 |
A |
3105 |
2970 |
5.80 |
|
|
|
| 6 |
A |
3098 |
2963 |
44.67 |
|
|
|
| 7 |
A |
3077 |
2943 |
49.54 |
|
|
|
| 8 |
A |
3064 |
2932 |
20.18 |
|
|
|
| 9 |
A |
3049 |
2917 |
78.04 |
|
|
|
| 10 |
A |
3042 |
2911 |
10.47 |
|
|
|
| 11 |
A |
3040 |
2908 |
22.84 |
|
|
|
| 12 |
A |
3016 |
2885 |
9.93 |
|
|
|
| 13 |
A |
1545 |
1478 |
1.10 |
|
|
|
| 14 |
A |
1526 |
1460 |
10.51 |
|
|
|
| 15 |
A |
1521 |
1455 |
2.49 |
|
|
|
| 16 |
A |
1519 |
1453 |
1.52 |
|
|
|
| 17 |
A |
1518 |
1453 |
1.86 |
|
|
|
| 18 |
A |
1507 |
1442 |
0.04 |
|
|
|
| 19 |
A |
1434 |
1372 |
3.34 |
|
|
|
| 20 |
A |
1407 |
1346 |
0.78 |
|
|
|
| 21 |
A |
1366 |
1307 |
1.78 |
|
|
|
| 22 |
A |
1352 |
1293 |
0.00 |
|
|
|
| 23 |
A |
1344 |
1286 |
0.03 |
|
|
|
| 24 |
A |
1333 |
1275 |
1.48 |
|
|
|
| 25 |
A |
1309 |
1252 |
0.07 |
|
|
|
| 26 |
A |
1275 |
1220 |
0.20 |
|
|
|
| 27 |
A |
1261 |
1206 |
0.67 |
|
|
|
| 28 |
A |
1229 |
1176 |
0.30 |
|
|
|
| 29 |
A |
1214 |
1162 |
0.39 |
|
|
|
| 30 |
A |
1175 |
1124 |
1.03 |
|
|
|
| 31 |
A |
1119 |
1070 |
0.02 |
|
|
|
| 32 |
A |
1066 |
1019 |
0.27 |
|
|
|
| 33 |
A |
1033 |
988 |
1.03 |
|
|
|
| 34 |
A |
1008 |
965 |
2.75 |
|
|
|
| 35 |
A |
986 |
943 |
0.93 |
|
|
|
| 36 |
A |
939 |
898 |
0.02 |
|
|
|
| 37 |
A |
929 |
889 |
0.11 |
|
|
|
| 38 |
A |
906 |
867 |
1.11 |
|
|
|
| 39 |
A |
863 |
826 |
1.27 |
|
|
|
| 40 |
A |
824 |
788 |
0.33 |
|
|
|
| 41 |
A |
763 |
730 |
0.99 |
|
|
|
| 42 |
A |
628 |
600 |
0.47 |
|
|
|
| 43 |
A |
531 |
508 |
0.64 |
|
|
|
| 44 |
A |
426 |
408 |
0.21 |
|
|
|
| 45 |
A |
301 |
288 |
0.06 |
|
|
|
| 46 |
A |
234 |
224 |
0.00 |
|
|
|
| 47 |
A |
179 |
171 |
0.01 |
|
|
|
| 48 |
A |
33 |
31 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 37283.4 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 35669.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.150 |
|
|
-0.002 |
| 2 |
C |
-0.124 |
|
|
0.313 |
| 3 |
H |
0.158 |
|
|
0.029 |
| 4 |
H |
0.154 |
|
|
0.054 |
| 5 |
C |
-0.314 |
|
|
-0.132 |
| 6 |
H |
0.158 |
|
|
0.029 |
| 7 |
H |
0.154 |
|
|
0.054 |
| 8 |
C |
-0.314 |
|
|
-0.132 |
| 9 |
H |
0.162 |
|
|
0.041 |
| 10 |
H |
0.162 |
|
|
0.045 |
| 11 |
C |
-0.325 |
|
|
-0.101 |
| 12 |
H |
0.161 |
|
|
0.045 |
| 13 |
H |
0.162 |
|
|
0.041 |
| 14 |
C |
-0.325 |
|
|
-0.100 |
| 15 |
H |
0.164 |
|
|
0.147 |
| 16 |
H |
0.164 |
|
|
0.147 |
| 17 |
H |
0.160 |
|
|
0.145 |
| 18 |
C |
-0.507 |
|
|
-0.622 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.020 |
0.000 |
-0.072 |
0.075 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
-0.026 |
-0.001 |
-0.047 |
0.054 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-40.069 |
-0.000 |
-0.187 |
| y |
-0.000 |
-39.636 |
-0.000 |
| z |
-0.187 |
-0.000 |
-38.664 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.919 |
-0.000 |
-0.187 |
| y |
-0.000 |
-0.269 |
-0.000 |
| z |
-0.187 |
-0.000 |
1.188 |
|
| Polar |
|---|
| 3z2-r2 | 2.376 |
|---|
| x2-y2 | -0.433 |
|---|
| xy | -0.000 |
|---|
| xz | -0.187 |
|---|
| yz | -0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
178.829 |
| (<r2>)1/2 |
13.373 |