return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H12 (Cyclopentane, methyl-)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-235.556339
Energy at 298.15K-235.570110
HF Energy-235.297690
Nuclear repulsion energy252.704031
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3148 2988 102.14      
2 A' 3139 2980 26.25      
3 A' 3124 2965 7.36      
4 A' 3098 2941 45.16      
5 A' 3071 2915 73.15      
6 A' 3066 2910 9.74      
7 A' 3038 2883 11.43      
8 A' 1571 1491 0.76      
9 A' 1552 1473 9.13      
10 A' 1545 1467 0.94      
11 A' 1457 1383 2.24      
12 A' 1427 1355 0.71      
13 A' 1362 1293 0.19      
14 A' 1353 1284 1.92      
15 A' 1278 1213 0.60      
16 A' 1244 1181 0.30      
17 A' 1127 1070 0.03      
18 A' 1072 1018 0.29      
19 A' 1041 988 0.65      
20 A' 952 903 0.04      
21 A' 909 863 0.88      
22 A' 869 825 0.87      
23 A' 774 735 0.63      
24 A' 537 510 0.49      
25 A' 437 415 0.16      
26 A' 182 173 0.01      
27 A" 3143 2984 33.31      
28 A" 3130 2971 8.98      
29 A" 3117 2959 45.19      
30 A" 3086 2929 19.80      
31 A" 3062 2907 21.79      
32 A" 1548 1469 2.11      
33 A" 1545 1466 0.99      
34 A" 1533 1455 0.02      
35 A" 1388 1317 1.98      
36 A" 1371 1301 0.03      
37 A" 1328 1261 0.04      
38 A" 1292 1226 0.23      
39 A" 1231 1168 0.44      
40 A" 1187 1127 0.75      
41 A" 1017 965 2.15      
42 A" 998 948 0.62      
43 A" 936 889 0.20      
44 A" 835 792 0.23      
45 A" 635 602 0.36      
46 A" 306 291 0.04      
47 A" 243 231 0.00      
48 A" 33 32 0.00      

Unscaled Zero Point Vibrational Energy (zpe) 37668.2 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 35754.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.20835 0.09752 0.07355

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.238 2.215 0.000
C2 0.457 0.705 0.000
C3 -0.167 -0.033 1.195
C4 -0.167 -0.033 -1.195
C5 -0.167 -1.520 -0.779
C6 -0.167 -1.520 0.779
H7 0.684 2.682 -0.884
H8 0.684 2.682 0.884
H9 -0.832 2.450 0.000
H10 1.538 0.505 0.000
H11 0.366 0.148 2.134
H12 0.366 0.148 -2.134
H13 -1.195 0.324 1.332
H14 -1.195 0.324 -1.332
H15 0.729 -2.020 -1.158
H16 0.729 -2.020 1.158
H17 -1.025 -2.057 -1.192
H18 -1.025 -2.057 1.192

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16 H17 H18
C11.52612.57792.57793.83703.83701.09491.09491.09562.14852.97402.97402.72162.72164.41874.41874.61194.6119
C21.52611.53631.53632.43832.43832.17782.17782.16951.09952.20722.20722.15612.15612.97392.97393.35343.3534
C32.57791.53632.38902.47081.54393.52382.86212.83482.14971.09473.37541.09752.75083.20762.18043.24482.1985
C42.57791.53632.38901.54392.47082.86213.52382.83482.14973.37541.09472.75081.09752.18043.20762.19853.2448
C53.83702.43832.47081.54391.55754.28854.59854.10022.75863.39802.21362.98512.18241.09432.19231.09362.2156
C63.83702.43831.54392.47081.55754.59854.28854.10022.75862.21363.39802.18242.98512.19231.09432.21561.0936
H71.09492.17783.52382.86214.28854.59851.76881.77032.50073.95402.84363.74243.04884.71085.12735.04725.4490
H81.09492.17782.86213.52384.59854.28851.76881.77032.50072.84363.95403.04883.74245.12734.71085.44905.0472
H91.09562.16952.83482.83484.10024.10021.77031.77033.06633.36003.36002.53532.53534.87514.87514.66624.6662
H102.14851.09952.14972.14972.75862.75862.50072.50073.06632.46042.46043.04593.04592.89332.89333.81453.8145
H112.97402.20721.09473.37543.39802.21363.95402.84363.36002.46044.26741.76423.80493.95862.40494.22552.7715
H122.97402.20723.37541.09472.21363.39802.84363.95403.36002.46044.26743.80491.76422.40493.95862.77154.2255
H132.72162.15611.09752.75082.98512.18243.74243.04882.53533.04591.76423.80492.66323.92433.03813.47352.3908
H142.72162.15612.75081.09752.18242.98513.04883.74242.53533.04593.80491.76422.66323.03813.92432.39083.4735
H154.41872.97393.20762.18041.09432.19234.71085.12734.87512.89333.95862.40493.92433.03812.31681.75502.9331
H164.41872.97392.18043.20762.19231.09435.12734.71084.87512.89332.40493.95863.03813.92432.31682.93311.7550
H174.61193.35343.24482.19851.09362.21565.04725.44904.66623.81454.22552.77153.47352.39081.75502.93312.3840
H184.61193.35342.19853.24482.21561.09365.44905.04724.66623.81452.77154.22552.39083.47352.93311.75502.3840

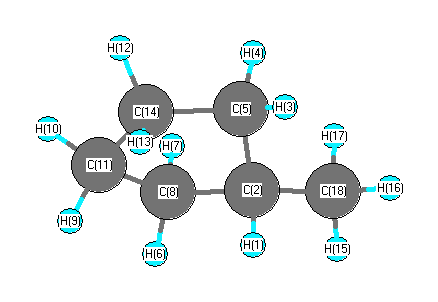
picture of Cyclopentane, methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
H1 C2 C5 150.047 H1 C2 C8 27.926
H1 C2 C18 138.751 C2 C5 H3 36.467
C2 C5 H4 37.556 C2 C5 C14 55.296
C2 C8 H6 23.041 C2 C8 H7 66.039
C2 C8 C11 50.026 C2 C18 H15 55.987
C2 C18 H16 62.247 C2 C18 H17 69.179
H3 C5 H4 68.623 H3 C5 C14 72.178
H4 C5 C14 28.146 C5 C2 C8 169.987
C5 C2 C18 41.321 C5 C14 C11 62.402
C5 C14 H12 67.269 C5 C14 H13 75.326
H6 C8 H7 88.588 H6 C8 C11 27.784
H7 C8 C11 116.062 C8 C2 C18 130.503
C8 C11 H9 31.792 C8 C11 H10 55.696
C8 C11 C14 66.705 H9 C11 H10 61.294
H9 C11 C14 40.857 H10 C11 C14 53.055
C11 C14 H12 92.651 C11 C14 H13 24.392
H12 C14 H13 117.043 H15 C18 H16 52.161
H15 C18 H17 36.743 H16 C18 H17 88.904
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C -0.458      
2 C -0.110      
3 C -0.286      
4 C -0.286      
5 C -0.297      
6 C -0.297      
7 H 0.149      
8 H 0.149      
9 H 0.145      
10 H 0.134      
11 H 0.145      
12 H 0.145      
13 H 0.139      
14 H 0.139      
15 H 0.147      
16 H 0.147      
17 H 0.148      
18 H 0.148      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 8.201 0.213 0.000
y 0.213 9.680 0.000
z 0.000 0.000 8.886


<r2> (average value of r2) Å2
<r2> 179.259
(<r2>)1/2 13.389