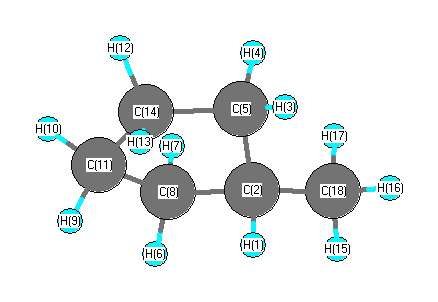
Vibrational Frequencies calculated at HSEh1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3144 |
2990 |
89.03 |
|
|
|
| 2 |
A' |
3133 |
2979 |
29.85 |
|
|
|
| 3 |
A' |
3121 |
2968 |
5.71 |
|
|
|
| 4 |
A' |
3091 |
2940 |
46.09 |
|
|
|
| 5 |
A' |
3062 |
2912 |
74.65 |
|
|
|
| 6 |
A' |
3057 |
2907 |
11.00 |
|
|
|
| 7 |
A' |
3029 |
2880 |
9.83 |
|
|
|
| 8 |
A' |
1549 |
1473 |
1.24 |
|
|
|
| 9 |
A' |
1528 |
1453 |
11.17 |
|
|
|
| 10 |
A' |
1522 |
1447 |
1.62 |
|
|
|
| 11 |
A' |
1438 |
1367 |
3.92 |
|
|
|
| 12 |
A' |
1412 |
1343 |
0.64 |
|
|
|
| 13 |
A' |
1349 |
1283 |
0.01 |
|
|
|
| 14 |
A' |
1336 |
1271 |
1.36 |
|
|
|
| 15 |
A' |
1266 |
1204 |
0.63 |
|
|
|
| 16 |
A' |
1234 |
1174 |
0.28 |
|
|
|
| 17 |
A' |
1128 |
1072 |
0.01 |
|
|
|
| 18 |
A' |
1075 |
1023 |
0.22 |
|
|
|
| 19 |
A' |
1037 |
986 |
1.08 |
|
|
|
| 20 |
A' |
945 |
898 |
0.02 |
|
|
|
| 21 |
A' |
914 |
869 |
1.10 |
|
|
|
| 22 |
A' |
871 |
828 |
1.36 |
|
|
|
| 23 |
A' |
768 |
730 |
0.96 |
|
|
|
| 24 |
A' |
535 |
509 |
0.60 |
|
|
|
| 25 |
A' |
435 |
413 |
0.22 |
|
|
|
| 26 |
A' |
183 |
174 |
0.01 |
|
|
|
| 27 |
A" |
3140 |
2986 |
30.31 |
|
|
|
| 28 |
A" |
3126 |
2973 |
10.42 |
|
|
|
| 29 |
A" |
3114 |
2961 |
43.92 |
|
|
|
| 30 |
A" |
3079 |
2928 |
19.30 |
|
|
|
| 31 |
A" |
3054 |
2904 |
22.13 |
|
|
|
| 32 |
A" |
1525 |
1450 |
2.78 |
|
|
|
| 33 |
A" |
1520 |
1446 |
2.10 |
|
|
|
| 34 |
A" |
1508 |
1434 |
0.02 |
|
|
|
| 35 |
A" |
1371 |
1304 |
1.68 |
|
|
|
| 36 |
A" |
1357 |
1290 |
0.01 |
|
|
|
| 37 |
A" |
1312 |
1248 |
0.07 |
|
|
|
| 38 |
A" |
1279 |
1216 |
0.24 |
|
|
|
| 39 |
A" |
1219 |
1159 |
0.43 |
|
|
|
| 40 |
A" |
1183 |
1125 |
0.87 |
|
|
|
| 41 |
A" |
1015 |
965 |
2.93 |
|
|
|
| 42 |
A" |
988 |
939 |
0.97 |
|
|
|
| 43 |
A" |
935 |
889 |
0.09 |
|
|
|
| 44 |
A" |
829 |
788 |
0.31 |
|
|
|
| 45 |
A" |
630 |
600 |
0.47 |
|
|
|
| 46 |
A" |
304 |
289 |
0.05 |
|
|
|
| 47 |
A" |
240 |
228 |
0.00 |
|
|
|
| 48 |
A" |
35 |
33 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 37460.5 cm
-1
Scaled (by 0.951) Zero Point Vibrational Energy (zpe) 35624.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.511 |
|
|
|
| 2 |
C |
-0.131 |
|
|
|
| 3 |
C |
-0.318 |
|
|
|
| 4 |
C |
-0.318 |
|
|
|
| 5 |
C |
-0.330 |
|
|
|
| 6 |
C |
-0.330 |
|
|
|
| 7 |
H |
0.167 |
|
|
|
| 8 |
H |
0.167 |
|
|
|
| 9 |
H |
0.161 |
|
|
|
| 10 |
H |
0.153 |
|
|
|
| 11 |
H |
0.161 |
|
|
|
| 12 |
H |
0.161 |
|
|
|
| 13 |
H |
0.156 |
|
|
|
| 14 |
H |
0.156 |
|
|
|
| 15 |
H |
0.164 |
|
|
|
| 16 |
H |
0.164 |
|
|
|
| 17 |
H |
0.164 |
|
|
|
| 18 |
H |
0.164 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.070 |
-0.030 |
0.000 |
0.077 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.749 |
0.006 |
0.000 |
| y |
0.006 |
-40.194 |
0.000 |
| z |
0.000 |
0.000 |
-39.735 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.216 |
0.006 |
0.000 |
| y |
0.006 |
-0.952 |
0.000 |
| z |
0.000 |
0.000 |
-0.263 |
|
| Polar |
|---|
| 3z2-r2 | -0.526 |
|---|
| x2-y2 | 1.445 |
|---|
| xy | 0.006 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.270 |
0.215 |
0.000 |
| y |
0.215 |
9.811 |
0.000 |
| z |
0.000 |
0.000 |
8.979 |
<r2> (average value of r
2) Å
2
| <r2> |
178.217 |
| (<r2>)1/2 |
13.350 |