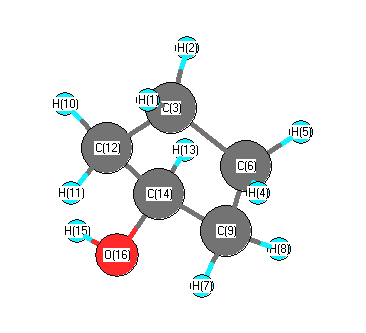
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3787 |
3623 |
14.25 |
|
|
|
| 2 |
A |
3136 |
3000 |
71.68 |
|
|
|
| 3 |
A |
3125 |
2989 |
16.98 |
|
|
|
| 4 |
A |
3111 |
2976 |
4.40 |
|
|
|
| 5 |
A |
3104 |
2969 |
41.58 |
|
|
|
| 6 |
A |
3082 |
2948 |
34.13 |
|
|
|
| 7 |
A |
3071 |
2938 |
31.02 |
|
|
|
| 8 |
A |
3066 |
2933 |
24.08 |
|
|
|
| 9 |
A |
3057 |
2924 |
31.46 |
|
|
|
| 10 |
A |
2977 |
2848 |
56.81 |
|
|
|
| 11 |
A |
1547 |
1480 |
2.58 |
|
|
|
| 12 |
A |
1522 |
1456 |
5.13 |
|
|
|
| 13 |
A |
1519 |
1453 |
2.47 |
|
|
|
| 14 |
A |
1507 |
1442 |
0.67 |
|
|
|
| 15 |
A |
1452 |
1389 |
3.56 |
|
|
|
| 16 |
A |
1395 |
1335 |
35.08 |
|
|
|
| 17 |
A |
1352 |
1294 |
0.11 |
|
|
|
| 18 |
A |
1344 |
1285 |
1.60 |
|
|
|
| 19 |
A |
1333 |
1275 |
9.02 |
|
|
|
| 20 |
A |
1312 |
1256 |
4.96 |
|
|
|
| 21 |
A |
1291 |
1235 |
19.67 |
|
|
|
| 22 |
A |
1273 |
1218 |
1.37 |
|
|
|
| 23 |
A |
1225 |
1172 |
25.54 |
|
|
|
| 24 |
A |
1211 |
1159 |
0.14 |
|
|
|
| 25 |
A |
1202 |
1150 |
2.69 |
|
|
|
| 26 |
A |
1131 |
1082 |
66.80 |
|
|
|
| 27 |
A |
1090 |
1043 |
32.73 |
|
|
|
| 28 |
A |
1067 |
1021 |
3.93 |
|
|
|
| 29 |
A |
995 |
952 |
1.30 |
|
|
|
| 30 |
A |
982 |
940 |
4.95 |
|
|
|
| 31 |
A |
969 |
927 |
6.02 |
|
|
|
| 32 |
A |
910 |
870 |
0.06 |
|
|
|
| 33 |
A |
892 |
854 |
0.13 |
|
|
|
| 34 |
A |
824 |
789 |
0.77 |
|
|
|
| 35 |
A |
771 |
737 |
2.43 |
|
|
|
| 36 |
A |
626 |
599 |
0.55 |
|
|
|
| 37 |
A |
541 |
517 |
5.57 |
|
|
|
| 38 |
A |
471 |
451 |
7.73 |
|
|
|
| 39 |
A |
359 |
343 |
18.23 |
|
|
|
| 40 |
A |
297 |
284 |
115.16 |
|
|
|
| 41 |
A |
181 |
173 |
1.02 |
|
|
|
| 42 |
A |
32 |
30 |
0.16 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 32067.0 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 30678.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.167 |
|
|
0.015 |
| 2 |
H |
0.164 |
|
|
0.014 |
| 3 |
C |
-0.331 |
|
|
0.045 |
| 4 |
H |
0.168 |
|
|
0.067 |
| 5 |
H |
0.165 |
|
|
0.065 |
| 6 |
C |
-0.331 |
|
|
-0.227 |
| 7 |
H |
0.176 |
|
|
0.062 |
| 8 |
H |
0.169 |
|
|
0.022 |
| 9 |
C |
-0.332 |
|
|
-0.024 |
| 10 |
H |
0.153 |
|
|
0.030 |
| 11 |
H |
0.170 |
|
|
0.071 |
| 12 |
C |
-0.347 |
|
|
-0.199 |
| 13 |
H |
0.134 |
|
|
-0.019 |
| 14 |
C |
0.109 |
|
|
0.347 |
| 15 |
H |
0.402 |
|
|
0.407 |
| 16 |
O |
-0.635 |
|
|
-0.676 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.534 |
1.196 |
0.808 |
1.538 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
-0.569 |
1.175 |
0.803 |
1.533 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.076 |
3.781 |
2.110 |
| y |
3.781 |
-37.009 |
0.329 |
| z |
2.110 |
0.329 |
-37.090 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.026 |
3.781 |
2.110 |
| y |
3.781 |
0.575 |
0.329 |
| z |
2.110 |
0.329 |
0.452 |
|
| Polar |
|---|
| 3z2-r2 | 0.904 |
|---|
| x2-y2 | -1.067 |
|---|
| xy | 3.781 |
|---|
| xz | 2.110 |
|---|
| yz | 0.329 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
165.753 |
| (<r2>)1/2 |
12.875 |