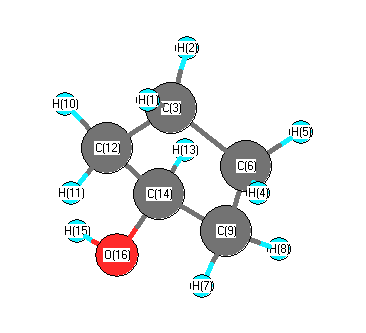
Vibrational Frequencies calculated at PBE1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3816 |
3626 |
16.91 |
|
|
|
| 2 |
A |
3154 |
2997 |
63.32 |
|
|
|
| 3 |
A |
3142 |
2986 |
14.56 |
|
|
|
| 4 |
A |
3128 |
2973 |
3.89 |
|
|
|
| 5 |
A |
3121 |
2966 |
39.39 |
|
|
|
| 6 |
A |
3098 |
2944 |
32.85 |
|
|
|
| 7 |
A |
3086 |
2933 |
27.28 |
|
|
|
| 8 |
A |
3081 |
2928 |
21.89 |
|
|
|
| 9 |
A |
3071 |
2919 |
29.56 |
|
|
|
| 10 |
A |
2992 |
2843 |
54.57 |
|
|
|
| 11 |
A |
1549 |
1472 |
3.07 |
|
|
|
| 12 |
A |
1523 |
1447 |
5.67 |
|
|
|
| 13 |
A |
1520 |
1444 |
3.03 |
|
|
|
| 14 |
A |
1508 |
1433 |
0.81 |
|
|
|
| 15 |
A |
1459 |
1387 |
3.53 |
|
|
|
| 16 |
A |
1402 |
1333 |
38.44 |
|
|
|
| 17 |
A |
1358 |
1290 |
0.09 |
|
|
|
| 18 |
A |
1348 |
1281 |
1.88 |
|
|
|
| 19 |
A |
1337 |
1271 |
10.46 |
|
|
|
| 20 |
A |
1316 |
1251 |
5.84 |
|
|
|
| 21 |
A |
1294 |
1230 |
19.96 |
|
|
|
| 22 |
A |
1279 |
1215 |
1.34 |
|
|
|
| 23 |
A |
1230 |
1169 |
27.13 |
|
|
|
| 24 |
A |
1217 |
1156 |
0.13 |
|
|
|
| 25 |
A |
1207 |
1147 |
1.89 |
|
|
|
| 26 |
A |
1147 |
1090 |
62.84 |
|
|
|
| 27 |
A |
1100 |
1046 |
32.24 |
|
|
|
| 28 |
A |
1078 |
1024 |
3.59 |
|
|
|
| 29 |
A |
999 |
950 |
1.10 |
|
|
|
| 30 |
A |
987 |
938 |
7.10 |
|
|
|
| 31 |
A |
973 |
925 |
4.10 |
|
|
|
| 32 |
A |
918 |
873 |
0.08 |
|
|
|
| 33 |
A |
902 |
857 |
0.11 |
|
|
|
| 34 |
A |
829 |
787 |
0.85 |
|
|
|
| 35 |
A |
773 |
735 |
2.29 |
|
|
|
| 36 |
A |
626 |
595 |
0.60 |
|
|
|
| 37 |
A |
544 |
517 |
5.71 |
|
|
|
| 38 |
A |
473 |
449 |
7.63 |
|
|
|
| 39 |
A |
362 |
344 |
19.37 |
|
|
|
| 40 |
A |
299 |
284 |
114.88 |
|
|
|
| 41 |
A |
183 |
174 |
0.98 |
|
|
|
| 42 |
A |
30 |
29 |
0.16 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 32229.4 cm
-1
Scaled (by 0.9503) Zero Point Vibrational Energy (zpe) 30627.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.173 |
|
|
|
| 2 |
H |
0.169 |
|
|
|
| 3 |
C |
-0.343 |
|
|
|
| 4 |
H |
0.174 |
|
|
|
| 5 |
H |
0.170 |
|
|
|
| 6 |
C |
-0.342 |
|
|
|
| 7 |
H |
0.182 |
|
|
|
| 8 |
H |
0.175 |
|
|
|
| 9 |
C |
-0.343 |
|
|
|
| 10 |
H |
0.159 |
|
|
|
| 11 |
H |
0.175 |
|
|
|
| 12 |
C |
-0.358 |
|
|
|
| 13 |
H |
0.139 |
|
|
|
| 14 |
C |
0.102 |
|
|
|
| 15 |
H |
0.405 |
|
|
|
| 16 |
O |
-0.638 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.526 |
1.207 |
0.791 |
1.536 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.145 |
3.799 |
2.053 |
| y |
3.799 |
-37.076 |
0.317 |
| z |
2.053 |
0.317 |
-37.179 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.018 |
3.799 |
2.053 |
| y |
3.799 |
0.586 |
0.317 |
| z |
2.053 |
0.317 |
0.432 |
|
| Polar |
|---|
| 3z2-r2 | 0.863 |
|---|
| x2-y2 | -1.069 |
|---|
| xy | 3.799 |
|---|
| xz | 2.053 |
|---|
| yz | 0.317 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.482 |
0.210 |
-0.019 |
| y |
0.210 |
8.011 |
0.088 |
| z |
-0.019 |
0.088 |
7.190 |
<r2> (average value of r
2) Å
2
| <r2> |
165.103 |
| (<r2>)1/2 |
12.849 |