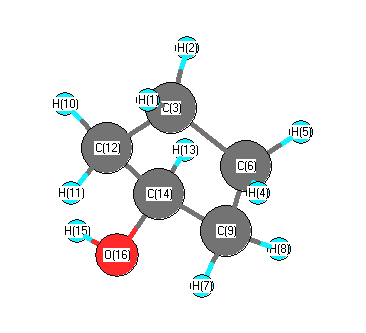
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3633 |
3573 |
5.99 |
|
|
|
| 2 |
A |
3060 |
3009 |
65.55 |
|
|
|
| 3 |
A |
3048 |
2998 |
15.67 |
|
|
|
| 4 |
A |
3034 |
2984 |
4.89 |
|
|
|
| 5 |
A |
3028 |
2978 |
42.24 |
|
|
|
| 6 |
A |
3006 |
2957 |
36.04 |
|
|
|
| 7 |
A |
2994 |
2945 |
24.81 |
|
|
|
| 8 |
A |
2989 |
2940 |
22.95 |
|
|
|
| 9 |
A |
2979 |
2930 |
30.62 |
|
|
|
| 10 |
A |
2886 |
2838 |
58.70 |
|
|
|
| 11 |
A |
1500 |
1475 |
2.27 |
|
|
|
| 12 |
A |
1476 |
1451 |
4.61 |
|
|
|
| 13 |
A |
1473 |
1448 |
1.94 |
|
|
|
| 14 |
A |
1463 |
1439 |
0.78 |
|
|
|
| 15 |
A |
1403 |
1380 |
3.52 |
|
|
|
| 16 |
A |
1348 |
1326 |
33.06 |
|
|
|
| 17 |
A |
1308 |
1287 |
0.24 |
|
|
|
| 18 |
A |
1296 |
1275 |
2.34 |
|
|
|
| 19 |
A |
1289 |
1268 |
7.87 |
|
|
|
| 20 |
A |
1268 |
1247 |
5.88 |
|
|
|
| 21 |
A |
1248 |
1228 |
15.54 |
|
|
|
| 22 |
A |
1235 |
1214 |
1.71 |
|
|
|
| 23 |
A |
1181 |
1162 |
17.41 |
|
|
|
| 24 |
A |
1172 |
1153 |
3.32 |
|
|
|
| 25 |
A |
1163 |
1143 |
1.35 |
|
|
|
| 26 |
A |
1086 |
1068 |
56.72 |
|
|
|
| 27 |
A |
1056 |
1039 |
34.69 |
|
|
|
| 28 |
A |
1035 |
1018 |
8.36 |
|
|
|
| 29 |
A |
968 |
952 |
0.95 |
|
|
|
| 30 |
A |
952 |
936 |
7.68 |
|
|
|
| 31 |
A |
940 |
925 |
4.15 |
|
|
|
| 32 |
A |
884 |
870 |
0.04 |
|
|
|
| 33 |
A |
869 |
854 |
0.07 |
|
|
|
| 34 |
A |
799 |
786 |
1.00 |
|
|
|
| 35 |
A |
752 |
740 |
2.14 |
|
|
|
| 36 |
A |
605 |
595 |
0.67 |
|
|
|
| 37 |
A |
524 |
516 |
5.15 |
|
|
|
| 38 |
A |
458 |
450 |
7.03 |
|
|
|
| 39 |
A |
350 |
344 |
21.30 |
|
|
|
| 40 |
A |
293 |
288 |
101.09 |
|
|
|
| 41 |
A |
177 |
174 |
0.92 |
|
|
|
| 42 |
A |
30 |
30 |
0.15 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31128.2 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 30614.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.153 |
|
|
|
| 2 |
H |
0.151 |
|
|
|
| 3 |
C |
-0.304 |
|
|
|
| 4 |
H |
0.155 |
|
|
|
| 5 |
H |
0.152 |
|
|
|
| 6 |
C |
-0.304 |
|
|
|
| 7 |
H |
0.163 |
|
|
|
| 8 |
H |
0.155 |
|
|
|
| 9 |
C |
-0.305 |
|
|
|
| 10 |
H |
0.139 |
|
|
|
| 11 |
H |
0.156 |
|
|
|
| 12 |
C |
-0.320 |
|
|
|
| 13 |
H |
0.120 |
|
|
|
| 14 |
C |
0.098 |
|
|
|
| 15 |
H |
0.383 |
|
|
|
| 16 |
O |
-0.591 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.483 |
1.160 |
0.745 |
1.461 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.160 |
3.699 |
1.941 |
| y |
3.699 |
-37.230 |
0.308 |
| z |
1.941 |
0.308 |
-37.393 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.848 |
3.699 |
1.941 |
| y |
3.699 |
0.546 |
0.308 |
| z |
1.941 |
0.308 |
0.302 |
|
| Polar |
|---|
| 3z2-r2 | 0.604 |
|---|
| x2-y2 | -0.929 |
|---|
| xy | 3.699 |
|---|
| xz | 1.941 |
|---|
| yz | 0.308 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.869 |
0.231 |
-0.009 |
| y |
0.231 |
8.311 |
0.096 |
| z |
-0.009 |
0.096 |
7.396 |
<r2> (average value of r
2) Å
2
| <r2> |
167.381 |
| (<r2>)1/2 |
12.938 |