Jump to
S1C2
Energy calculated at B3PW91/6-31G*
| | hartrees |
|---|
| Energy at 0K | -342.266854 |
| Energy at 298.15K | |
| HF Energy | -342.266854 |
| Nuclear repulsion energy | 245.630125 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3078 |
2945 |
55.49 |
|
|
|
| 2 |
A1 |
1967 |
1882 |
583.61 |
|
|
|
| 3 |
A1 |
1573 |
1505 |
3.43 |
|
|
|
| 4 |
A1 |
1412 |
1350 |
1.58 |
|
|
|
| 5 |
A1 |
1227 |
1174 |
233.72 |
|
|
|
| 6 |
A1 |
961 |
920 |
8.28 |
|
|
|
| 7 |
A1 |
848 |
811 |
30.59 |
|
|
|
| 8 |
A1 |
727 |
695 |
0.88 |
|
|
|
| 9 |
A2 |
3106 |
2972 |
0.00 |
|
|
|
| 10 |
A2 |
1262 |
1208 |
0.00 |
|
|
|
| 11 |
A2 |
1179 |
1128 |
0.00 |
|
|
|
| 12 |
A2 |
169i |
162i |
0.00 |
|
|
|
| 13 |
B1 |
3128 |
2992 |
50.35 |
|
|
|
| 14 |
B1 |
1279 |
1223 |
0.84 |
|
|
|
| 15 |
B1 |
869 |
831 |
0.64 |
|
|
|
| 16 |
B1 |
755 |
722 |
22.75 |
|
|
|
| 17 |
B1 |
146 |
140 |
1.73 |
|
|
|
| 18 |
B2 |
3071 |
2938 |
29.62 |
|
|
|
| 19 |
B2 |
1560 |
1492 |
2.59 |
|
|
|
| 20 |
B2 |
1424 |
1363 |
17.37 |
|
|
|
| 21 |
B2 |
1214 |
1162 |
6.54 |
|
|
|
| 22 |
B2 |
1064 |
1018 |
315.88 |
|
|
|
| 23 |
B2 |
756 |
723 |
1.17 |
|
|
|
| 24 |
B2 |
506 |
485 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16471.9 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 15758.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/6-31G*
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.864 |
| O2 |
0.000 |
0.000 |
2.051 |
| O3 |
0.000 |
1.129 |
0.057 |
| O4 |
0.000 |
-1.129 |
0.057 |
| C5 |
0.000 |
0.775 |
-1.285 |
| C6 |
0.000 |
-0.775 |
-1.285 |
| H7 |
-0.889 |
1.191 |
-1.772 |
| H8 |
0.889 |
1.191 |
-1.772 |
| H9 |
0.889 |
-1.191 |
-1.772 |
| H10 |
-0.889 |
-1.191 |
-1.772 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
O3 |
O4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.1873 | 1.3879 | 1.3879 | 2.2845 | 2.2845 | 3.0259 | 3.0259 | 3.0259 | 3.0259 |
O2 | 1.1873 | | 2.2918 | 2.2918 | 3.4251 | 3.4251 | 4.1018 | 4.1018 | 4.1018 | 4.1018 | O3 | 1.3879 | 2.2918 | | 2.2582 | 1.3879 | 2.3295 | 2.0343 | 2.0343 | 3.0850 | 3.0850 | O4 | 1.3879 | 2.2918 | 2.2582 | | 2.3295 | 1.3879 | 3.0850 | 3.0850 | 2.0343 | 2.0343 | C5 | 2.2845 | 3.4251 | 1.3879 | 2.3295 | | 1.5501 | 1.0955 | 1.0955 | 2.2119 | 2.2119 | C6 | 2.2845 | 3.4251 | 2.3295 | 1.3879 | 1.5501 | | 2.2119 | 2.2119 | 1.0955 | 1.0955 | H7 | 3.0259 | 4.1018 | 2.0343 | 3.0850 | 1.0955 | 2.2119 | | 1.7777 | 2.9722 | 2.3820 | H8 | 3.0259 | 4.1018 | 2.0343 | 3.0850 | 1.0955 | 2.2119 | 1.7777 | | 2.3820 | 2.9722 | H9 | 3.0259 | 4.1018 | 3.0850 | 2.0343 | 2.2119 | 1.0955 | 2.9722 | 2.3820 | | 1.7777 | H10 | 3.0259 | 4.1018 | 3.0850 | 2.0343 | 2.2119 | 1.0955 | 2.3820 | 2.9722 | 1.7777 | |
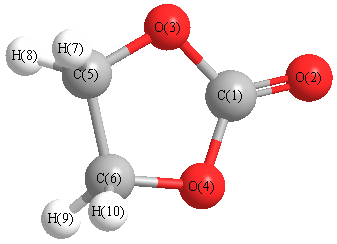 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O3 |
C5 |
110.777 |
|
C1 |
O4 |
C6 |
110.777 |
| O2 |
C1 |
O3 |
125.556 |
|
O2 |
C1 |
O4 |
125.556 |
| O3 |
C1 |
O4 |
108.888 |
|
O3 |
C5 |
C6 |
104.779 |
| O3 |
C5 |
H7 |
109.441 |
|
O3 |
C5 |
H8 |
109.441 |
| O4 |
C6 |
C5 |
104.779 |
|
O4 |
C6 |
H9 |
109.441 |
| O4 |
C6 |
H10 |
109.441 |
|
C5 |
C6 |
H9 |
112.313 |
| C5 |
C6 |
H10 |
112.313 |
|
C6 |
C5 |
H7 |
112.313 |
| C6 |
C5 |
H8 |
112.313 |
|
H7 |
C5 |
H8 |
108.466 |
| H9 |
C6 |
H10 |
108.466 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.811 |
|
|
|
| 2 |
O |
-0.442 |
|
|
|
| 3 |
O |
-0.455 |
|
|
|
| 4 |
O |
-0.455 |
|
|
|
| 5 |
C |
-0.112 |
|
|
|
| 6 |
C |
-0.112 |
|
|
|
| 7 |
H |
0.191 |
|
|
|
| 8 |
H |
0.191 |
|
|
|
| 9 |
H |
0.191 |
|
|
|
| 10 |
H |
0.191 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-4.847 |
4.847 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-31.744 |
0.000 |
0.000 |
| y |
0.000 |
-35.761 |
0.000 |
| z |
0.000 |
0.000 |
-35.194 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.733 |
0.000 |
0.000 |
| y |
0.000 |
-2.291 |
0.000 |
| z |
0.000 |
0.000 |
-1.442 |
|
| Polar |
|---|
| 3z2-r2 | -2.884 |
|---|
| x2-y2 | 4.017 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.158 |
0.000 |
0.000 |
| y |
0.000 |
5.027 |
0.000 |
| z |
0.000 |
0.000 |
7.228 |
<r2> (average value of r
2) Å
2
| <r2> |
128.387 |
| (<r2>)1/2 |
11.331 |
Jump to
S1C1
Energy calculated at B3PW91/6-31G*
| | hartrees |
|---|
| Energy at 0K | -342.267442 |
| Energy at 298.15K | -342.273794 |
| HF Energy | -342.267442 |
| Nuclear repulsion energy | 246.134536 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3133 |
2998 |
17.65 |
|
|
|
| 2 |
A |
3057 |
2925 |
30.42 |
|
|
|
| 3 |
A |
1970 |
1885 |
568.97 |
|
|
|
| 4 |
A |
1561 |
1494 |
3.66 |
|
|
|
| 5 |
A |
1416 |
1355 |
6.68 |
|
|
|
| 6 |
A |
1268 |
1213 |
21.51 |
|
|
|
| 7 |
A |
1224 |
1171 |
162.71 |
|
|
|
| 8 |
A |
1165 |
1115 |
40.85 |
|
|
|
| 9 |
A |
964 |
922 |
3.70 |
|
|
|
| 10 |
A |
844 |
807 |
33.92 |
|
|
|
| 11 |
A |
724 |
693 |
1.06 |
|
|
|
| 12 |
A |
120 |
115 |
0.50 |
|
|
|
| 13 |
B |
3144 |
3007 |
32.19 |
|
|
|
| 14 |
B |
3062 |
2929 |
50.46 |
|
|
|
| 15 |
B |
1552 |
1485 |
3.56 |
|
|
|
| 16 |
B |
1419 |
1357 |
15.42 |
|
|
|
| 17 |
B |
1281 |
1225 |
3.09 |
|
|
|
| 18 |
B |
1192 |
1140 |
5.50 |
|
|
|
| 19 |
B |
1059 |
1013 |
296.44 |
|
|
|
| 20 |
B |
906 |
867 |
6.78 |
|
|
|
| 21 |
B |
764 |
731 |
23.83 |
|
|
|
| 22 |
B |
696 |
666 |
0.86 |
|
|
|
| 23 |
B |
507 |
485 |
0.04 |
|
|
|
| 24 |
B |
153 |
147 |
1.58 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16590.3 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 15871.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/6-31G*
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.860 |
| O2 |
0.000 |
0.000 |
2.047 |
| O3 |
0.000 |
1.129 |
0.048 |
| O4 |
0.000 |
-1.129 |
0.048 |
| C5 |
0.185 |
0.746 |
-1.276 |
| C6 |
-0.185 |
-0.746 |
-1.276 |
| H7 |
-0.459 |
1.347 |
-1.923 |
| H8 |
1.232 |
0.909 |
-1.566 |
| H9 |
0.459 |
-1.347 |
-1.923 |
| H10 |
-1.232 |
-0.909 |
-1.566 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
O3 |
O4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.1870 | 1.3909 | 1.3909 | 2.2700 | 2.2700 | 3.1258 | 2.8681 | 3.1258 | 2.8681 |
O2 | 1.1870 | | 2.2959 | 2.2959 | 3.4106 | 3.4106 | 4.2174 | 3.9233 | 4.2174 | 3.9233 | O3 | 1.3909 | 2.2959 | | 2.2587 | 1.3909 | 2.3030 | 2.0355 | 2.0419 | 3.1982 | 2.8767 | O4 | 1.3909 | 2.2959 | 2.2587 | | 2.3030 | 1.3909 | 3.1982 | 2.8767 | 2.0355 | 2.0419 | C5 | 2.2700 | 3.4106 | 1.3909 | 2.3030 | | 1.5370 | 1.0931 | 1.0981 | 2.2077 | 2.1979 | C6 | 2.2700 | 3.4106 | 2.3030 | 1.3909 | 1.5370 | | 2.2077 | 2.1979 | 1.0931 | 1.0981 | H7 | 3.1258 | 4.2174 | 2.0355 | 3.1982 | 1.0931 | 2.2077 | | 1.7826 | 2.8460 | 2.4116 | H8 | 2.8681 | 3.9233 | 2.0419 | 2.8767 | 1.0981 | 2.1979 | 1.7826 | | 2.4116 | 3.0619 | H9 | 3.1258 | 4.2174 | 3.1982 | 2.0355 | 2.2077 | 1.0931 | 2.8460 | 2.4116 | | 1.7826 | H10 | 2.8681 | 3.9233 | 2.8767 | 2.0419 | 2.1979 | 1.0981 | 2.4116 | 3.0619 | 1.7826 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O3 |
C5 |
109.375 |
|
C1 |
O4 |
C6 |
109.375 |
| O2 |
C1 |
O3 |
125.712 |
|
O2 |
C1 |
O4 |
125.712 |
| O3 |
C1 |
O4 |
108.576 |
|
O3 |
C5 |
C6 |
103.620 |
| O3 |
C5 |
H7 |
109.476 |
|
O3 |
C5 |
H8 |
109.687 |
| O4 |
C6 |
C5 |
103.620 |
|
O4 |
C6 |
H9 |
109.476 |
| O4 |
C6 |
H10 |
109.687 |
|
C5 |
C6 |
H9 |
113.066 |
| C5 |
C6 |
H10 |
111.965 |
|
C6 |
C5 |
H7 |
113.066 |
| C6 |
C5 |
H8 |
111.965 |
|
H7 |
C5 |
H8 |
108.886 |
| H9 |
C6 |
H10 |
108.886 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.816 |
|
|
|
| 2 |
O |
-0.443 |
|
|
|
| 3 |
O |
-0.455 |
|
|
|
| 4 |
O |
-0.455 |
|
|
|
| 5 |
C |
-0.114 |
|
|
|
| 6 |
C |
-0.114 |
|
|
|
| 7 |
H |
0.195 |
|
|
|
| 8 |
H |
0.188 |
|
|
|
| 9 |
H |
0.195 |
|
|
|
| 10 |
H |
0.188 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-4.794 |
4.794 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-31.793 |
0.205 |
0.000 |
| y |
0.205 |
-35.781 |
0.000 |
| z |
0.000 |
0.000 |
-35.036 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.615 |
0.205 |
0.000 |
| y |
0.205 |
-2.366 |
0.000 |
| z |
0.000 |
0.000 |
-1.249 |
|
| Polar |
|---|
| 3z2-r2 | -2.498 |
|---|
| x2-y2 | 3.987 |
|---|
| xy | 0.205 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.229 |
0.130 |
0.000 |
| y |
0.130 |
4.979 |
0.000 |
| z |
0.000 |
0.000 |
7.201 |
<r2> (average value of r
2) Å
2
| <r2> |
127.423 |
| (<r2>)1/2 |
11.288 |
