Jump to
S1C2
Energy calculated at HSEh1PBE/6-31G*
| | hartrees |
|---|
| Energy at 0K | -342.057187 |
| Energy at 298.15K | |
| HF Energy | -342.057187 |
| Nuclear repulsion energy | 246.165045 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3093 |
2941 |
53.88 |
|
|
|
| 2 |
A1 |
1984 |
1887 |
598.08 |
|
|
|
| 3 |
A1 |
1577 |
1500 |
3.45 |
|
|
|
| 4 |
A1 |
1417 |
1348 |
1.85 |
|
|
|
| 5 |
A1 |
1245 |
1184 |
238.82 |
|
|
|
| 6 |
A1 |
971 |
923 |
8.69 |
|
|
|
| 7 |
A1 |
860 |
817 |
30.75 |
|
|
|
| 8 |
A1 |
733 |
697 |
0.85 |
|
|
|
| 9 |
A2 |
3122 |
2969 |
0.00 |
|
|
|
| 10 |
A2 |
1267 |
1205 |
0.00 |
|
|
|
| 11 |
A2 |
1183 |
1125 |
0.00 |
|
|
|
| 12 |
A2 |
174i |
165i |
0.00 |
|
|
|
| 13 |
B1 |
3143 |
2989 |
48.23 |
|
|
|
| 14 |
B1 |
1284 |
1221 |
0.89 |
|
|
|
| 15 |
B1 |
872 |
829 |
0.78 |
|
|
|
| 16 |
B1 |
763 |
725 |
23.57 |
|
|
|
| 17 |
B1 |
146 |
139 |
1.68 |
|
|
|
| 18 |
B2 |
3086 |
2935 |
28.50 |
|
|
|
| 19 |
B2 |
1564 |
1487 |
3.06 |
|
|
|
| 20 |
B2 |
1430 |
1360 |
18.89 |
|
|
|
| 21 |
B2 |
1230 |
1170 |
8.66 |
|
|
|
| 22 |
B2 |
1079 |
1026 |
316.32 |
|
|
|
| 23 |
B2 |
763 |
726 |
0.82 |
|
|
|
| 24 |
B2 |
511 |
486 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16574.1 cm
-1
Scaled (by 0.951) Zero Point Vibrational Energy (zpe) 15762.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G*
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.862 |
| O2 |
0.000 |
0.000 |
2.047 |
| O3 |
0.000 |
1.126 |
0.057 |
| O4 |
0.000 |
-1.126 |
0.057 |
| C5 |
0.000 |
0.774 |
-1.282 |
| C6 |
0.000 |
-0.774 |
-1.282 |
| H7 |
-0.888 |
1.189 |
-1.769 |
| H8 |
0.888 |
1.189 |
-1.769 |
| H9 |
0.888 |
-1.189 |
-1.769 |
| H10 |
-0.888 |
-1.189 |
-1.769 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
O3 |
O4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.1853 | 1.3842 | 1.3842 | 2.2790 | 2.2790 | 3.0208 | 3.0208 | 3.0208 | 3.0208 |
O2 | 1.1853 | | 2.2868 | 2.2868 | 3.4177 | 3.4177 | 4.0948 | 4.0948 | 4.0948 | 4.0948 | O3 | 1.3842 | 2.2868 | | 2.2520 | 1.3842 | 2.3239 | 2.0316 | 2.0316 | 3.0793 | 3.0793 | O4 | 1.3842 | 2.2868 | 2.2520 | | 2.3239 | 1.3842 | 3.0793 | 3.0793 | 2.0316 | 2.0316 | C5 | 2.2790 | 3.4177 | 1.3842 | 2.3239 | | 1.5474 | 1.0950 | 1.0950 | 2.2087 | 2.2087 | C6 | 2.2790 | 3.4177 | 2.3239 | 1.3842 | 1.5474 | | 2.2087 | 2.2087 | 1.0950 | 1.0950 | H7 | 3.0208 | 4.0948 | 2.0316 | 3.0793 | 1.0950 | 2.2087 | | 1.7767 | 2.9682 | 2.3778 | H8 | 3.0208 | 4.0948 | 2.0316 | 3.0793 | 1.0950 | 2.2087 | 1.7767 | | 2.3778 | 2.9682 | H9 | 3.0208 | 4.0948 | 3.0793 | 2.0316 | 2.2087 | 1.0950 | 2.9682 | 2.3778 | | 1.7767 | H10 | 3.0208 | 4.0948 | 3.0793 | 2.0316 | 2.2087 | 1.0950 | 2.3778 | 2.9682 | 1.7767 | |
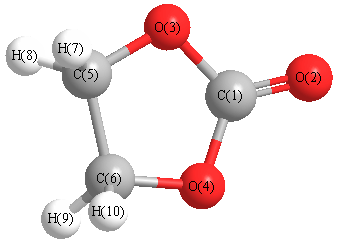 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O3 |
C5 |
110.821 |
|
C1 |
O4 |
C6 |
110.821 |
| O2 |
C1 |
O3 |
125.565 |
|
O2 |
C1 |
O4 |
125.565 |
| O3 |
C1 |
O4 |
108.870 |
|
O3 |
C5 |
C6 |
104.744 |
| O3 |
C5 |
H7 |
109.508 |
|
O3 |
C5 |
H8 |
109.508 |
| O4 |
C6 |
C5 |
104.744 |
|
O4 |
C6 |
H9 |
109.508 |
| O4 |
C6 |
H10 |
109.508 |
|
C5 |
C6 |
H9 |
112.282 |
| C5 |
C6 |
H10 |
112.282 |
|
C6 |
C5 |
H7 |
112.282 |
| C6 |
C5 |
H8 |
112.282 |
|
H7 |
C5 |
H8 |
108.436 |
| H9 |
C6 |
H10 |
108.436 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.812 |
|
|
|
| 2 |
O |
-0.440 |
|
|
|
| 3 |
O |
-0.456 |
|
|
|
| 4 |
O |
-0.456 |
|
|
|
| 5 |
C |
-0.115 |
|
|
|
| 6 |
C |
-0.115 |
|
|
|
| 7 |
H |
0.193 |
|
|
|
| 8 |
H |
0.193 |
|
|
|
| 9 |
H |
0.193 |
|
|
|
| 10 |
H |
0.193 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-4.838 |
4.838 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-31.799 |
0.000 |
0.000 |
| y |
0.000 |
-35.827 |
0.000 |
| z |
0.000 |
0.000 |
-35.278 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.754 |
0.000 |
0.000 |
| y |
0.000 |
-2.289 |
0.000 |
| z |
0.000 |
0.000 |
-1.465 |
|
| Polar |
|---|
| 3z2-r2 | -2.930 |
|---|
| x2-y2 | 4.029 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.147 |
0.000 |
0.000 |
| y |
0.000 |
4.990 |
0.000 |
| z |
0.000 |
0.000 |
7.168 |
<r2> (average value of r
2) Å
2
| <r2> |
127.974 |
| (<r2>)1/2 |
11.313 |
Jump to
S1C1
Energy calculated at HSEh1PBE/6-31G*
| | hartrees |
|---|
| Energy at 0K | -342.057855 |
| Energy at 298.15K | -342.064240 |
| HF Energy | -342.057855 |
| Nuclear repulsion energy | 246.698953 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3149 |
2994 |
16.78 |
|
|
|
| 2 |
A |
3071 |
2920 |
29.29 |
|
|
|
| 3 |
A |
1988 |
1890 |
582.33 |
|
|
|
| 4 |
A |
1564 |
1488 |
3.71 |
|
|
|
| 5 |
A |
1422 |
1352 |
7.75 |
|
|
|
| 6 |
A |
1273 |
1211 |
26.96 |
|
|
|
| 7 |
A |
1240 |
1179 |
170.53 |
|
|
|
| 8 |
A |
1171 |
1113 |
31.56 |
|
|
|
| 9 |
A |
972 |
925 |
3.61 |
|
|
|
| 10 |
A |
855 |
813 |
34.30 |
|
|
|
| 11 |
A |
731 |
695 |
1.04 |
|
|
|
| 12 |
A |
127 |
120 |
0.53 |
|
|
|
| 13 |
B |
3159 |
3004 |
30.28 |
|
|
|
| 14 |
B |
3076 |
2925 |
49.00 |
|
|
|
| 15 |
B |
1555 |
1479 |
4.16 |
|
|
|
| 16 |
B |
1424 |
1354 |
16.82 |
|
|
|
| 17 |
B |
1287 |
1224 |
3.48 |
|
|
|
| 18 |
B |
1205 |
1146 |
6.73 |
|
|
|
| 19 |
B |
1072 |
1020 |
297.52 |
|
|
|
| 20 |
B |
913 |
868 |
5.66 |
|
|
|
| 21 |
B |
773 |
735 |
24.79 |
|
|
|
| 22 |
B |
698 |
664 |
0.77 |
|
|
|
| 23 |
B |
511 |
486 |
0.02 |
|
|
|
| 24 |
B |
153 |
145 |
1.52 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16693.0 cm
-1
Scaled (by 0.951) Zero Point Vibrational Energy (zpe) 15875.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G*
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.857 |
| O2 |
0.000 |
0.000 |
2.042 |
| O3 |
0.000 |
1.126 |
0.047 |
| O4 |
0.000 |
-1.126 |
0.047 |
| C5 |
0.190 |
0.743 |
-1.273 |
| C6 |
-0.190 |
-0.743 |
-1.273 |
| H7 |
-0.446 |
1.348 |
-1.924 |
| H8 |
1.239 |
0.898 |
-1.557 |
| H9 |
0.446 |
-1.348 |
-1.924 |
| H10 |
-1.239 |
-0.898 |
-1.557 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
O3 |
O4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.1849 | 1.3874 | 1.3874 | 2.2637 | 2.2637 | 3.1224 | 2.8578 | 3.1224 | 2.8578 |
O2 | 1.1849 | | 2.2909 | 2.2909 | 3.4023 | 3.4023 | 4.2124 | 3.9104 | 4.2124 | 3.9104 | O3 | 1.3874 | 2.2909 | | 2.2527 | 1.3874 | 2.2961 | 2.0330 | 2.0392 | 3.1944 | 2.8643 | O4 | 1.3874 | 2.2909 | 2.2527 | | 2.2961 | 1.3874 | 3.1944 | 2.8643 | 2.0330 | 2.0392 | C5 | 2.2637 | 3.4023 | 1.3874 | 2.2961 | | 1.5336 | 1.0926 | 1.0977 | 2.2046 | 2.1939 | C6 | 2.2637 | 3.4023 | 2.2961 | 1.3874 | 1.5336 | | 2.2046 | 2.1939 | 1.0926 | 1.0977 | H7 | 3.1224 | 4.2124 | 2.0330 | 3.1944 | 1.0926 | 2.2046 | | 1.7819 | 2.8390 | 2.4095 | H8 | 2.8578 | 3.9104 | 2.0392 | 2.8643 | 1.0977 | 2.1939 | 1.7819 | | 2.4095 | 3.0597 | H9 | 3.1224 | 4.2124 | 3.1944 | 2.0330 | 2.2046 | 1.0926 | 2.8390 | 2.4095 | | 1.7819 | H10 | 2.8578 | 3.9104 | 2.8643 | 2.0392 | 2.1939 | 1.0977 | 2.4095 | 3.0597 | 1.7819 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O3 |
C5 |
109.336 |
|
C1 |
O4 |
C6 |
109.336 |
| O2 |
C1 |
O3 |
125.724 |
|
O2 |
C1 |
O4 |
125.724 |
| O3 |
C1 |
O4 |
108.553 |
|
O3 |
C5 |
C6 |
103.529 |
| O3 |
C5 |
H7 |
109.548 |
|
O3 |
C5 |
H8 |
109.737 |
| O4 |
C6 |
C5 |
103.529 |
|
O4 |
C6 |
H9 |
109.548 |
| O4 |
C6 |
H10 |
109.737 |
|
C5 |
C6 |
H9 |
113.095 |
| C5 |
C6 |
H10 |
111.914 |
|
C6 |
C5 |
H7 |
113.095 |
| C6 |
C5 |
H8 |
111.914 |
|
H7 |
C5 |
H8 |
108.880 |
| H9 |
C6 |
H10 |
108.880 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.817 |
|
|
|
| 2 |
O |
-0.441 |
|
|
|
| 3 |
O |
-0.457 |
|
|
|
| 4 |
O |
-0.457 |
|
|
|
| 5 |
C |
-0.117 |
|
|
|
| 6 |
C |
-0.117 |
|
|
|
| 7 |
H |
0.197 |
|
|
|
| 8 |
H |
0.189 |
|
|
|
| 9 |
H |
0.197 |
|
|
|
| 10 |
H |
0.189 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-4.783 |
4.783 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-31.854 |
0.206 |
0.000 |
| y |
0.206 |
-35.844 |
0.000 |
| z |
0.000 |
0.000 |
-35.109 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.623 |
0.206 |
0.000 |
| y |
0.206 |
-2.363 |
0.000 |
| z |
0.000 |
0.000 |
-1.260 |
|
| Polar |
|---|
| 3z2-r2 | -2.521 |
|---|
| x2-y2 | 3.990 |
|---|
| xy | 0.206 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.220 |
0.131 |
0.000 |
| y |
0.131 |
4.941 |
0.000 |
| z |
0.000 |
0.000 |
7.140 |
<r2> (average value of r
2) Å
2
| <r2> |
126.959 |
| (<r2>)1/2 |
11.268 |
