Jump to
S1C2
Energy calculated at PBEPBEultrafine/6-31G*
| | hartrees |
|---|
| Energy at 0K | -342.023020 |
| Energy at 298.15K | |
| HF Energy | -342.023020 |
| Nuclear repulsion energy | 243.333848 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
2994 |
2944 |
57.33 |
|
|
|
| 2 |
A1 |
1898 |
1866 |
501.40 |
|
|
|
| 3 |
A1 |
1523 |
1498 |
3.15 |
|
|
|
| 4 |
A1 |
1359 |
1336 |
1.53 |
|
|
|
| 5 |
A1 |
1157 |
1138 |
202.71 |
|
|
|
| 6 |
A1 |
932 |
917 |
7.22 |
|
|
|
| 7 |
A1 |
805 |
792 |
29.22 |
|
|
|
| 8 |
A1 |
700 |
689 |
0.74 |
|
|
|
| 9 |
A2 |
3018 |
2969 |
0.00 |
|
|
|
| 10 |
A2 |
1216 |
1196 |
0.00 |
|
|
|
| 11 |
A2 |
1139 |
1120 |
0.00 |
|
|
|
| 12 |
A2 |
167i |
164i |
0.00 |
|
|
|
| 13 |
B1 |
3041 |
2990 |
50.72 |
|
|
|
| 14 |
B1 |
1232 |
1212 |
0.44 |
|
|
|
| 15 |
B1 |
842 |
829 |
0.32 |
|
|
|
| 16 |
B1 |
711 |
699 |
15.22 |
|
|
|
| 17 |
B1 |
149 |
146 |
1.57 |
|
|
|
| 18 |
B2 |
2987 |
2938 |
30.43 |
|
|
|
| 19 |
B2 |
1511 |
1486 |
1.75 |
|
|
|
| 20 |
B2 |
1371 |
1348 |
14.00 |
|
|
|
| 21 |
B2 |
1148 |
1129 |
2.99 |
|
|
|
| 22 |
B2 |
1007 |
990 |
290.83 |
|
|
|
| 23 |
B2 |
725 |
713 |
2.99 |
|
|
|
| 24 |
B2 |
485 |
477 |
0.14 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 15891.1 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 15628.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBEultrafine/6-31G*
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.871 |
| O2 |
0.000 |
0.000 |
2.070 |
| O3 |
0.000 |
1.143 |
0.058 |
| O4 |
0.000 |
-1.143 |
0.058 |
| C5 |
0.000 |
0.779 |
-1.297 |
| C6 |
0.000 |
-0.779 |
-1.297 |
| H7 |
-0.896 |
1.198 |
-1.788 |
| H8 |
0.896 |
1.198 |
-1.788 |
| H9 |
0.896 |
-1.198 |
-1.788 |
| H10 |
-0.896 |
-1.198 |
-1.788 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
O3 |
O4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.1989 | 1.4026 | 1.4026 | 2.3032 | 2.3032 | 3.0505 | 3.0505 | 3.0505 | 3.0505 |
O2 | 1.1989 | | 2.3138 | 2.3138 | 3.4553 | 3.4553 | 4.1373 | 4.1373 | 4.1373 | 4.1373 | O3 | 1.4026 | 2.3138 | | 2.2860 | 1.4026 | 2.3514 | 2.0523 | 2.0523 | 3.1127 | 3.1127 | O4 | 1.4026 | 2.3138 | 2.2860 | | 2.3514 | 1.4026 | 3.1127 | 3.1127 | 2.0523 | 2.0523 | C5 | 2.3032 | 3.4553 | 1.4026 | 2.3514 | | 1.5580 | 1.1042 | 1.1042 | 2.2253 | 2.2253 | C6 | 2.3032 | 3.4553 | 2.3514 | 1.4026 | 1.5580 | | 2.2253 | 2.2253 | 1.1042 | 1.1042 | H7 | 3.0505 | 4.1373 | 2.0523 | 3.1127 | 1.1042 | 2.2253 | | 1.7919 | 2.9918 | 2.3959 | H8 | 3.0505 | 4.1373 | 2.0523 | 3.1127 | 1.1042 | 2.2253 | 1.7919 | | 2.3959 | 2.9918 | H9 | 3.0505 | 4.1373 | 3.1127 | 2.0523 | 2.2253 | 1.1042 | 2.9918 | 2.3959 | | 1.7919 | H10 | 3.0505 | 4.1373 | 3.1127 | 2.0523 | 2.2253 | 1.1042 | 2.3959 | 2.9918 | 1.7919 | |
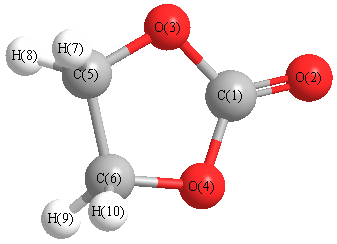 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O3 |
C5 |
110.382 |
|
C1 |
O4 |
C6 |
110.382 |
| O2 |
C1 |
O3 |
125.422 |
|
O2 |
C1 |
O4 |
125.422 |
| O3 |
C1 |
O4 |
109.155 |
|
O3 |
C5 |
C6 |
105.040 |
| O3 |
C5 |
H7 |
109.329 |
|
O3 |
C5 |
H8 |
109.329 |
| O4 |
C6 |
C5 |
105.040 |
|
O4 |
C6 |
H9 |
109.329 |
| O4 |
C6 |
H10 |
109.329 |
|
C5 |
C6 |
H9 |
112.296 |
| C5 |
C6 |
H10 |
112.296 |
|
C6 |
C5 |
H7 |
112.296 |
| C6 |
C5 |
H8 |
112.296 |
|
H7 |
C5 |
H8 |
108.461 |
| H9 |
C6 |
H10 |
108.461 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.721 |
|
|
|
| 2 |
O |
-0.406 |
|
|
|
| 3 |
O |
-0.406 |
|
|
|
| 4 |
O |
-0.406 |
|
|
|
| 5 |
C |
-0.109 |
|
|
|
| 6 |
C |
-0.109 |
|
|
|
| 7 |
H |
0.179 |
|
|
|
| 8 |
H |
0.179 |
|
|
|
| 9 |
H |
0.179 |
|
|
|
| 10 |
H |
0.179 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-4.638 |
4.638 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-31.979 |
0.000 |
0.000 |
| y |
0.000 |
-35.698 |
0.000 |
| z |
0.000 |
0.000 |
-35.161 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.451 |
0.000 |
0.000 |
| y |
0.000 |
-2.128 |
0.000 |
| z |
0.000 |
0.000 |
-1.322 |
|
| Polar |
|---|
| 3z2-r2 | -2.644 |
|---|
| x2-y2 | 3.719 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.261 |
0.000 |
0.000 |
| y |
0.000 |
5.266 |
0.000 |
| z |
0.000 |
0.000 |
7.632 |
<r2> (average value of r
2) Å
2
| <r2> |
130.378 |
| (<r2>)1/2 |
11.418 |
Jump to
S1C1
Energy calculated at PBEPBEultrafine/6-31G*
| | hartrees |
|---|
| Energy at 0K | -342.023774 |
| Energy at 298.15K | -342.030029 |
| HF Energy | -342.023774 |
| Nuclear repulsion energy | 243.895609 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3050 |
3000 |
19.01 |
|
|
|
| 2 |
A |
2970 |
2921 |
29.95 |
|
|
|
| 3 |
A |
1900 |
1868 |
486.89 |
|
|
|
| 4 |
A |
1510 |
1485 |
3.28 |
|
|
|
| 5 |
A |
1364 |
1341 |
6.69 |
|
|
|
| 6 |
A |
1222 |
1202 |
14.66 |
|
|
|
| 7 |
A |
1163 |
1144 |
101.39 |
|
|
|
| 8 |
A |
1117 |
1099 |
78.48 |
|
|
|
| 9 |
A |
936 |
920 |
3.41 |
|
|
|
| 10 |
A |
800 |
787 |
32.28 |
|
|
|
| 11 |
A |
698 |
686 |
0.92 |
|
|
|
| 12 |
A |
138 |
136 |
0.50 |
|
|
|
| 13 |
B |
3060 |
3009 |
30.25 |
|
|
|
| 14 |
B |
2976 |
2927 |
54.12 |
|
|
|
| 15 |
B |
1502 |
1477 |
2.83 |
|
|
|
| 16 |
B |
1364 |
1342 |
12.15 |
|
|
|
| 17 |
B |
1233 |
1213 |
2.29 |
|
|
|
| 18 |
B |
1127 |
1108 |
3.56 |
|
|
|
| 19 |
B |
1001 |
985 |
261.06 |
|
|
|
| 20 |
B |
876 |
862 |
14.34 |
|
|
|
| 21 |
B |
723 |
711 |
16.39 |
|
|
|
| 22 |
B |
665 |
654 |
1.29 |
|
|
|
| 23 |
B |
485 |
477 |
0.26 |
|
|
|
| 24 |
B |
157 |
155 |
1.42 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16017.3 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 15753.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBEultrafine/6-31G*
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.866 |
| O2 |
0.000 |
0.000 |
2.064 |
| O3 |
0.000 |
1.143 |
0.047 |
| O4 |
0.000 |
-1.143 |
0.047 |
| C5 |
0.198 |
0.746 |
-1.287 |
| C6 |
-0.198 |
-0.746 |
-1.287 |
| H7 |
-0.433 |
1.360 |
-1.948 |
| H8 |
1.259 |
0.892 |
-1.565 |
| H9 |
0.433 |
-1.360 |
-1.948 |
| H10 |
-1.259 |
-0.892 |
-1.565 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
O3 |
O4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.1986 | 1.4061 | 1.4061 | 2.2866 | 2.2866 | 3.1553 | 2.8791 | 3.1553 | 2.8791 |
O2 | 1.1986 | | 2.3186 | 2.3186 | 3.4388 | 3.4388 | 4.2589 | 3.9436 | 4.2589 | 3.9436 | O3 | 1.4061 | 2.3186 | | 2.2869 | 1.4061 | 2.3211 | 2.0535 | 2.0610 | 3.2308 | 2.8859 | O4 | 1.4061 | 2.3186 | 2.2869 | | 2.3211 | 1.4061 | 3.2308 | 2.8859 | 2.0535 | 2.0610 | C5 | 2.2866 | 3.4388 | 1.4061 | 2.3211 | | 1.5428 | 1.1016 | 1.1071 | 2.2198 | 2.2096 | C6 | 2.2866 | 3.4388 | 2.3211 | 1.4061 | 1.5428 | | 2.2198 | 2.2096 | 1.1016 | 1.1071 | H7 | 3.1553 | 4.2589 | 2.0535 | 3.2308 | 1.1016 | 2.2198 | | 1.7976 | 2.8552 | 2.4294 | H8 | 2.8791 | 3.9436 | 2.0610 | 2.8859 | 1.1071 | 2.2096 | 1.7976 | | 2.4294 | 3.0865 | H9 | 3.1553 | 4.2589 | 3.2308 | 2.0535 | 2.2198 | 1.1016 | 2.8552 | 2.4294 | | 1.7976 | H10 | 2.8791 | 3.9436 | 2.8859 | 2.0610 | 2.2096 | 1.1071 | 2.4294 | 3.0865 | 1.7976 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O3 |
C5 |
108.795 |
|
C1 |
O4 |
C6 |
108.795 |
| O2 |
C1 |
O3 |
125.593 |
|
O2 |
C1 |
O4 |
125.593 |
| O3 |
C1 |
O4 |
108.814 |
|
O3 |
C5 |
C6 |
103.733 |
| O3 |
C5 |
H7 |
109.345 |
|
O3 |
C5 |
H8 |
109.612 |
| O4 |
C6 |
C5 |
103.733 |
|
O4 |
C6 |
H9 |
109.345 |
| O4 |
C6 |
H10 |
109.612 |
|
C5 |
C6 |
H9 |
113.102 |
| C5 |
C6 |
H10 |
111.941 |
|
C6 |
C5 |
H7 |
113.102 |
| C6 |
C5 |
H8 |
111.941 |
|
H7 |
C5 |
H8 |
108.958 |
| H9 |
C6 |
H10 |
108.958 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.727 |
|
|
|
| 2 |
O |
-0.407 |
|
|
|
| 3 |
O |
-0.406 |
|
|
|
| 4 |
O |
-0.406 |
|
|
|
| 5 |
C |
-0.110 |
|
|
|
| 6 |
C |
-0.110 |
|
|
|
| 7 |
H |
0.182 |
|
|
|
| 8 |
H |
0.176 |
|
|
|
| 9 |
H |
0.182 |
|
|
|
| 10 |
H |
0.176 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-4.579 |
4.579 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-32.039 |
0.196 |
0.000 |
| y |
0.196 |
-35.721 |
0.000 |
| z |
0.000 |
0.000 |
-34.991 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.317 |
0.196 |
0.000 |
| y |
0.196 |
-2.206 |
0.000 |
| z |
0.000 |
0.000 |
-1.111 |
|
| Polar |
|---|
| 3z2-r2 | -2.222 |
|---|
| x2-y2 | 3.682 |
|---|
| xy | 0.196 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.350 |
0.152 |
0.000 |
| y |
0.152 |
5.209 |
0.000 |
| z |
0.000 |
0.000 |
7.596 |
<r2> (average value of r
2) Å
2
| <r2> |
129.275 |
| (<r2>)1/2 |
11.370 |
