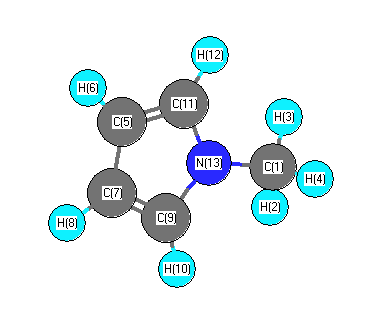
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3211 |
3158 |
1.34 |
|
|
|
| 2 |
A |
3203 |
3150 |
8.57 |
|
|
|
| 3 |
A |
3191 |
3139 |
5.71 |
|
|
|
| 4 |
A |
3182 |
3129 |
1.70 |
|
|
|
| 5 |
A |
3085 |
3034 |
11.17 |
|
|
|
| 6 |
A |
3051 |
3001 |
20.38 |
|
|
|
| 7 |
A |
2979 |
2930 |
64.43 |
|
|
|
| 8 |
A |
1534 |
1508 |
0.82 |
|
|
|
| 9 |
A |
1517 |
1492 |
30.43 |
|
|
|
| 10 |
A |
1493 |
1469 |
6.33 |
|
|
|
| 11 |
A |
1469 |
1445 |
12.23 |
|
|
|
| 12 |
A |
1424 |
1401 |
9.20 |
|
|
|
| 13 |
A |
1414 |
1390 |
9.15 |
|
|
|
| 14 |
A |
1389 |
1366 |
0.30 |
|
|
|
| 15 |
A |
1303 |
1281 |
35.01 |
|
|
|
| 16 |
A |
1258 |
1237 |
0.26 |
|
|
|
| 17 |
A |
1117 |
1099 |
6.79 |
|
|
|
| 18 |
A |
1089 |
1071 |
21.68 |
|
|
|
| 19 |
A |
1083 |
1065 |
10.50 |
|
|
|
| 20 |
A |
1070 |
1053 |
1.03 |
|
|
|
| 21 |
A |
1042 |
1025 |
6.35 |
|
|
|
| 22 |
A |
963 |
947 |
7.23 |
|
|
|
| 23 |
A |
856 |
842 |
0.01 |
|
|
|
| 24 |
A |
818 |
805 |
0.11 |
|
|
|
| 25 |
A |
755 |
742 |
0.11 |
|
|
|
| 26 |
A |
697 |
685 |
88.62 |
|
|
|
| 27 |
A |
664 |
653 |
5.08 |
|
|
|
| 28 |
A |
624 |
614 |
0.02 |
|
|
|
| 29 |
A |
606 |
596 |
0.00 |
|
|
|
| 30 |
A |
599 |
590 |
3.79 |
|
|
|
| 31 |
A |
350 |
345 |
0.53 |
|
|
|
| 32 |
A |
184 |
181 |
3.22 |
|
|
|
| 33 |
A |
90 |
88 |
0.06 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 23654.9 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 23264.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.369 |
|
|
|
| 2 |
H |
0.181 |
|
|
|
| 3 |
H |
0.181 |
|
|
|
| 4 |
H |
0.184 |
|
|
|
| 5 |
C |
-0.198 |
|
|
|
| 6 |
H |
0.127 |
|
|
|
| 7 |
C |
-0.198 |
|
|
|
| 8 |
H |
0.127 |
|
|
|
| 9 |
C |
0.011 |
|
|
|
| 10 |
H |
0.140 |
|
|
|
| 11 |
C |
0.011 |
|
|
|
| 12 |
H |
0.140 |
|
|
|
| 13 |
N |
-0.338 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.094 |
0.000 |
0.006 |
2.094 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-31.224 |
0.000 |
-0.081 |
| y |
0.000 |
-32.784 |
0.000 |
| z |
-0.081 |
0.000 |
-39.049 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.693 |
0.000 |
-0.081 |
| y |
0.000 |
2.353 |
0.000 |
| z |
-0.081 |
0.000 |
-7.045 |
|
| Polar |
|---|
| 3z2-r2 | -14.091 |
|---|
| x2-y2 | 1.560 |
|---|
| xy | 0.000 |
|---|
| xz | -0.081 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.747 |
-0.000 |
0.020 |
| y |
-0.000 |
9.220 |
0.000 |
| z |
0.020 |
0.000 |
4.196 |
<r2> (average value of r
2) Å
2
| <r2> |
144.214 |
| (<r2>)1/2 |
12.009 |