Jump to
S1C2
Energy calculated at CID/6-31G*
| | hartrees |
|---|
| Energy at 0K | -342.159508 |
| Energy at 298.15K | -342.164512 |
| Nuclear repulsion energy | 273.439210 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3431 |
3169 |
0.37 |
|
|
|
| 2 |
A' |
3404 |
3145 |
0.17 |
|
|
|
| 3 |
A' |
3390 |
3131 |
1.85 |
|
|
|
| 4 |
A' |
3121 |
2883 |
89.32 |
|
|
|
| 5 |
A' |
1935 |
1787 |
310.42 |
|
|
|
| 6 |
A' |
1740 |
1607 |
4.55 |
|
|
|
| 7 |
A' |
1625 |
1501 |
52.47 |
|
|
|
| 8 |
A' |
1533 |
1416 |
37.90 |
|
|
|
| 9 |
A' |
1490 |
1377 |
3.49 |
|
|
|
| 10 |
A' |
1398 |
1292 |
37.24 |
|
|
|
| 11 |
A' |
1326 |
1224 |
10.67 |
|
|
|
| 12 |
A' |
1276 |
1179 |
20.21 |
|
|
|
| 13 |
A' |
1184 |
1094 |
6.11 |
|
|
|
| 14 |
A' |
1100 |
1016 |
36.52 |
|
|
|
| 15 |
A' |
1013 |
935 |
24.32 |
|
|
|
| 16 |
A' |
944 |
872 |
10.93 |
|
|
|
| 17 |
A' |
804 |
742 |
74.48 |
|
|
|
| 18 |
A' |
526 |
486 |
1.53 |
|
|
|
| 19 |
A' |
213 |
196 |
6.69 |
|
|
|
| 20 |
A" |
1072 |
991 |
0.10 |
|
|
|
| 21 |
A" |
928 |
858 |
0.84 |
|
|
|
| 22 |
A" |
899 |
830 |
7.20 |
|
|
|
| 23 |
A" |
831 |
768 |
63.32 |
|
|
|
| 24 |
A" |
675 |
624 |
5.90 |
|
|
|
| 25 |
A" |
629 |
581 |
7.17 |
|
|
|
| 26 |
A" |
290 |
268 |
14.60 |
|
|
|
| 27 |
A" |
135 |
125 |
2.78 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 18454.8 cm
-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 17046.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CID/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
0.487 |
-0.895 |
0.000 |
| C2 |
-0.570 |
-1.720 |
0.000 |
| C3 |
-1.731 |
-1.028 |
0.000 |
| C4 |
0.000 |
0.366 |
0.000 |
| C5 |
-1.355 |
0.345 |
0.000 |
| C6 |
0.935 |
1.486 |
0.000 |
| O7 |
2.132 |
1.376 |
0.000 |
| H8 |
-0.346 |
-2.770 |
0.000 |
| H9 |
-2.723 |
-1.438 |
0.000 |
| H10 |
-2.006 |
1.201 |
0.000 |
| H11 |
0.447 |
2.470 |
0.000 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
C5 |
C6 |
O7 |
H8 |
H9 |
H10 |
H11 |
| O1 | | 1.3414 | 2.2222 | 1.3519 | 2.2213 | 2.4226 | 2.8049 | 2.0512 | 3.2561 | 3.2572 | 3.3651 |
C2 | 1.3414 | | 1.3514 | 2.1627 | 2.2097 | 3.5418 | 4.1104 | 1.0728 | 2.1714 | 3.2550 | 4.3117 | C3 | 2.2222 | 1.3514 | | 2.2225 | 1.4238 | 3.6643 | 4.5507 | 2.2248 | 1.0735 | 2.2460 | 4.1205 | C4 | 1.3519 | 2.1627 | 2.2225 | | 1.3555 | 1.4589 | 2.3599 | 3.1543 | 3.2664 | 2.1727 | 2.1508 | C5 | 2.2213 | 2.2097 | 1.4238 | 1.3555 | | 2.5587 | 3.6371 | 3.2741 | 2.2473 | 1.0749 | 2.7860 | C6 | 2.4226 | 3.5418 | 3.6643 | 1.4589 | 2.5587 | | 1.2023 | 4.4438 | 4.6830 | 2.9546 | 1.0984 | O7 | 2.8049 | 4.1104 | 4.5507 | 2.3599 | 3.6371 | 1.2023 | | 4.8305 | 5.6124 | 4.1419 | 2.0089 | H8 | 2.0512 | 1.0728 | 2.2248 | 3.1543 | 3.2741 | 4.4438 | 4.8305 | | 2.7246 | 4.3033 | 5.2989 | H9 | 3.2561 | 2.1714 | 1.0735 | 3.2664 | 2.2473 | 4.6830 | 5.6124 | 2.7246 | | 2.7347 | 5.0318 | H10 | 3.2572 | 3.2550 | 2.2460 | 2.1727 | 1.0749 | 2.9546 | 4.1419 | 4.3033 | 2.7347 | | 2.7614 | H11 | 3.3651 | 4.3117 | 4.1205 | 2.1508 | 2.7860 | 1.0984 | 2.0089 | 5.2989 | 5.0318 | 2.7614 | |
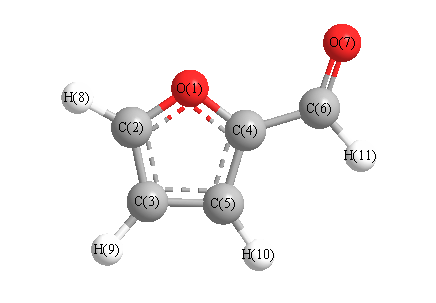 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C3 |
111.227 |
|
O1 |
C2 |
H8 |
115.908 |
| O1 |
C4 |
C5 |
110.256 |
|
O1 |
C4 |
C6 |
119.011 |
| C2 |
O1 |
C4 |
106.836 |
|
C2 |
C3 |
C5 |
105.515 |
| C2 |
C3 |
H9 |
126.752 |
|
C3 |
C2 |
H8 |
132.865 |
| C3 |
C5 |
C4 |
106.167 |
|
C3 |
C5 |
H10 |
127.472 |
| C4 |
C5 |
H10 |
126.361 |
|
C4 |
C6 |
O7 |
124.658 |
| C4 |
C6 |
H11 |
113.755 |
|
C5 |
C3 |
H9 |
127.733 |
| C5 |
C4 |
C6 |
130.733 |
|
O7 |
C6 |
H11 |
121.587 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CID/6-31G*
| | hartrees |
|---|
| Energy at 0K | -342.161547 |
| Energy at 298.15K | -342.166528 |
| Nuclear repulsion energy | 272.537325 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3431 |
3169 |
0.46 |
|
|
|
| 2 |
A' |
3412 |
3152 |
0.40 |
|
|
|
| 3 |
A' |
3398 |
3138 |
1.11 |
|
|
|
| 4 |
A' |
3142 |
2902 |
82.04 |
|
|
|
| 5 |
A' |
1927 |
1780 |
261.80 |
|
|
|
| 6 |
A' |
1750 |
1616 |
44.58 |
|
|
|
| 7 |
A' |
1619 |
1495 |
103.12 |
|
|
|
| 8 |
A' |
1541 |
1423 |
8.16 |
|
|
|
| 9 |
A' |
1486 |
1373 |
15.00 |
|
|
|
| 10 |
A' |
1346 |
1243 |
6.04 |
|
|
|
| 11 |
A' |
1338 |
1236 |
31.15 |
|
|
|
| 12 |
A' |
1257 |
1161 |
21.41 |
|
|
|
| 13 |
A' |
1187 |
1096 |
19.27 |
|
|
|
| 14 |
A' |
1093 |
1010 |
32.46 |
|
|
|
| 15 |
A' |
1029 |
950 |
13.55 |
|
|
|
| 16 |
A' |
943 |
871 |
7.76 |
|
|
|
| 17 |
A' |
794 |
734 |
88.62 |
|
|
|
| 18 |
A' |
520 |
480 |
1.45 |
|
|
|
| 19 |
A' |
214 |
198 |
7.70 |
|
|
|
| 20 |
A" |
1081 |
999 |
0.29 |
|
|
|
| 21 |
A" |
932 |
861 |
2.22 |
|
|
|
| 22 |
A" |
908 |
839 |
7.02 |
|
|
|
| 23 |
A" |
831 |
767 |
60.14 |
|
|
|
| 24 |
A" |
667 |
616 |
6.71 |
|
|
|
| 25 |
A" |
627 |
580 |
8.70 |
|
|
|
| 26 |
A" |
251 |
232 |
14.90 |
|
|
|
| 27 |
A" |
157 |
145 |
6.12 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 18438.9 cm
-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 17032.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CID/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
1.241 |
-0.268 |
0.000 |
| C2 |
1.075 |
-1.600 |
0.000 |
| C3 |
-0.237 |
-1.925 |
0.000 |
| C4 |
0.000 |
0.281 |
0.000 |
| C5 |
-0.943 |
-0.690 |
0.000 |
| C6 |
-0.082 |
1.738 |
0.000 |
| O7 |
-1.127 |
2.337 |
0.000 |
| H8 |
1.974 |
-2.187 |
0.000 |
| H9 |
-0.650 |
-2.916 |
0.000 |
| H10 |
-2.005 |
-0.531 |
0.000 |
| H11 |
0.884 |
2.258 |
0.000 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
C5 |
C6 |
O7 |
H8 |
H9 |
H10 |
H11 |
| O1 | | 1.3429 | 2.2206 | 1.3572 | 2.2250 | 2.4032 | 3.5202 | 2.0538 | 3.2542 | 3.2574 | 2.5506 |
C2 | 1.3429 | | 1.3517 | 2.1673 | 2.2149 | 3.5337 | 4.5114 | 1.0727 | 2.1697 | 3.2613 | 3.8629 | C3 | 2.2206 | 1.3517 | | 2.2190 | 1.4236 | 3.6668 | 4.3541 | 2.2259 | 1.0737 | 2.2522 | 4.3305 | C4 | 1.3572 | 2.1673 | 2.2190 | | 1.3536 | 1.4595 | 2.3443 | 3.1600 | 3.2628 | 2.1637 | 2.1654 | C5 | 2.2250 | 2.2149 | 1.4236 | 1.3536 | | 2.5762 | 3.0320 | 3.2790 | 2.2460 | 1.0738 | 3.4680 | C6 | 2.4032 | 3.5337 | 3.6668 | 1.4595 | 2.5762 | | 1.2042 | 4.4307 | 4.6891 | 2.9750 | 1.0971 | O7 | 3.5202 | 4.5114 | 4.3541 | 2.3443 | 3.0320 | 1.2042 | | 5.4841 | 5.2748 | 2.9996 | 2.0128 | H8 | 2.0538 | 1.0727 | 2.2259 | 3.1600 | 3.2790 | 4.4307 | 5.4841 | | 2.7232 | 4.3099 | 4.5758 | H9 | 3.2542 | 2.1697 | 1.0737 | 3.2628 | 2.2460 | 4.6891 | 5.2748 | 2.7232 | | 2.7436 | 5.3966 | H10 | 3.2574 | 3.2613 | 2.2522 | 2.1637 | 1.0738 | 2.9750 | 2.9996 | 4.3099 | 2.7436 | | 4.0161 | H11 | 2.5506 | 3.8629 | 4.3305 | 2.1654 | 3.4680 | 1.0971 | 2.0128 | 4.5758 | 5.3966 | 4.0161 | |
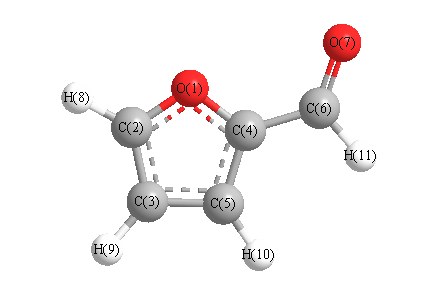 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C3 |
110.993 |
|
O1 |
C2 |
H8 |
116.029 |
| O1 |
C4 |
C5 |
110.325 |
|
O1 |
C4 |
C6 |
117.077 |
| C2 |
O1 |
C4 |
106.768 |
|
C2 |
C3 |
C5 |
105.869 |
| C2 |
C3 |
H9 |
126.530 |
|
C3 |
C2 |
H8 |
132.978 |
| C3 |
C5 |
C4 |
106.045 |
|
C3 |
C5 |
H10 |
128.256 |
| C4 |
C5 |
H10 |
125.699 |
|
C4 |
C6 |
O7 |
123.024 |
| C4 |
C6 |
H11 |
115.037 |
|
C5 |
C3 |
H9 |
127.600 |
| C5 |
C4 |
C6 |
132.598 |
|
O7 |
C6 |
H11 |
121.939 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
