Jump to
S1C2
Energy calculated at B3PW91/6-31G*
| | hartrees |
|---|
| Energy at 0K | -343.214086 |
| Energy at 298.15K | -343.218916 |
| Nuclear repulsion energy | 271.708141 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3311 |
3168 |
0.41 |
|
|
|
| 2 |
A' |
3290 |
3147 |
0.27 |
|
|
|
| 3 |
A' |
3278 |
3136 |
1.26 |
|
|
|
| 4 |
A' |
2932 |
2805 |
116.86 |
|
|
|
| 5 |
A' |
1808 |
1730 |
308.81 |
|
|
|
| 6 |
A' |
1628 |
1558 |
7.24 |
|
|
|
| 7 |
A' |
1530 |
1463 |
49.89 |
|
|
|
| 8 |
A' |
1455 |
1392 |
38.88 |
|
|
|
| 9 |
A' |
1412 |
1351 |
0.02 |
|
|
|
| 10 |
A' |
1322 |
1265 |
47.27 |
|
|
|
| 11 |
A' |
1263 |
1208 |
2.60 |
|
|
|
| 12 |
A' |
1218 |
1165 |
6.84 |
|
|
|
| 13 |
A' |
1127 |
1078 |
10.23 |
|
|
|
| 14 |
A' |
1055 |
1009 |
40.51 |
|
|
|
| 15 |
A' |
959 |
917 |
18.59 |
|
|
|
| 16 |
A' |
898 |
860 |
10.89 |
|
|
|
| 17 |
A' |
771 |
737 |
63.83 |
|
|
|
| 18 |
A' |
503 |
481 |
1.03 |
|
|
|
| 19 |
A' |
199 |
191 |
5.36 |
|
|
|
| 20 |
A" |
1009 |
966 |
0.00 |
|
|
|
| 21 |
A" |
894 |
855 |
1.14 |
|
|
|
| 22 |
A" |
839 |
803 |
6.30 |
|
|
|
| 23 |
A" |
779 |
746 |
57.42 |
|
|
|
| 24 |
A" |
657 |
629 |
1.55 |
|
|
|
| 25 |
A" |
609 |
583 |
7.09 |
|
|
|
| 26 |
A" |
292 |
280 |
11.19 |
|
|
|
| 27 |
A" |
137 |
131 |
1.37 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17587.4 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 16825.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
0.507 |
-0.896 |
0.000 |
| C2 |
-0.541 |
-1.740 |
0.000 |
| C3 |
-1.726 |
-1.057 |
0.000 |
| C4 |
0.000 |
0.369 |
0.000 |
| C5 |
-1.373 |
0.318 |
0.000 |
| C6 |
0.911 |
1.502 |
0.000 |
| O7 |
2.124 |
1.430 |
0.000 |
| H8 |
-0.297 |
-2.792 |
0.000 |
| H9 |
-2.718 |
-1.486 |
0.000 |
| H10 |
-2.041 |
1.170 |
0.000 |
| H11 |
0.382 |
2.481 |
0.000 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
C5 |
C6 |
O7 |
H8 |
H9 |
H10 |
H11 |
| O1 | | 1.3458 | 2.2389 | 1.3636 | 2.2388 | 2.4322 | 2.8326 | 2.0591 | 3.2787 | 3.2808 | 3.3793 |
C2 | 1.3458 | | 1.3670 | 2.1772 | 2.2197 | 3.5521 | 4.1410 | 1.0805 | 2.1912 | 3.2730 | 4.3200 | C3 | 2.2389 | 1.3670 | | 2.2385 | 1.4196 | 3.6740 | 4.5826 | 2.2479 | 1.0811 | 2.2486 | 4.1177 | C4 | 1.3636 | 2.1772 | 2.2385 | | 1.3740 | 1.4536 | 2.3738 | 3.1751 | 3.2906 | 2.1924 | 2.1456 | C5 | 2.2388 | 2.2197 | 1.4196 | 1.3740 | | 2.5723 | 3.6691 | 3.2914 | 2.2504 | 1.0820 | 2.7849 | C6 | 2.4322 | 3.5521 | 3.6740 | 1.4536 | 2.5723 | | 1.2151 | 4.4606 | 4.7006 | 2.9705 | 1.1120 | O7 | 2.8326 | 4.1410 | 4.5826 | 2.3738 | 3.6691 | 1.2151 | | 4.8664 | 5.6518 | 4.1729 | 2.0339 | H8 | 2.0591 | 1.0805 | 2.2479 | 3.1751 | 3.2914 | 4.4606 | 4.8664 | | 2.7509 | 4.3287 | 5.3161 | H9 | 3.2787 | 2.1912 | 1.0811 | 3.2906 | 2.2504 | 4.7006 | 5.6518 | 2.7509 | | 2.7405 | 5.0343 | H10 | 3.2808 | 3.2730 | 2.2486 | 2.1924 | 1.0820 | 2.9705 | 4.1729 | 4.3287 | 2.7405 | | 2.7552 | H11 | 3.3793 | 4.3200 | 4.1177 | 2.1456 | 2.7849 | 1.1120 | 2.0339 | 5.3161 | 5.0343 | 2.7552 | |
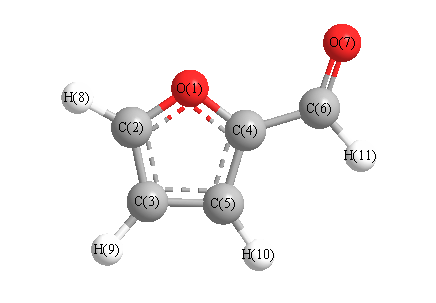 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C3 |
111.234 |
|
O1 |
C2 |
H8 |
115.708 |
| O1 |
C4 |
C5 |
109.730 |
|
O1 |
C4 |
C6 |
119.354 |
| C2 |
O1 |
C4 |
106.947 |
|
C2 |
C3 |
C5 |
105.588 |
| C2 |
C3 |
H9 |
126.635 |
|
C3 |
C2 |
H8 |
133.058 |
| C3 |
C5 |
C4 |
106.501 |
|
C3 |
C5 |
H10 |
127.494 |
| C4 |
C5 |
H10 |
126.005 |
|
C4 |
C6 |
O7 |
125.385 |
| C4 |
C6 |
H11 |
112.830 |
|
C5 |
C3 |
H9 |
127.777 |
| C5 |
C4 |
C6 |
130.916 |
|
O7 |
C6 |
H11 |
121.785 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.411 |
|
|
|
| 2 |
C |
0.090 |
|
|
|
| 3 |
C |
-0.227 |
|
|
|
| 4 |
C |
0.275 |
|
|
|
| 5 |
C |
-0.210 |
|
|
|
| 6 |
C |
0.175 |
|
|
|
| 7 |
O |
-0.410 |
|
|
|
| 8 |
H |
0.202 |
|
|
|
| 9 |
H |
0.189 |
|
|
|
| 10 |
H |
0.194 |
|
|
|
| 11 |
H |
0.133 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-3.994 |
-1.196 |
0.000 |
4.170 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-43.000 |
-2.815 |
0.000 |
| y |
-2.815 |
-33.081 |
0.000 |
| z |
0.000 |
0.000 |
-40.353 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.283 |
-2.815 |
0.000 |
| y |
-2.815 |
8.595 |
0.000 |
| z |
0.000 |
0.000 |
-2.312 |
|
| Polar |
|---|
| 3z2-r2 | -4.624 |
|---|
| x2-y2 | -9.919 |
|---|
| xy | -2.815 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.508 |
1.955 |
0.000 |
| y |
1.955 |
10.372 |
0.000 |
| z |
0.000 |
0.000 |
3.331 |
<r2> (average value of r
2) Å
2
| <r2> |
190.212 |
| (<r2>)1/2 |
13.792 |
Jump to
S1C1
Energy calculated at B3PW91/6-31G*
| | hartrees |
|---|
| Energy at 0K | -343.215649 |
| Energy at 298.15K | -343.220462 |
| Nuclear repulsion energy | 270.900633 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3311 |
3168 |
0.60 |
|
|
|
| 2 |
A' |
3300 |
3157 |
0.16 |
|
|
|
| 3 |
A' |
3283 |
3141 |
1.01 |
|
|
|
| 4 |
A' |
2954 |
2826 |
110.03 |
|
|
|
| 5 |
A' |
1802 |
1724 |
255.92 |
|
|
|
| 6 |
A' |
1635 |
1564 |
53.19 |
|
|
|
| 7 |
A' |
1520 |
1454 |
117.97 |
|
|
|
| 8 |
A' |
1465 |
1402 |
2.24 |
|
|
|
| 9 |
A' |
1419 |
1357 |
13.32 |
|
|
|
| 10 |
A' |
1283 |
1227 |
28.98 |
|
|
|
| 11 |
A' |
1259 |
1204 |
0.97 |
|
|
|
| 12 |
A' |
1203 |
1151 |
12.95 |
|
|
|
| 13 |
A' |
1132 |
1083 |
22.40 |
|
|
|
| 14 |
A' |
1048 |
1003 |
37.81 |
|
|
|
| 15 |
A' |
974 |
932 |
6.76 |
|
|
|
| 16 |
A' |
898 |
859 |
7.57 |
|
|
|
| 17 |
A' |
759 |
726 |
71.57 |
|
|
|
| 18 |
A' |
499 |
477 |
1.00 |
|
|
|
| 19 |
A' |
202 |
193 |
6.56 |
|
|
|
| 20 |
A" |
1017 |
973 |
0.08 |
|
|
|
| 21 |
A" |
903 |
864 |
1.54 |
|
|
|
| 22 |
A" |
845 |
808 |
8.46 |
|
|
|
| 23 |
A" |
777 |
743 |
52.65 |
|
|
|
| 24 |
A" |
647 |
619 |
1.53 |
|
|
|
| 25 |
A" |
610 |
583 |
10.00 |
|
|
|
| 26 |
A" |
253 |
242 |
12.26 |
|
|
|
| 27 |
A" |
161 |
154 |
2.60 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17578.3 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 16817.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
1.251 |
-0.266 |
0.000 |
| C2 |
1.087 |
-1.602 |
0.000 |
| C3 |
-0.240 |
-1.934 |
0.000 |
| C4 |
0.000 |
0.289 |
0.000 |
| C5 |
-0.947 |
-0.704 |
0.000 |
| C6 |
-0.087 |
1.740 |
0.000 |
| O7 |
-1.139 |
2.351 |
0.000 |
| H8 |
1.995 |
-2.188 |
0.000 |
| H9 |
-0.652 |
-2.933 |
0.000 |
| H10 |
-2.017 |
-0.547 |
0.000 |
| H11 |
0.896 |
2.258 |
0.000 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
C5 |
C6 |
O7 |
H8 |
H9 |
H10 |
H11 |
| O1 | | 1.3466 | 2.2370 | 1.3688 | 2.2420 | 2.4110 | 3.5442 | 2.0613 | 3.2767 | 3.2804 | 2.5484 |
C2 | 1.3466 | | 1.3677 | 2.1815 | 2.2244 | 3.5424 | 4.5371 | 1.0804 | 2.1900 | 3.2788 | 3.8648 | C3 | 2.2370 | 1.3677 | | 2.2351 | 1.4188 | 3.6765 | 4.3776 | 2.2493 | 1.0812 | 2.2541 | 4.3422 | C4 | 1.3688 | 2.1815 | 2.2351 | | 1.3722 | 1.4536 | 2.3557 | 3.1806 | 3.2871 | 2.1832 | 2.1630 | C5 | 2.2420 | 2.2244 | 1.4188 | 1.3722 | | 2.5908 | 3.0605 | 3.2958 | 2.2487 | 1.0809 | 3.4882 | C6 | 2.4110 | 3.5424 | 3.6765 | 1.4536 | 2.5908 | | 1.2170 | 4.4456 | 4.7069 | 2.9926 | 1.1105 | O7 | 3.5442 | 4.5371 | 4.3776 | 2.3557 | 3.0605 | 1.2170 | | 5.5160 | 5.3062 | 3.0277 | 2.0371 | H8 | 2.0613 | 1.0804 | 2.2493 | 3.1806 | 3.2958 | 4.4456 | 5.5160 | | 2.7498 | 4.3348 | 4.5799 | H9 | 3.2767 | 2.1900 | 1.0812 | 3.2871 | 2.2487 | 4.7069 | 5.3062 | 2.7498 | | 2.7489 | 5.4165 | H10 | 3.2804 | 3.2788 | 2.2541 | 2.1832 | 1.0809 | 2.9926 | 3.0277 | 4.3348 | 2.7489 | | 4.0434 | H11 | 2.5484 | 3.8648 | 4.3422 | 2.1630 | 3.4882 | 1.1105 | 2.0371 | 4.5799 | 5.4165 | 4.0434 | |
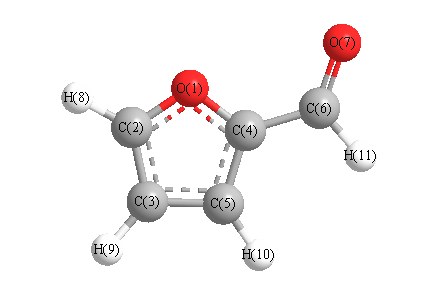 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C3 |
111.003 |
|
O1 |
C2 |
H8 |
115.845 |
| O1 |
C4 |
C5 |
109.763 |
|
O1 |
C4 |
C6 |
117.322 |
| C2 |
O1 |
C4 |
106.908 |
|
C2 |
C3 |
C5 |
105.914 |
| C2 |
C3 |
H9 |
126.422 |
|
C3 |
C2 |
H8 |
133.152 |
| C3 |
C5 |
C4 |
106.411 |
|
C3 |
C5 |
H10 |
128.263 |
| C4 |
C5 |
H10 |
125.326 |
|
C4 |
C6 |
O7 |
123.546 |
| C4 |
C6 |
H11 |
114.378 |
|
C5 |
C3 |
H9 |
127.663 |
| C5 |
C4 |
C6 |
132.915 |
|
O7 |
C6 |
H11 |
122.076 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.431 |
|
|
|
| 2 |
C |
0.084 |
|
|
|
| 3 |
C |
-0.226 |
|
|
|
| 4 |
C |
0.267 |
|
|
|
| 5 |
C |
-0.190 |
|
|
|
| 6 |
C |
0.171 |
|
|
|
| 7 |
O |
-0.414 |
|
|
|
| 8 |
H |
0.200 |
|
|
|
| 9 |
H |
0.189 |
|
|
|
| 10 |
H |
0.207 |
|
|
|
| 11 |
H |
0.143 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.482 |
-3.294 |
0.000 |
3.612 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-37.826 |
3.770 |
0.000 |
| y |
3.770 |
-39.122 |
0.000 |
| z |
0.000 |
0.000 |
-40.320 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.895 |
3.770 |
0.000 |
| y |
3.770 |
-0.049 |
0.000 |
| z |
0.000 |
0.000 |
-1.846 |
|
| Polar |
|---|
| 3z2-r2 | -3.693 |
|---|
| x2-y2 | 1.296 |
|---|
| xy | 3.770 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.633 |
-1.139 |
0.000 |
| y |
-1.139 |
12.209 |
0.000 |
| z |
0.000 |
0.000 |
3.331 |
<r2> (average value of r
2) Å
2
| <r2> |
192.501 |
| (<r2>)1/2 |
13.874 |
