Jump to
S1C2
Energy calculated at B2PLYP/6-31G*
| | hartrees |
|---|
| Energy at 0K | -343.006474 |
| Energy at 298.15K | -343.011268 |
| HF Energy | -342.676036 |
| Nuclear repulsion energy | 271.140846 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3328 |
3159 |
0.21 |
134.99 |
0.15 |
0.25 |
| 2 |
A' |
3305 |
3137 |
0.30 |
70.08 |
0.34 |
0.51 |
| 3 |
A' |
3292 |
3124 |
1.31 |
73.66 |
0.65 |
0.79 |
| 4 |
A' |
2966 |
2815 |
112.98 |
124.98 |
0.31 |
0.48 |
| 5 |
A' |
1773 |
1683 |
266.86 |
130.31 |
0.32 |
0.48 |
| 6 |
A' |
1623 |
1540 |
5.99 |
3.87 |
0.49 |
0.66 |
| 7 |
A' |
1530 |
1452 |
46.91 |
93.25 |
0.34 |
0.51 |
| 8 |
A' |
1453 |
1379 |
35.65 |
45.85 |
0.39 |
0.56 |
| 9 |
A' |
1419 |
1347 |
0.01 |
17.90 |
0.13 |
0.23 |
| 10 |
A' |
1320 |
1253 |
46.35 |
5.97 |
0.35 |
0.52 |
| 11 |
A' |
1268 |
1203 |
3.52 |
9.50 |
0.45 |
0.62 |
| 12 |
A' |
1212 |
1151 |
9.24 |
4.59 |
0.14 |
0.24 |
| 13 |
A' |
1124 |
1067 |
6.31 |
9.68 |
0.43 |
0.60 |
| 14 |
A' |
1062 |
1008 |
38.41 |
4.11 |
0.34 |
0.51 |
| 15 |
A' |
954 |
905 |
18.92 |
6.36 |
0.12 |
0.21 |
| 16 |
A' |
901 |
855 |
10.03 |
7.00 |
0.73 |
0.84 |
| 17 |
A' |
770 |
731 |
64.53 |
1.82 |
0.70 |
0.83 |
| 18 |
A' |
503 |
477 |
1.05 |
7.71 |
0.32 |
0.49 |
| 19 |
A' |
201 |
191 |
5.34 |
0.95 |
0.66 |
0.80 |
| 20 |
A" |
1007 |
956 |
0.05 |
6.27 |
0.75 |
0.86 |
| 21 |
A" |
863 |
819 |
0.85 |
1.48 |
0.75 |
0.86 |
| 22 |
A" |
825 |
783 |
8.34 |
0.44 |
0.75 |
0.86 |
| 23 |
A" |
773 |
734 |
53.09 |
0.78 |
0.75 |
0.86 |
| 24 |
A" |
645 |
612 |
3.32 |
2.92 |
0.75 |
0.86 |
| 25 |
A" |
602 |
571 |
5.34 |
0.12 |
0.75 |
0.86 |
| 26 |
A" |
286 |
272 |
10.97 |
0.68 |
0.75 |
0.86 |
| 27 |
A" |
134 |
127 |
1.44 |
1.71 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 17568.8 cm
-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 16676.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
0.502 |
-0.900 |
0.000 |
| C2 |
-0.562 |
-1.738 |
0.000 |
| C3 |
-1.739 |
-1.041 |
0.000 |
| C4 |
0.000 |
0.374 |
0.000 |
| C5 |
-1.374 |
0.334 |
0.000 |
| C6 |
0.927 |
1.497 |
0.000 |
| O7 |
2.144 |
1.402 |
0.000 |
| H8 |
-0.326 |
-2.790 |
0.000 |
| H9 |
-2.734 |
-1.459 |
0.000 |
| H10 |
-2.034 |
1.189 |
0.000 |
| H11 |
0.418 |
2.482 |
0.000 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
C5 |
C6 |
O7 |
H8 |
H9 |
H10 |
H11 |
| O1 | | 1.3546 | 2.2455 | 1.3695 | 2.2456 | 2.4338 | 2.8266 | 2.0643 | 3.2841 | 3.2862 | 3.3827 |
C2 | 1.3546 | | 1.3678 | 2.1856 | 2.2253 | 3.5607 | 4.1443 | 1.0786 | 2.1895 | 3.2766 | 4.3321 | C3 | 2.2455 | 1.3678 | | 2.2419 | 1.4223 | 3.6801 | 4.5868 | 2.2485 | 1.0794 | 2.2496 | 4.1306 | C4 | 1.3695 | 2.1856 | 2.2419 | | 1.3747 | 1.4555 | 2.3770 | 3.1815 | 3.2919 | 2.1916 | 2.1487 | C5 | 2.2456 | 2.2253 | 1.4223 | 1.3747 | | 2.5778 | 3.6762 | 3.2953 | 2.2507 | 1.0805 | 2.7974 | C6 | 2.4338 | 3.5607 | 3.6801 | 1.4555 | 2.5778 | | 1.2209 | 4.4665 | 4.7051 | 2.9769 | 1.1086 | O7 | 2.8266 | 4.1443 | 4.5868 | 2.3770 | 3.6762 | 1.2209 | | 4.8655 | 5.6546 | 4.1836 | 2.0359 | H8 | 2.0643 | 1.0786 | 2.2485 | 3.1815 | 3.2953 | 4.4665 | 4.8655 | | 2.7506 | 4.3307 | 5.3246 | H9 | 3.2841 | 2.1895 | 1.0794 | 3.2919 | 2.2507 | 4.7051 | 5.6546 | 2.7506 | | 2.7396 | 5.0467 | H10 | 3.2862 | 3.2766 | 2.2496 | 2.1916 | 1.0805 | 2.9769 | 4.1836 | 4.3307 | 2.7396 | | 2.7724 | H11 | 3.3827 | 4.3321 | 4.1306 | 2.1487 | 2.7974 | 1.1086 | 2.0359 | 5.3246 | 5.0467 | 2.7724 | |
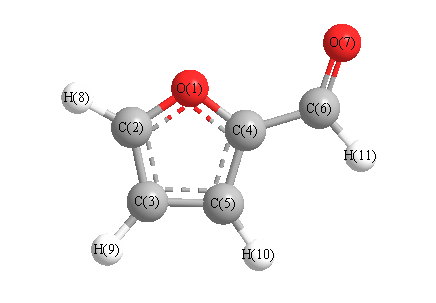 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C3 |
111.142 |
|
O1 |
C2 |
H8 |
115.609 |
| O1 |
C4 |
C5 |
109.834 |
|
O1 |
C4 |
C6 |
118.949 |
| C2 |
O1 |
C4 |
106.706 |
|
C2 |
C3 |
C5 |
105.778 |
| C2 |
C3 |
H9 |
126.534 |
|
C3 |
C2 |
H8 |
133.249 |
| C3 |
C5 |
C4 |
106.539 |
|
C3 |
C5 |
H10 |
127.470 |
| C4 |
C5 |
H10 |
125.992 |
|
C4 |
C6 |
O7 |
125.057 |
| C4 |
C6 |
H11 |
113.166 |
|
C5 |
C3 |
H9 |
127.687 |
| C5 |
C4 |
C6 |
131.217 |
|
O7 |
C6 |
H11 |
121.777 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.419 |
|
|
|
| 2 |
C |
0.098 |
|
|
|
| 3 |
C |
-0.208 |
|
|
|
| 4 |
C |
0.267 |
|
|
|
| 5 |
C |
-0.194 |
|
|
|
| 6 |
C |
0.184 |
|
|
|
| 7 |
O |
-0.398 |
|
|
|
| 8 |
H |
0.189 |
|
|
|
| 9 |
H |
0.177 |
|
|
|
| 10 |
H |
0.181 |
|
|
|
| 11 |
H |
0.124 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-3.952 |
-1.026 |
0.000 |
4.083 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-43.518 |
-2.582 |
0.004 |
| y |
-2.582 |
-33.455 |
0.000 |
| z |
0.004 |
0.000 |
-40.570 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.506 |
-2.582 |
0.004 |
| y |
-2.582 |
8.589 |
0.000 |
| z |
0.004 |
0.000 |
-2.083 |
|
| Polar |
|---|
| 3z2-r2 | -4.166 |
|---|
| x2-y2 | -10.063 |
|---|
| xy | -2.582 |
|---|
| xz | 0.004 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.462 |
1.847 |
-0.000 |
| y |
1.847 |
10.155 |
-0.000 |
| z |
-0.000 |
-0.000 |
3.278 |
<r2> (average value of r
2) Å
2
| <r2> |
190.914 |
| (<r2>)1/2 |
13.817 |
Jump to
S1C1
Energy calculated at B2PLYP/6-31G*
| | hartrees |
|---|
| Energy at 0K | -343.008212 |
| Energy at 298.15K | -343.012989 |
| HF Energy | -342.677891 |
| Nuclear repulsion energy | 270.288362 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3328 |
3159 |
0.29 |
142.48 |
0.15 |
0.26 |
| 2 |
A' |
3313 |
3144 |
0.27 |
41.65 |
0.23 |
0.37 |
| 3 |
A' |
3298 |
3131 |
1.02 |
86.57 |
0.70 |
0.82 |
| 4 |
A' |
2991 |
2839 |
103.55 |
132.82 |
0.31 |
0.48 |
| 5 |
A' |
1766 |
1676 |
213.94 |
98.03 |
0.34 |
0.51 |
| 6 |
A' |
1632 |
1549 |
55.56 |
23.45 |
0.35 |
0.52 |
| 7 |
A' |
1521 |
1444 |
109.96 |
172.65 |
0.32 |
0.48 |
| 8 |
A' |
1465 |
1391 |
1.82 |
6.88 |
0.75 |
0.86 |
| 9 |
A' |
1422 |
1349 |
13.59 |
29.27 |
0.39 |
0.56 |
| 10 |
A' |
1287 |
1222 |
30.01 |
4.26 |
0.41 |
0.58 |
| 11 |
A' |
1254 |
1190 |
3.02 |
4.16 |
0.35 |
0.52 |
| 12 |
A' |
1199 |
1138 |
11.22 |
6.26 |
0.12 |
0.22 |
| 13 |
A' |
1126 |
1069 |
17.95 |
12.06 |
0.45 |
0.62 |
| 14 |
A' |
1054 |
1000 |
35.57 |
2.21 |
0.22 |
0.36 |
| 15 |
A' |
970 |
921 |
8.70 |
6.62 |
0.15 |
0.26 |
| 16 |
A' |
901 |
855 |
6.83 |
6.16 |
0.75 |
0.86 |
| 17 |
A' |
761 |
722 |
72.63 |
2.75 |
0.66 |
0.80 |
| 18 |
A' |
497 |
472 |
0.91 |
6.31 |
0.36 |
0.53 |
| 19 |
A' |
203 |
192 |
6.70 |
0.29 |
0.43 |
0.60 |
| 20 |
A" |
1016 |
965 |
0.22 |
7.39 |
0.75 |
0.86 |
| 21 |
A" |
874 |
829 |
1.63 |
1.77 |
0.75 |
0.86 |
| 22 |
A" |
832 |
789 |
11.19 |
0.65 |
0.75 |
0.86 |
| 23 |
A" |
768 |
729 |
47.98 |
0.70 |
0.75 |
0.86 |
| 24 |
A" |
636 |
604 |
3.69 |
1.54 |
0.75 |
0.86 |
| 25 |
A" |
602 |
571 |
7.06 |
0.33 |
0.75 |
0.86 |
| 26 |
A" |
249 |
236 |
12.27 |
1.65 |
0.75 |
0.86 |
| 27 |
A" |
158 |
150 |
2.77 |
1.26 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 17560.4 cm
-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 16668.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
1.257 |
-0.265 |
0.000 |
| C2 |
1.089 |
-1.610 |
0.000 |
| C3 |
-0.241 |
-1.935 |
0.000 |
| C4 |
0.000 |
0.290 |
0.000 |
| C5 |
-0.948 |
-0.702 |
0.000 |
| C6 |
-0.085 |
1.743 |
0.000 |
| O7 |
-1.145 |
2.352 |
0.000 |
| H8 |
1.994 |
-2.196 |
0.000 |
| H9 |
-0.655 |
-2.932 |
0.000 |
| H10 |
-2.016 |
-0.545 |
0.000 |
| H11 |
0.892 |
2.263 |
0.000 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
C5 |
C6 |
O7 |
H8 |
H9 |
H10 |
H11 |
| O1 | | 1.3556 | 2.2436 | 1.3746 | 2.2488 | 2.4156 | 3.5530 | 2.0666 | 3.2819 | 3.2858 | 2.5542 |
C2 | 1.3556 | | 1.3685 | 2.1900 | 2.2302 | 3.5527 | 4.5488 | 1.0785 | 2.1882 | 3.2827 | 3.8780 | C3 | 2.2436 | 1.3685 | | 2.2384 | 1.4216 | 3.6816 | 4.3820 | 2.2500 | 1.0796 | 2.2553 | 4.3483 | C4 | 1.3746 | 2.1900 | 2.2384 | | 1.3728 | 1.4554 | 2.3589 | 3.1870 | 3.2882 | 2.1824 | 2.1651 | C5 | 2.2488 | 2.2302 | 1.4216 | 1.3728 | | 2.5934 | 3.0611 | 3.2999 | 2.2490 | 1.0795 | 3.4901 | C6 | 2.4156 | 3.5527 | 3.6816 | 1.4554 | 2.5934 | | 1.2228 | 4.4541 | 4.7099 | 2.9941 | 1.1068 | O7 | 3.5530 | 4.5488 | 4.3820 | 2.3589 | 3.0611 | 1.2228 | | 5.5265 | 5.3072 | 3.0253 | 2.0393 | H8 | 2.0666 | 1.0785 | 2.2500 | 3.1870 | 3.2999 | 4.4541 | 5.5265 | | 2.7497 | 4.3371 | 4.5930 | H9 | 3.2819 | 2.1882 | 1.0796 | 3.2882 | 2.2490 | 4.7099 | 5.3072 | 2.7497 | | 2.7482 | 5.4206 | H10 | 3.2858 | 3.2827 | 2.2553 | 2.1824 | 1.0795 | 2.9941 | 3.0253 | 4.3371 | 2.7482 | | 4.0426 | H11 | 2.5542 | 3.8780 | 4.3483 | 2.1651 | 3.4901 | 1.1068 | 2.0393 | 4.5930 | 5.4206 | 4.0426 | |
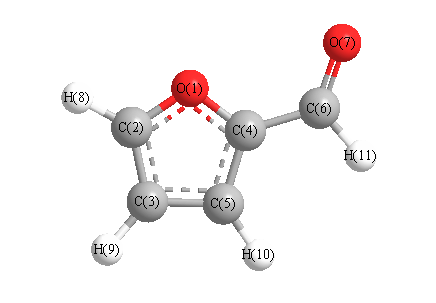 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C3 |
110.895 |
|
O1 |
C2 |
H8 |
115.744 |
| O1 |
C4 |
C5 |
109.876 |
|
O1 |
C4 |
C6 |
117.171 |
| C2 |
O1 |
C4 |
106.671 |
|
C2 |
C3 |
C5 |
106.117 |
| C2 |
C3 |
H9 |
126.314 |
|
C3 |
C2 |
H8 |
133.361 |
| C3 |
C5 |
C4 |
106.442 |
|
C3 |
C5 |
H10 |
128.253 |
| C4 |
C5 |
H10 |
125.304 |
|
C4 |
C6 |
O7 |
123.232 |
| C4 |
C6 |
H11 |
114.664 |
|
C5 |
C3 |
H9 |
127.569 |
| C5 |
C4 |
C6 |
132.953 |
|
O7 |
C6 |
H11 |
122.104 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.439 |
|
|
|
| 2 |
C |
0.092 |
|
|
|
| 3 |
C |
-0.206 |
|
|
|
| 4 |
C |
0.261 |
|
|
|
| 5 |
C |
-0.179 |
|
|
|
| 6 |
C |
0.183 |
|
|
|
| 7 |
O |
-0.404 |
|
|
|
| 8 |
H |
0.187 |
|
|
|
| 9 |
H |
0.176 |
|
|
|
| 10 |
H |
0.194 |
|
|
|
| 11 |
H |
0.133 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.437 |
-3.164 |
0.000 |
3.475 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.439 |
3.773 |
0.001 |
| y |
3.773 |
-39.447 |
0.003 |
| z |
0.001 |
0.003 |
-40.539 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.553 |
3.773 |
0.001 |
| y |
3.773 |
0.042 |
0.003 |
| z |
0.001 |
0.003 |
-1.596 |
|
| Polar |
|---|
| 3z2-r2 | -3.191 |
|---|
| x2-y2 | 1.007 |
|---|
| xy | 3.773 |
|---|
| xz | 0.001 |
|---|
| yz | 0.003 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.602 |
-1.114 |
0.000 |
| y |
-1.114 |
11.970 |
-0.000 |
| z |
0.000 |
-0.000 |
3.278 |
<r2> (average value of r
2) Å
2
| <r2> |
193.354 |
| (<r2>)1/2 |
13.905 |
