Jump to
S1C2
Energy calculated at HSEh1PBE/6-31G*
| | hartrees |
|---|
| Energy at 0K | -342.991695 |
| Energy at 298.15K | -342.996551 |
| HF Energy | -342.991695 |
| Nuclear repulsion energy | 272.200754 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3324 |
3162 |
0.27 |
141.78 |
0.13 |
0.23 |
| 2 |
A' |
3303 |
3141 |
0.16 |
59.03 |
0.37 |
0.54 |
| 3 |
A' |
3291 |
3130 |
1.12 |
77.86 |
0.68 |
0.81 |
| 4 |
A' |
2950 |
2806 |
112.82 |
123.89 |
0.32 |
0.49 |
| 5 |
A' |
1827 |
1737 |
313.74 |
123.55 |
0.32 |
0.49 |
| 6 |
A' |
1642 |
1562 |
7.80 |
7.51 |
0.46 |
0.63 |
| 7 |
A' |
1541 |
1465 |
51.57 |
94.08 |
0.33 |
0.50 |
| 8 |
A' |
1464 |
1393 |
39.92 |
39.39 |
0.43 |
0.60 |
| 9 |
A' |
1416 |
1347 |
0.17 |
13.81 |
0.12 |
0.21 |
| 10 |
A' |
1336 |
1271 |
45.67 |
6.43 |
0.35 |
0.51 |
| 11 |
A' |
1270 |
1208 |
3.28 |
9.89 |
0.49 |
0.66 |
| 12 |
A' |
1226 |
1166 |
6.75 |
3.12 |
0.12 |
0.22 |
| 13 |
A' |
1133 |
1077 |
10.59 |
10.52 |
0.34 |
0.51 |
| 14 |
A' |
1058 |
1006 |
41.17 |
3.40 |
0.36 |
0.53 |
| 15 |
A' |
967 |
920 |
18.68 |
6.12 |
0.12 |
0.22 |
| 16 |
A' |
903 |
859 |
11.06 |
7.09 |
0.74 |
0.85 |
| 17 |
A' |
776 |
738 |
63.95 |
1.54 |
0.66 |
0.79 |
| 18 |
A' |
506 |
481 |
1.11 |
7.81 |
0.33 |
0.49 |
| 19 |
A' |
199 |
189 |
5.56 |
0.93 |
0.67 |
0.80 |
| 20 |
A" |
1016 |
966 |
0.00 |
6.11 |
0.75 |
0.86 |
| 21 |
A" |
899 |
855 |
1.16 |
1.17 |
0.75 |
0.86 |
| 22 |
A" |
845 |
803 |
5.84 |
0.24 |
0.75 |
0.86 |
| 23 |
A" |
784 |
746 |
58.05 |
0.70 |
0.75 |
0.86 |
| 24 |
A" |
661 |
629 |
1.75 |
3.12 |
0.75 |
0.86 |
| 25 |
A" |
613 |
583 |
7.18 |
0.20 |
0.75 |
0.86 |
| 26 |
A" |
294 |
280 |
11.35 |
0.62 |
0.75 |
0.86 |
| 27 |
A" |
138 |
132 |
1.38 |
1.73 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 17691.1 cm
-1
Scaled (by 0.951) Zero Point Vibrational Energy (zpe) 16824.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
0.505 |
-0.894 |
0.000 |
| C2 |
-0.542 |
-1.735 |
0.000 |
| C3 |
-1.724 |
-1.052 |
0.000 |
| C4 |
0.000 |
0.369 |
0.000 |
| C5 |
-1.371 |
0.321 |
0.000 |
| C6 |
0.912 |
1.498 |
0.000 |
| O7 |
2.122 |
1.421 |
0.000 |
| H8 |
-0.299 |
-2.787 |
0.000 |
| H9 |
-2.716 |
-1.481 |
0.000 |
| H10 |
-2.037 |
1.174 |
0.000 |
| H11 |
0.387 |
2.478 |
0.000 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
C5 |
C6 |
O7 |
H8 |
H9 |
H10 |
H11 |
| O1 | | 1.3428 | 2.2351 | 1.3599 | 2.2350 | 2.4265 | 2.8235 | 2.0567 | 3.2744 | 3.2763 | 3.3736 |
C2 | 1.3428 | | 1.3652 | 2.1721 | 2.2166 | 3.5448 | 4.1296 | 1.0801 | 2.1891 | 3.2698 | 4.3135 | C3 | 2.2351 | 1.3652 | | 2.2345 | 1.4183 | 3.6682 | 4.5730 | 2.2454 | 1.0806 | 2.2477 | 4.1132 | C4 | 1.3599 | 2.1721 | 2.2345 | | 1.3717 | 1.4517 | 2.3685 | 3.1698 | 3.2864 | 2.1899 | 2.1441 | C5 | 2.2350 | 2.2166 | 1.4183 | 1.3717 | | 2.5684 | 3.6619 | 3.2879 | 2.2491 | 1.0816 | 2.7821 | C6 | 2.4265 | 3.5448 | 3.6682 | 1.4517 | 2.5684 | | 1.2126 | 4.4530 | 4.6947 | 2.9664 | 1.1113 | O7 | 2.8235 | 4.1296 | 4.5730 | 2.3685 | 3.6619 | 1.2126 | | 4.8544 | 5.6419 | 4.1661 | 2.0318 | H8 | 2.0567 | 1.0801 | 2.2454 | 3.1698 | 3.2879 | 4.4530 | 4.8544 | | 2.7478 | 4.3251 | 5.3091 | H9 | 3.2744 | 2.1891 | 1.0806 | 3.2864 | 2.2491 | 4.6947 | 5.6419 | 2.7478 | | 2.7403 | 5.0300 | H10 | 3.2763 | 3.2698 | 2.2477 | 2.1899 | 1.0816 | 2.9664 | 4.1661 | 4.3251 | 2.7403 | | 2.7521 | H11 | 3.3736 | 4.3135 | 4.1132 | 2.1441 | 2.7821 | 1.1113 | 2.0318 | 5.3091 | 5.0300 | 2.7521 | |
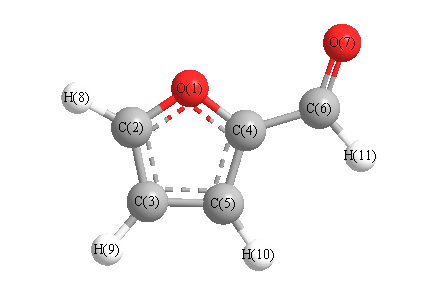 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C3 |
111.247 |
|
O1 |
C2 |
H8 |
115.753 |
| O1 |
C4 |
C5 |
109.811 |
|
O1 |
C4 |
C6 |
119.284 |
| C2 |
O1 |
C4 |
106.967 |
|
C2 |
C3 |
C5 |
105.548 |
| C2 |
C3 |
H9 |
126.638 |
|
C3 |
C2 |
H8 |
132.999 |
| C3 |
C5 |
C4 |
106.426 |
|
C3 |
C5 |
H10 |
127.571 |
| C4 |
C5 |
H10 |
126.003 |
|
C4 |
C6 |
O7 |
125.250 |
| C4 |
C6 |
H11 |
112.887 |
|
C5 |
C3 |
H9 |
127.813 |
| C5 |
C4 |
C6 |
130.905 |
|
O7 |
C6 |
H11 |
121.863 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.412 |
|
|
|
| 2 |
C |
0.090 |
|
|
|
| 3 |
C |
-0.232 |
|
|
|
| 4 |
C |
0.268 |
|
|
|
| 5 |
C |
-0.211 |
|
|
|
| 6 |
C |
0.178 |
|
|
|
| 7 |
O |
-0.408 |
|
|
|
| 8 |
H |
0.204 |
|
|
|
| 9 |
H |
0.192 |
|
|
|
| 10 |
H |
0.196 |
|
|
|
| 11 |
H |
0.135 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-3.976 |
-1.172 |
0.000 |
4.146 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-43.067 |
-2.788 |
0.000 |
| y |
-2.788 |
-33.104 |
0.000 |
| z |
0.000 |
0.000 |
-40.436 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.297 |
-2.788 |
0.000 |
| y |
-2.788 |
8.647 |
0.000 |
| z |
0.000 |
0.000 |
-2.350 |
|
| Polar |
|---|
| 3z2-r2 | -4.700 |
|---|
| x2-y2 | -9.962 |
|---|
| xy | -2.788 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.451 |
1.914 |
0.000 |
| y |
1.914 |
10.272 |
0.000 |
| z |
0.000 |
0.000 |
3.327 |
<r2> (average value of r
2) Å
2
| <r2> |
189.597 |
| (<r2>)1/2 |
13.769 |
Jump to
S1C1
Energy calculated at HSEh1PBE/6-31G*
| | hartrees |
|---|
| Energy at 0K | -342.993228 |
| Energy at 298.15K | -342.998068 |
| HF Energy | -342.993228 |
| Nuclear repulsion energy | 271.376597 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3325 |
3162 |
0.42 |
152.11 |
0.13 |
0.23 |
| 2 |
A' |
3313 |
3150 |
0.36 |
30.76 |
0.32 |
0.49 |
| 3 |
A' |
3296 |
3135 |
0.86 |
87.78 |
0.71 |
0.83 |
| 4 |
A' |
2971 |
2825 |
106.28 |
134.36 |
0.32 |
0.49 |
| 5 |
A' |
1821 |
1732 |
261.67 |
96.11 |
0.35 |
0.51 |
| 6 |
A' |
1649 |
1568 |
54.07 |
21.14 |
0.36 |
0.53 |
| 7 |
A' |
1532 |
1456 |
118.34 |
159.15 |
0.31 |
0.48 |
| 8 |
A' |
1474 |
1401 |
2.64 |
10.30 |
0.72 |
0.84 |
| 9 |
A' |
1424 |
1354 |
13.70 |
21.55 |
0.39 |
0.56 |
| 10 |
A' |
1289 |
1226 |
29.41 |
4.85 |
0.34 |
0.51 |
| 11 |
A' |
1273 |
1210 |
0.38 |
3.85 |
0.51 |
0.68 |
| 12 |
A' |
1211 |
1151 |
12.87 |
2.95 |
0.13 |
0.23 |
| 13 |
A' |
1138 |
1082 |
22.72 |
13.81 |
0.31 |
0.48 |
| 14 |
A' |
1050 |
999 |
39.04 |
1.72 |
0.22 |
0.36 |
| 15 |
A' |
981 |
933 |
6.74 |
6.48 |
0.16 |
0.28 |
| 16 |
A' |
902 |
858 |
7.46 |
6.20 |
0.75 |
0.86 |
| 17 |
A' |
763 |
725 |
71.90 |
2.88 |
0.60 |
0.75 |
| 18 |
A' |
503 |
478 |
1.10 |
6.32 |
0.37 |
0.54 |
| 19 |
A' |
202 |
192 |
6.71 |
0.30 |
0.42 |
0.59 |
| 20 |
A" |
1024 |
974 |
0.11 |
7.41 |
0.75 |
0.86 |
| 21 |
A" |
908 |
864 |
1.62 |
1.52 |
0.75 |
0.86 |
| 22 |
A" |
851 |
809 |
7.75 |
0.46 |
0.75 |
0.86 |
| 23 |
A" |
782 |
744 |
53.42 |
0.69 |
0.75 |
0.86 |
| 24 |
A" |
651 |
619 |
1.74 |
1.45 |
0.75 |
0.86 |
| 25 |
A" |
614 |
584 |
10.03 |
0.49 |
0.75 |
0.86 |
| 26 |
A" |
255 |
242 |
12.39 |
1.63 |
0.75 |
0.86 |
| 27 |
A" |
163 |
155 |
2.60 |
1.31 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 17681.1 cm
-1
Scaled (by 0.951) Zero Point Vibrational Energy (zpe) 16814.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
1.250 |
-0.259 |
0.000 |
| C2 |
1.094 |
-1.594 |
0.000 |
| C3 |
-0.230 |
-1.931 |
0.000 |
| C4 |
0.000 |
0.288 |
0.000 |
| C5 |
-0.942 |
-0.706 |
0.000 |
| C6 |
-0.095 |
1.737 |
0.000 |
| O7 |
-1.150 |
2.339 |
0.000 |
| H8 |
2.004 |
-2.175 |
0.000 |
| H9 |
-0.636 |
-2.933 |
0.000 |
| H10 |
-2.012 |
-0.552 |
0.000 |
| H11 |
0.883 |
2.260 |
0.000 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
C5 |
C6 |
O7 |
H8 |
H9 |
H10 |
H11 |
| O1 | | 1.3437 | 2.2331 | 1.3649 | 2.2381 | 2.4071 | 3.5371 | 2.0588 | 3.2723 | 3.2757 | 2.5459 |
C2 | 1.3437 | | 1.3659 | 2.1765 | 2.2214 | 3.5363 | 4.5275 | 1.0800 | 2.1878 | 3.2759 | 3.8596 | C3 | 2.2331 | 1.3659 | | 2.2311 | 1.4175 | 3.6704 | 4.3680 | 2.2469 | 1.0808 | 2.2535 | 4.3367 | C4 | 1.3649 | 2.1765 | 2.2311 | | 1.3699 | 1.4518 | 2.3511 | 3.1753 | 3.2829 | 2.1805 | 2.1609 | C5 | 2.2381 | 2.2214 | 1.4175 | 1.3699 | | 2.5855 | 3.0518 | 3.2924 | 2.2474 | 1.0806 | 3.4831 | C6 | 2.4071 | 3.5363 | 3.6704 | 1.4518 | 2.5855 | | 1.2145 | 4.4395 | 4.7005 | 2.9856 | 1.1099 | O7 | 3.5371 | 4.5275 | 4.3680 | 2.3511 | 3.0518 | 1.2145 | | 5.5065 | 5.2961 | 3.0167 | 2.0350 | H8 | 2.0588 | 1.0800 | 2.2469 | 3.1753 | 3.2924 | 4.4395 | 5.5065 | | 2.7469 | 4.3315 | 4.5747 | H9 | 3.2723 | 2.1878 | 1.0808 | 3.2829 | 2.2474 | 4.7005 | 5.2961 | 2.7469 | | 2.7491 | 5.4106 | H10 | 3.2757 | 3.2759 | 2.2535 | 2.1805 | 1.0806 | 2.9856 | 3.0167 | 4.3315 | 2.7491 | | 4.0365 | H11 | 2.5459 | 3.8596 | 4.3367 | 2.1609 | 3.4831 | 1.1099 | 2.0350 | 4.5747 | 5.4106 | 4.0365 | |
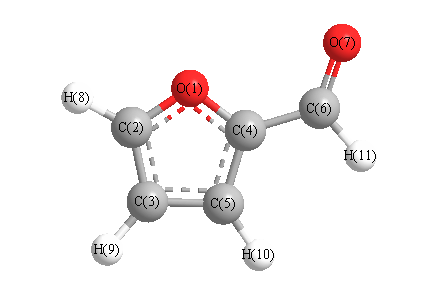 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C3 |
111.003 |
|
O1 |
C2 |
H8 |
115.881 |
| O1 |
C4 |
C5 |
109.843 |
|
O1 |
C4 |
C6 |
117.396 |
| C2 |
O1 |
C4 |
106.939 |
|
C2 |
C3 |
C5 |
105.881 |
| C2 |
C3 |
H9 |
126.420 |
|
C3 |
C2 |
H8 |
133.116 |
| C3 |
C5 |
C4 |
106.333 |
|
C3 |
C5 |
H10 |
128.370 |
| C4 |
C5 |
H10 |
125.296 |
|
C4 |
C6 |
O7 |
123.477 |
| C4 |
C6 |
H11 |
114.380 |
|
C5 |
C3 |
H9 |
127.698 |
| C5 |
C4 |
C6 |
132.760 |
|
O7 |
C6 |
H11 |
122.144 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.432 |
|
|
|
| 2 |
C |
0.084 |
|
|
|
| 3 |
C |
-0.231 |
|
|
|
| 4 |
C |
0.260 |
|
|
|
| 5 |
C |
-0.191 |
|
|
|
| 6 |
C |
0.175 |
|
|
|
| 7 |
O |
-0.413 |
|
|
|
| 8 |
H |
0.203 |
|
|
|
| 9 |
H |
0.192 |
|
|
|
| 10 |
H |
0.209 |
|
|
|
| 11 |
H |
0.144 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.509 |
-3.260 |
0.000 |
3.592 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-37.921 |
3.763 |
0.000 |
| y |
3.763 |
-39.117 |
0.000 |
| z |
0.000 |
0.000 |
-40.403 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.839 |
3.763 |
0.000 |
| y |
3.763 |
0.045 |
0.000 |
| z |
0.000 |
0.000 |
-1.884 |
|
| Polar |
|---|
| 3z2-r2 | -3.768 |
|---|
| x2-y2 | 1.196 |
|---|
| xy | 3.763 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.605 |
-1.141 |
0.000 |
| y |
-1.141 |
12.081 |
0.000 |
| z |
0.000 |
0.000 |
3.326 |
<r2> (average value of r
2) Å
2
| <r2> |
191.909 |
| (<r2>)1/2 |
13.853 |
