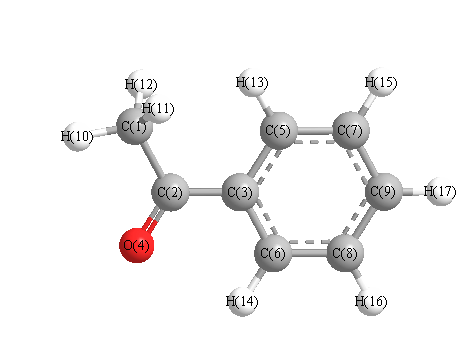
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3234 |
3094 |
6.29 |
|
|
|
| 2 |
A' |
3224 |
3084 |
14.43 |
|
|
|
| 3 |
A' |
3215 |
3076 |
23.36 |
|
|
|
| 4 |
A' |
3206 |
3067 |
7.24 |
|
|
|
| 5 |
A' |
3196 |
3057 |
0.07 |
|
|
|
| 6 |
A' |
3186 |
3048 |
10.14 |
|
|
|
| 7 |
A' |
3062 |
2930 |
2.30 |
|
|
|
| 8 |
A' |
1797 |
1719 |
164.39 |
|
|
|
| 9 |
A' |
1670 |
1598 |
20.44 |
|
|
|
| 10 |
A' |
1651 |
1579 |
7.74 |
|
|
|
| 11 |
A' |
1540 |
1473 |
1.32 |
|
|
|
| 12 |
A' |
1496 |
1431 |
19.52 |
|
|
|
| 13 |
A' |
1493 |
1428 |
9.25 |
|
|
|
| 14 |
A' |
1405 |
1344 |
60.26 |
|
|
|
| 15 |
A' |
1387 |
1327 |
3.29 |
|
|
|
| 16 |
A' |
1348 |
1290 |
4.13 |
|
|
|
| 17 |
A' |
1298 |
1242 |
182.61 |
|
|
|
| 18 |
A' |
1210 |
1158 |
13.71 |
|
|
|
| 19 |
A' |
1193 |
1142 |
0.64 |
|
|
|
| 20 |
A' |
1118 |
1069 |
6.68 |
|
|
|
| 21 |
A' |
1102 |
1055 |
1.25 |
|
|
|
| 22 |
A' |
1057 |
1011 |
4.69 |
|
|
|
| 23 |
A' |
1018 |
974 |
0.75 |
|
|
|
| 24 |
A' |
969 |
927 |
25.21 |
|
|
|
| 25 |
A' |
746 |
713 |
0.72 |
|
|
|
| 26 |
A' |
628 |
601 |
1.01 |
|
|
|
| 27 |
A' |
595 |
569 |
25.59 |
|
|
|
| 28 |
A' |
466 |
445 |
0.75 |
|
|
|
| 29 |
A' |
365 |
349 |
0.81 |
|
|
|
| 30 |
A' |
215 |
205 |
4.88 |
|
|
|
| 31 |
A" |
3131 |
2996 |
7.63 |
|
|
|
| 32 |
A" |
1502 |
1437 |
11.35 |
|
|
|
| 33 |
A" |
1053 |
1007 |
1.67 |
|
|
|
| 34 |
A" |
1011 |
967 |
0.43 |
|
|
|
| 35 |
A" |
988 |
945 |
0.38 |
|
|
|
| 36 |
A" |
946 |
905 |
2.46 |
|
|
|
| 37 |
A" |
867 |
829 |
0.18 |
|
|
|
| 38 |
A" |
777 |
744 |
34.85 |
|
|
|
| 39 |
A" |
706 |
675 |
30.23 |
|
|
|
| 40 |
A" |
602 |
576 |
9.32 |
|
|
|
| 41 |
A" |
429 |
410 |
0.06 |
|
|
|
| 42 |
A" |
413 |
395 |
0.01 |
|
|
|
| 43 |
A" |
167 |
160 |
0.34 |
|
|
|
| 44 |
A" |
154 |
147 |
0.02 |
|
|
|
| 45 |
A" |
64 |
61 |
3.33 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30447.2 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 29128.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.618 |
|
|
|
| 2 |
C |
0.424 |
|
|
|
| 3 |
C |
0.039 |
|
|
|
| 4 |
O |
-0.454 |
|
|
|
| 5 |
C |
-0.203 |
|
|
|
| 6 |
C |
-0.179 |
|
|
|
| 7 |
C |
-0.167 |
|
|
|
| 8 |
C |
-0.166 |
|
|
|
| 9 |
C |
-0.152 |
|
|
|
| 10 |
H |
0.209 |
|
|
|
| 11 |
H |
0.195 |
|
|
|
| 12 |
H |
0.195 |
|
|
|
| 13 |
H |
0.167 |
|
|
|
| 14 |
H |
0.198 |
|
|
|
| 15 |
H |
0.169 |
|
|
|
| 16 |
H |
0.172 |
|
|
|
| 17 |
H |
0.171 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.850 |
-2.368 |
0.000 |
3.005 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-46.508 |
5.416 |
0.000 |
| y |
5.416 |
-52.481 |
0.000 |
| z |
0.000 |
0.000 |
-54.157 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
6.811 |
5.416 |
0.000 |
| y |
5.416 |
-2.149 |
0.000 |
| z |
0.000 |
0.000 |
-4.662 |
|
| Polar |
|---|
| 3z2-r2 | -9.325 |
|---|
| x2-y2 | 5.973 |
|---|
| xy | 5.416 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
13.658 |
0.628 |
0.000 |
| y |
0.628 |
16.809 |
0.000 |
| z |
0.000 |
0.000 |
5.308 |
<r2> (average value of r
2) Å
2
| <r2> |
344.949 |
| (<r2>)1/2 |
18.573 |