return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3COC6H5 (acetophenone)

using model chemistry: QCISD/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at QCISD/6-31G*
 hartrees
Energy at 0K-383.714102
Energy at 298.15K-383.722309
HF Energy-382.472885
Nuclear repulsion energy400.265826
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3237 3083 3.95      
2 A' 3224 3071 12.32      
3 A' 3215 3062 24.43      
4 A' 3205 3052 9.21      
5 A' 3194 3042 0.04      
6 A' 3188 3036 10.81      
7 A' 3073 2927 2.59      
8 A' 1794 1708 130.36      
9 A' 1696 1615 12.07      
10 A' 1674 1594 7.66      
11 A' 1563 1489 0.67      
12 A' 1525 1452 8.98      
13 A' 1513 1441 19.99      
14 A' 1440 1371 49.33      
15 A' 1371 1306 4.67      
16 A' 1331 1267 78.80      
17 A' 1320 1257 74.82      
18 A' 1233 1174 7.91      
19 A' 1205 1148 0.65      
20 A' 1133 1079 4.55      
21 A' 1121 1068 1.05      
22 A' 1067 1016 3.18      
23 A' 1026 977 0.38      
24 A' 989 942 17.74      
25 A' 753 717 1.50      
26 A' 634 604 0.68      
27 A' 601 573 27.20      
28 A' 469 447 0.75      
29 A' 372 354 0.79      
30 A' 225 214 4.89      
31 A" 3145 2995 9.20      
32 A" 1527 1454 10.08      
33 A" 1066 1015 1.49      
34 A" 944 899 0.86      
35 A" 926 882 0.38      
36 A" 903 860 0.74      
37 A" 856 815 0.27      
38 A" 752 716 61.28      
39 A" 621 592 18.04      
40 A" 568 541 2.02      
41 A" 418 398 0.03      
42 A" 403 384 0.00      
43 A" 183 174 0.14      
44 A" 154 147 0.02      
45 A" 52 50 3.37      

Unscaled Zero Point Vibrational Energy (zpe) 30454.9 cm-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 29002.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/6-31G*
ABC
0.12257 0.04038 0.03055

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.664 2.210 0.000
C2 0.237 1.695 0.000
C3 0.000 0.210 0.000
O4 -0.708 2.475 0.000
C5 1.048 -0.720 0.000
C6 -1.327 -0.246 0.000
C7 0.772 -2.090 0.000
C8 -1.600 -1.612 0.000
C9 -0.550 -2.537 0.000
H10 1.637 3.302 0.000
H11 2.206 1.857 0.887
H12 2.206 1.857 -0.887
H13 2.084 -0.388 0.000
H14 -2.129 0.488 0.000
H15 1.592 -2.807 0.000
H16 -2.633 -1.959 0.000
H17 -0.764 -3.605 0.000

Atom - Atom Distances (Å)
  C1 C2 C3 O4 C5 C6 C7 C8 C9 H10 H11 H12 H13 H14 H15 H16 H17
C11.51712.60132.38762.99383.87014.39095.02645.23781.09301.09731.09732.63114.16525.01715.98686.3013
C21.51711.50321.22642.54722.49273.82223.78324.30442.13132.16552.16552.78362.65574.70114.64615.3935
C32.60131.50322.37331.40141.40312.42642.42552.80203.49832.89222.89222.16842.14663.41153.41143.8912
O42.38761.22642.37333.64642.79114.79954.18395.01502.48663.10873.10873.99972.44255.76164.83416.0808
C52.99382.54721.40143.64642.42151.39722.79462.41994.06522.96132.96131.08833.39862.15663.88363.4067
C63.87012.49271.40312.79112.42152.79381.39312.41884.62324.20624.20623.41391.08743.88272.15393.4057
C74.39093.82222.42644.79951.39722.79382.42041.39635.46094.29184.29182.14893.88101.08903.40762.1578
C85.02643.78322.42554.18392.79461.39312.42041.39935.88485.22615.22613.88282.16593.40861.08902.1614
C95.23784.30442.80205.01502.41992.41881.39631.39936.23545.26235.26233.40003.41232.15912.16111.0892
H101.09302.13133.49832.48664.06524.62325.46095.88486.23541.78861.78863.71714.70066.10946.77567.3126
H111.09732.16552.89223.10872.96134.20624.29185.22615.26231.78861.77312.41664.63144.78716.22616.2803
H121.09732.16552.89223.10872.96134.20624.29185.22615.26231.78861.77312.41664.63144.78716.22616.2803
H132.63112.78362.16843.99971.08833.41392.14893.88283.40003.71712.41662.41664.30292.46884.97174.2968
H144.16522.65572.14662.44253.39861.08743.88102.16593.41234.70064.63144.63144.30294.97002.49884.3149
H155.01714.70113.41155.76162.15663.88271.08903.40862.15916.10944.78714.78712.46884.97004.30882.4872
H165.98684.64613.41144.83413.88362.15393.40761.08902.16116.77566.22616.22614.97172.49884.30882.4904
H176.30135.39353.89126.08083.40673.40572.15782.16141.08927.31266.28036.28034.29684.31492.48722.4904

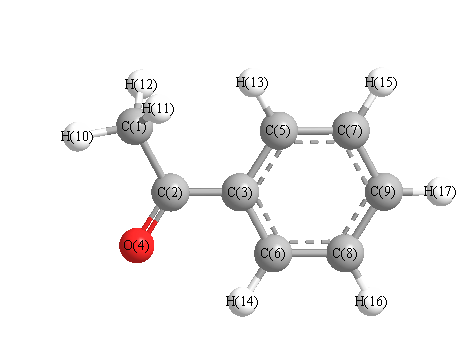
picture of acetophenone state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 118.920 C1 C2 O4 120.618
C2 C1 H10 108.389 C2 C1 H11 110.826
C2 C1 H12 110.826 C2 C3 C5 122.520
C2 C3 C6 118.074 C3 C2 O4 120.462
C3 C5 C7 120.228 C3 C5 H13 120.622
C3 C6 C8 120.322 C3 C6 H14 118.514
C5 C3 C6 119.407 C5 C7 C9 120.054
C5 C7 H15 119.818 C6 C8 C9 120.040
C6 C8 H16 119.906 C7 C5 H13 119.150
C7 C9 C8 119.949 C7 C9 H17 119.987
C8 C6 H14 121.163 C8 C9 H17 120.064
C9 C7 H15 120.128 C9 C8 H16 120.054
H10 C1 H11 109.494 H10 C1 H12 109.494
H11 C1 H12 107.799
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability