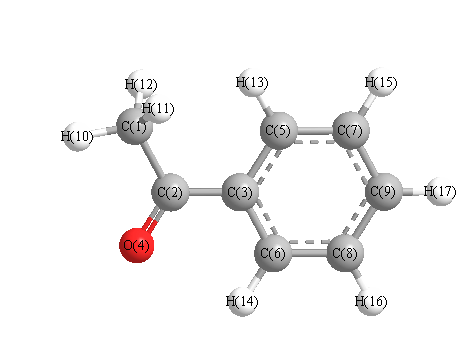
Vibrational Frequencies calculated at HSEh1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3248 |
3089 |
6.56 |
|
|
|
| 2 |
A' |
3239 |
3080 |
14.01 |
|
|
|
| 3 |
A' |
3230 |
3072 |
21.61 |
|
|
|
| 4 |
A' |
3221 |
3064 |
4.97 |
|
|
|
| 5 |
A' |
3212 |
3054 |
0.17 |
|
|
|
| 6 |
A' |
3202 |
3045 |
9.23 |
|
|
|
| 7 |
A' |
3075 |
2924 |
2.07 |
|
|
|
| 8 |
A' |
1817 |
1728 |
168.45 |
|
|
|
| 9 |
A' |
1683 |
1601 |
20.68 |
|
|
|
| 10 |
A' |
1664 |
1582 |
7.68 |
|
|
|
| 11 |
A' |
1548 |
1472 |
1.77 |
|
|
|
| 12 |
A' |
1503 |
1429 |
20.38 |
|
|
|
| 13 |
A' |
1494 |
1421 |
8.88 |
|
|
|
| 14 |
A' |
1409 |
1340 |
70.54 |
|
|
|
| 15 |
A' |
1396 |
1328 |
3.88 |
|
|
|
| 16 |
A' |
1352 |
1285 |
3.34 |
|
|
|
| 17 |
A' |
1310 |
1246 |
174.69 |
|
|
|
| 18 |
A' |
1213 |
1154 |
13.00 |
|
|
|
| 19 |
A' |
1195 |
1136 |
0.65 |
|
|
|
| 20 |
A' |
1122 |
1067 |
7.19 |
|
|
|
| 21 |
A' |
1108 |
1054 |
1.21 |
|
|
|
| 22 |
A' |
1063 |
1011 |
4.96 |
|
|
|
| 23 |
A' |
1023 |
973 |
0.73 |
|
|
|
| 24 |
A' |
975 |
927 |
22.21 |
|
|
|
| 25 |
A' |
751 |
714 |
0.82 |
|
|
|
| 26 |
A' |
630 |
599 |
1.01 |
|
|
|
| 27 |
A' |
598 |
569 |
26.30 |
|
|
|
| 28 |
A' |
468 |
445 |
0.78 |
|
|
|
| 29 |
A' |
367 |
349 |
0.86 |
|
|
|
| 30 |
A' |
217 |
206 |
4.96 |
|
|
|
| 31 |
A" |
3146 |
2992 |
6.63 |
|
|
|
| 32 |
A" |
1503 |
1430 |
11.69 |
|
|
|
| 33 |
A" |
1056 |
1004 |
1.64 |
|
|
|
| 34 |
A" |
1016 |
967 |
0.44 |
|
|
|
| 35 |
A" |
993 |
944 |
0.39 |
|
|
|
| 36 |
A" |
950 |
904 |
2.49 |
|
|
|
| 37 |
A" |
872 |
829 |
0.18 |
|
|
|
| 38 |
A" |
780 |
742 |
34.92 |
|
|
|
| 39 |
A" |
710 |
675 |
30.52 |
|
|
|
| 40 |
A" |
604 |
575 |
9.60 |
|
|
|
| 41 |
A" |
430 |
409 |
0.05 |
|
|
|
| 42 |
A" |
415 |
395 |
0.01 |
|
|
|
| 43 |
A" |
170 |
162 |
0.33 |
|
|
|
| 44 |
A" |
154 |
147 |
0.02 |
|
|
|
| 45 |
A" |
66 |
63 |
3.36 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30598.9 cm
-1
Scaled (by 0.951) Zero Point Vibrational Energy (zpe) 29099.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.622 |
|
|
|
| 2 |
C |
0.425 |
|
|
|
| 3 |
C |
0.028 |
|
|
|
| 4 |
O |
-0.452 |
|
|
|
| 5 |
C |
-0.201 |
|
|
|
| 6 |
C |
-0.178 |
|
|
|
| 7 |
C |
-0.171 |
|
|
|
| 8 |
C |
-0.169 |
|
|
|
| 9 |
C |
-0.155 |
|
|
|
| 10 |
H |
0.211 |
|
|
|
| 11 |
H |
0.197 |
|
|
|
| 12 |
H |
0.197 |
|
|
|
| 13 |
H |
0.170 |
|
|
|
| 14 |
H |
0.201 |
|
|
|
| 15 |
H |
0.172 |
|
|
|
| 16 |
H |
0.175 |
|
|
|
| 17 |
H |
0.174 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.851 |
-2.365 |
0.000 |
3.003 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-46.601 |
5.403 |
0.000 |
| y |
5.403 |
-52.567 |
0.000 |
| z |
0.000 |
0.000 |
-54.288 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
6.826 |
5.403 |
0.000 |
| y |
5.403 |
-2.122 |
0.000 |
| z |
0.000 |
0.000 |
-4.704 |
|
| Polar |
|---|
| 3z2-r2 | -9.408 |
|---|
| x2-y2 | 5.965 |
|---|
| xy | 5.403 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
13.591 |
0.608 |
0.000 |
| y |
0.608 |
16.681 |
0.000 |
| z |
0.000 |
0.000 |
5.297 |
<r2> (average value of r
2) Å
2
| <r2> |
343.824 |
| (<r2>)1/2 |
18.542 |