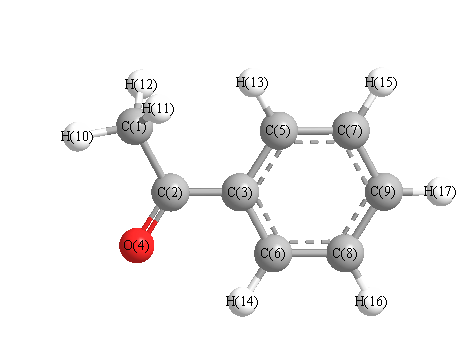
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3148 |
3096 |
9.58 |
|
|
|
| 2 |
A' |
3141 |
3089 |
15.73 |
|
|
|
| 3 |
A' |
3133 |
3081 |
21.73 |
|
|
|
| 4 |
A' |
3124 |
3072 |
6.38 |
|
|
|
| 5 |
A' |
3114 |
3063 |
0.12 |
|
|
|
| 6 |
A' |
3110 |
3059 |
10.50 |
|
|
|
| 7 |
A' |
2987 |
2937 |
2.40 |
|
|
|
| 8 |
A' |
1719 |
1691 |
137.74 |
|
|
|
| 9 |
A' |
1613 |
1587 |
21.24 |
|
|
|
| 10 |
A' |
1595 |
1569 |
7.85 |
|
|
|
| 11 |
A' |
1490 |
1466 |
1.64 |
|
|
|
| 12 |
A' |
1451 |
1427 |
18.03 |
|
|
|
| 13 |
A' |
1449 |
1425 |
9.32 |
|
|
|
| 14 |
A' |
1374 |
1351 |
2.03 |
|
|
|
| 15 |
A' |
1354 |
1331 |
56.39 |
|
|
|
| 16 |
A' |
1305 |
1283 |
3.93 |
|
|
|
| 17 |
A' |
1256 |
1235 |
181.11 |
|
|
|
| 18 |
A' |
1175 |
1156 |
16.69 |
|
|
|
| 19 |
A' |
1160 |
1141 |
1.02 |
|
|
|
| 20 |
A' |
1086 |
1068 |
6.95 |
|
|
|
| 21 |
A' |
1067 |
1050 |
0.82 |
|
|
|
| 22 |
A' |
1028 |
1011 |
4.78 |
|
|
|
| 23 |
A' |
989 |
973 |
0.84 |
|
|
|
| 24 |
A' |
936 |
920 |
28.66 |
|
|
|
| 25 |
A' |
724 |
712 |
0.60 |
|
|
|
| 26 |
A' |
611 |
600 |
0.88 |
|
|
|
| 27 |
A' |
577 |
567 |
21.83 |
|
|
|
| 28 |
A' |
452 |
445 |
0.60 |
|
|
|
| 29 |
A' |
356 |
350 |
0.79 |
|
|
|
| 30 |
A' |
211 |
208 |
4.59 |
|
|
|
| 31 |
A" |
3053 |
3003 |
7.05 |
|
|
|
| 32 |
A" |
1457 |
1433 |
10.63 |
|
|
|
| 33 |
A" |
1016 |
999 |
1.47 |
|
|
|
| 34 |
A" |
968 |
952 |
0.48 |
|
|
|
| 35 |
A" |
943 |
928 |
0.43 |
|
|
|
| 36 |
A" |
906 |
891 |
2.00 |
|
|
|
| 37 |
A" |
834 |
820 |
0.18 |
|
|
|
| 38 |
A" |
751 |
739 |
30.11 |
|
|
|
| 39 |
A" |
683 |
672 |
27.61 |
|
|
|
| 40 |
A" |
583 |
573 |
7.20 |
|
|
|
| 41 |
A" |
414 |
407 |
0.09 |
|
|
|
| 42 |
A" |
398 |
392 |
0.00 |
|
|
|
| 43 |
A" |
165 |
162 |
0.48 |
|
|
|
| 44 |
A" |
148 |
145 |
0.01 |
|
|
|
| 45 |
A" |
66 |
65 |
2.89 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 29559.0 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 29071.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.587 |
|
|
|
| 2 |
C |
0.378 |
|
|
|
| 3 |
C |
0.081 |
|
|
|
| 4 |
O |
-0.420 |
|
|
|
| 5 |
C |
-0.186 |
|
|
|
| 6 |
C |
-0.164 |
|
|
|
| 7 |
C |
-0.142 |
|
|
|
| 8 |
C |
-0.140 |
|
|
|
| 9 |
C |
-0.128 |
|
|
|
| 10 |
H |
0.193 |
|
|
|
| 11 |
H |
0.185 |
|
|
|
| 12 |
H |
0.185 |
|
|
|
| 13 |
H |
0.140 |
|
|
|
| 14 |
H |
0.168 |
|
|
|
| 15 |
H |
0.144 |
|
|
|
| 16 |
H |
0.147 |
|
|
|
| 17 |
H |
0.146 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.772 |
-2.302 |
0.000 |
2.905 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-46.971 |
5.213 |
0.000 |
| y |
5.213 |
-52.538 |
0.000 |
| z |
0.000 |
0.000 |
-54.186 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
6.391 |
5.213 |
0.000 |
| y |
5.213 |
-1.960 |
0.000 |
| z |
0.000 |
0.000 |
-4.431 |
|
| Polar |
|---|
| 3z2-r2 | -8.862 |
|---|
| x2-y2 | 5.567 |
|---|
| xy | 5.213 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
14.100 |
0.776 |
0.000 |
| y |
0.776 |
17.649 |
0.000 |
| z |
0.000 |
0.000 |
5.373 |
<r2> (average value of r
2) Å
2
| <r2> |
348.594 |
| (<r2>)1/2 |
18.671 |