return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H5NO2 (Nitrobenzene)

using model chemistry: MP3/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP3/6-31G*
 hartrees
Energy at 0K-435.469736
Energy at 298.15K-435.476699
HF Energy-434.171093
Nuclear repulsion energy412.822750
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP3/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3294 3092 2.99      
2 A1 3263 3063 9.52      
3 A1 3241 3042 0.59      
4 A1 1701 1596 1.76      
5 A1 1567 1471 0.53      
6 A1 1513 1420 205.75      
7 A1 1238 1162 1.47      
8 A1 1182 1110 14.08      
9 A1 1075 1009 4.71      
10 A1 1038 974 0.28      
11 A1 891 836 35.05      
12 A1 712 668 16.89      
13 A1 414 388 2.97      
14 A2 936 879 0.00      
15 A2 851 798 0.00      
16 A2 405 380 0.00      
17 A2 39 37 0.00      
18 B1 928 871 1.02      
19 B1 910 854 2.18      
20 B1 784 736 21.10      
21 B1 717 673 85.88      
22 B1 544 511 0.00      
23 B1 429 403 0.14      
24 B1 174 164 1.13      
25 B2 3294 3091 0.93      
26 B2 3254 3054 10.23      
27 B2 1804 1693 228.17      
28 B2 1703 1599 37.55      
29 B2 1537 1443 3.95      
30 B2 1374 1289 0.09      
31 B2 1344 1261 11.82      
32 B2 1214 1140 1.24      
33 B2 1136 1066 6.16      
34 B2 635 596 0.03      
35 B2 540 507 1.35      
36 B2 271 255 1.24      

Unscaled Zero Point Vibrational Energy (zpe) 22974.8 cm-1
Scaled (by 0.9386) Zero Point Vibrational Energy (zpe) 21564.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP3/6-31G*
ABC
0.13266 0.04308 0.03252

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP3/6-31G*

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 0.243
C2 0.000 1.222 -0.423
C3 0.000 1.210 -1.816
C4 0.000 0.000 -2.512
C5 0.000 -1.210 -1.816
C6 0.000 -1.222 -0.423
N7 0.000 0.000 1.713
O8 0.000 -1.083 2.282
O9 0.000 1.083 2.282
H10 0.000 2.146 0.141
H11 0.000 2.151 -2.358
H12 0.000 0.000 -3.598
H13 0.000 -2.151 -2.358
H14 0.000 -2.146 0.141

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 N7 O8 O9 H10 H11 H12 H13 H14
C11.39142.38852.75512.38851.39141.46962.30852.30852.14893.37523.84113.37522.1489
C21.39141.39312.42002.80272.44342.46033.55332.70851.08302.14653.40213.88843.4150
C32.38851.39311.39612.42062.80273.73054.69594.10002.16921.08572.15423.40483.8855
C42.75512.42001.39611.39612.42004.22464.91474.91473.41242.15671.08612.15673.4124
C52.38852.80272.42061.39611.39313.73054.10004.69593.88553.40482.15421.08572.1692
C61.39142.44342.80272.42001.39312.46032.70853.55333.41503.88843.40212.14651.0830
N71.46962.46033.73054.22463.73052.46031.22311.22312.66054.60395.31074.60392.6605
O82.30853.55334.69594.91474.10002.70851.22312.16513.87445.65555.97894.76132.3910
O92.30852.70854.10004.91474.69593.55331.22312.16512.39104.76135.97895.65553.8744
H102.14891.08302.16923.41243.88553.41502.66053.87442.39102.49854.31124.97114.2929
H113.37522.14651.08572.15673.40483.88844.60395.65554.76132.49852.48314.30234.9711
H123.84113.40212.15421.08612.15423.40215.31075.97895.97894.31122.48312.48314.3112
H133.37523.88843.40482.15671.08572.14654.60394.76135.65554.97114.30232.48312.4985
H142.14893.41503.88553.41242.16921.08302.66052.39103.87444.29294.97114.31122.4985

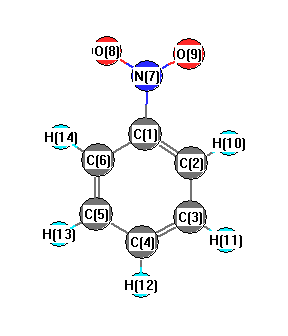
picture of Nitrobenzene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 118.129 C1 C2 H10 120.037
C1 C6 C5 118.129 C1 C6 H14 120.037
C1 N7 O8 117.742 C1 N7 O9 117.742
C2 C1 C6 122.805 C2 C1 N7 118.597
C2 C3 C4 120.369 C2 C3 H11 119.463
C3 C2 H10 121.835 C3 C4 C5 120.200
C3 C4 H12 119.900 C4 C3 H11 120.168
C4 C5 C6 120.369 C4 C5 H13 120.168
C5 C4 H12 119.900 C5 C6 H14 121.835
C6 C1 N7 118.597 C6 C5 H13 119.463
O8 N7 O9 124.515
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability