Jump to
S1C2
Energy calculated at HSEh1PBE/6-311+G(3df,2p)
| | hartrees |
|---|
| Energy at 0K | -191.770441 |
| Energy at 298.15K | |
| HF Energy | -191.770441 |
| Nuclear repulsion energy | 103.553598 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-311+G(3df,2p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3242 |
3242 |
3.16 |
64.78 |
0.51 |
0.68 |
| 2 |
A' |
3189 |
3189 |
0.95 |
100.35 |
0.29 |
0.44 |
| 3 |
A' |
3146 |
3146 |
4.30 |
78.95 |
0.09 |
0.16 |
| 4 |
A' |
2892 |
2892 |
84.96 |
104.06 |
0.28 |
0.44 |
| 5 |
A' |
1812 |
1812 |
274.42 |
81.37 |
0.38 |
0.55 |
| 6 |
A' |
1693 |
1693 |
2.67 |
52.33 |
0.12 |
0.22 |
| 7 |
A' |
1453 |
1453 |
14.04 |
5.54 |
0.50 |
0.67 |
| 8 |
A' |
1388 |
1388 |
6.79 |
20.12 |
0.30 |
0.46 |
| 9 |
A' |
1297 |
1297 |
3.66 |
9.15 |
0.16 |
0.27 |
| 10 |
A' |
1178 |
1178 |
39.01 |
12.35 |
0.61 |
0.76 |
| 11 |
A' |
925 |
925 |
19.43 |
1.52 |
0.00 |
0.00 |
| 12 |
A' |
576 |
576 |
5.05 |
6.02 |
0.17 |
0.29 |
| 13 |
A' |
320 |
320 |
10.31 |
1.20 |
0.32 |
0.49 |
| 14 |
A" |
1041 |
1041 |
8.23 |
0.62 |
0.75 |
0.86 |
| 15 |
A" |
1035 |
1035 |
5.14 |
2.21 |
0.75 |
0.86 |
| 16 |
A" |
1005 |
1005 |
45.01 |
2.43 |
0.75 |
0.86 |
| 17 |
A" |
616 |
616 |
10.49 |
0.60 |
0.75 |
0.86 |
| 18 |
A" |
168 |
168 |
4.44 |
0.51 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 13488.4 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 13488.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-311+G(3df,2p)
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.153 |
-0.739 |
0.000 |
| C2 |
0.000 |
0.719 |
0.000 |
| C3 |
1.209 |
1.271 |
0.000 |
| O4 |
-1.211 |
-1.313 |
0.000 |
| H5 |
0.804 |
-1.305 |
0.000 |
| H6 |
-0.911 |
1.308 |
0.000 |
| H7 |
1.358 |
2.343 |
0.000 |
| H8 |
2.100 |
0.651 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.4666 | 2.4284 | 1.2034 | 1.1114 | 2.1828 | 3.4326 | 2.6477 |
C2 | 1.4666 | | 1.3293 | 2.3654 | 2.1776 | 1.0844 | 2.1165 | 2.1014 | C3 | 2.4284 | 1.3293 | | 3.5402 | 2.6070 | 2.1208 | 1.0823 | 1.0854 | O4 | 1.2034 | 2.3654 | 3.5402 | | 2.0152 | 2.6372 | 4.4678 | 3.8498 | H5 | 1.1114 | 2.1776 | 2.6070 | 2.0152 | | 3.1247 | 3.6892 | 2.3460 | H6 | 2.1828 | 1.0844 | 2.1208 | 2.6372 | 3.1247 | | 2.4938 | 3.0821 | H7 | 3.4326 | 2.1165 | 1.0823 | 4.4678 | 3.6892 | 2.4938 | | 1.8479 | H8 | 2.6477 | 2.1014 | 1.0854 | 3.8498 | 2.3460 | 3.0821 | 1.8479 | |
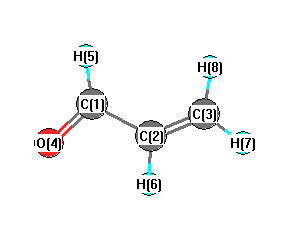 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
120.503 |
|
C1 |
C2 |
H6 |
116.863 |
| C2 |
C1 |
O4 |
124.430 |
|
C2 |
C1 |
H5 |
114.573 |
| C2 |
C3 |
H7 |
122.386 |
|
C2 |
C3 |
H8 |
120.651 |
| C3 |
C2 |
H6 |
122.635 |
|
O4 |
C1 |
H5 |
120.997 |
| H7 |
C3 |
H8 |
116.963 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-311+G(3df,2p)
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.451 |
|
|
|
| 2 |
C |
0.036 |
|
|
|
| 3 |
C |
-0.323 |
|
|
|
| 4 |
O |
-0.632 |
|
|
|
| 5 |
H |
0.090 |
|
|
|
| 6 |
H |
0.116 |
|
|
|
| 7 |
H |
0.135 |
|
|
|
| 8 |
H |
0.127 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.577 |
2.189 |
0.000 |
3.382 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-24.675 |
-2.710 |
0.000 |
| y |
-2.710 |
-23.985 |
0.000 |
| z |
0.000 |
0.000 |
-25.066 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.149 |
-2.710 |
0.000 |
| y |
-2.710 |
0.885 |
0.000 |
| z |
0.000 |
0.000 |
-0.736 |
|
| Polar |
|---|
| 3z2-r2 | -1.472 |
|---|
| x2-y2 | -0.690 |
|---|
| xy | -2.710 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.620 |
1.744 |
0.000 |
| y |
1.744 |
7.152 |
0.000 |
| z |
0.000 |
0.000 |
4.101 |
<r2> (average value of r
2) Å
2
| <r2> |
82.913 |
| (<r2>)1/2 |
9.106 |
Jump to
S1C1
Energy calculated at HSEh1PBE/6-311+G(3df,2p)
| | hartrees |
|---|
| Energy at 0K | -191.766647 |
| Energy at 298.15K | |
| HF Energy | -191.766647 |
| Nuclear repulsion energy | 105.046558 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-311+G(3df,2p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3252 |
3252 |
0.97 |
53.95 |
0.61 |
0.76 |
| 2 |
A' |
3178 |
3178 |
8.83 |
127.51 |
0.22 |
0.36 |
| 3 |
A' |
3155 |
3155 |
4.51 |
59.65 |
0.12 |
0.21 |
| 4 |
A' |
2917 |
2917 |
130.05 |
167.86 |
0.30 |
0.46 |
| 5 |
A' |
1815 |
1815 |
132.96 |
21.76 |
0.59 |
0.74 |
| 6 |
A' |
1684 |
1684 |
67.47 |
68.69 |
0.13 |
0.22 |
| 7 |
A' |
1434 |
1434 |
41.09 |
5.77 |
0.75 |
0.86 |
| 8 |
A' |
1424 |
1424 |
7.12 |
9.31 |
0.12 |
0.21 |
| 9 |
A' |
1313 |
1313 |
2.45 |
20.14 |
0.27 |
0.42 |
| 10 |
A' |
1069 |
1069 |
4.35 |
3.50 |
0.69 |
0.81 |
| 11 |
A' |
942 |
942 |
62.94 |
6.30 |
0.08 |
0.15 |
| 12 |
A' |
685 |
685 |
12.49 |
0.64 |
0.75 |
0.86 |
| 13 |
A' |
283 |
283 |
7.01 |
3.46 |
0.32 |
0.49 |
| 14 |
A" |
1046 |
1046 |
1.24 |
4.63 |
0.75 |
0.86 |
| 15 |
A" |
1030 |
1030 |
40.24 |
0.29 |
0.75 |
0.86 |
| 16 |
A" |
1022 |
1022 |
9.26 |
0.92 |
0.75 |
0.86 |
| 17 |
A" |
558 |
558 |
10.15 |
0.78 |
0.75 |
0.86 |
| 18 |
A" |
144 |
144 |
6.37 |
0.20 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 13475.0 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 13475.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-311+G(3df,2p)
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.882 |
-0.293 |
0.000 |
| C2 |
0.000 |
0.892 |
0.000 |
| C3 |
1.324 |
0.775 |
0.000 |
| O4 |
-0.494 |
-1.433 |
0.000 |
| H5 |
-1.969 |
-0.074 |
0.000 |
| H6 |
-0.483 |
1.864 |
0.000 |
| H7 |
1.975 |
1.639 |
0.000 |
| H8 |
1.781 |
-0.208 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.4779 | 2.4509 | 1.2039 | 1.1088 | 2.1938 | 3.4494 | 2.6645 |
C2 | 1.4779 | | 1.3288 | 2.3769 | 2.1933 | 1.0850 | 2.1114 | 2.0932 | C3 | 2.4509 | 1.3288 | | 2.8595 | 3.4003 | 2.1094 | 1.0823 | 1.0840 | O4 | 1.2039 | 2.3769 | 2.8595 | | 2.0060 | 3.2966 | 3.9410 | 2.5833 | H5 | 1.1088 | 2.1933 | 3.4003 | 2.0060 | | 2.4418 | 4.2999 | 3.7523 | H6 | 2.1938 | 1.0850 | 2.1094 | 3.2966 | 2.4418 | | 2.4680 | 3.0687 | H7 | 3.4494 | 2.1114 | 1.0823 | 3.9410 | 4.2999 | 2.4680 | | 1.8575 | H8 | 2.6645 | 2.0932 | 1.0840 | 2.5833 | 3.7523 | 3.0687 | 1.8575 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
121.588 |
|
C1 |
C2 |
H6 |
116.907 |
| C2 |
C1 |
O4 |
124.512 |
|
C2 |
C1 |
H5 |
115.233 |
| C2 |
C3 |
H7 |
121.923 |
|
C2 |
C3 |
H8 |
120.014 |
| C3 |
C2 |
H6 |
121.505 |
|
O4 |
C1 |
H5 |
120.255 |
| H7 |
C3 |
H8 |
118.062 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-311+G(3df,2p)
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.340 |
|
|
|
| 2 |
C |
0.088 |
|
|
|
| 3 |
C |
-0.281 |
|
|
|
| 4 |
O |
-0.638 |
|
|
|
| 5 |
H |
0.100 |
|
|
|
| 6 |
H |
0.102 |
|
|
|
| 7 |
H |
0.138 |
|
|
|
| 8 |
H |
0.150 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.247 |
2.767 |
0.000 |
2.778 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-20.200 |
-0.642 |
0.000 |
| y |
-0.642 |
-26.447 |
0.000 |
| z |
0.000 |
0.000 |
-25.028 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.537 |
-0.642 |
0.000 |
| y |
-0.642 |
-3.832 |
0.000 |
| z |
0.000 |
0.000 |
-1.705 |
|
| Polar |
|---|
| 3z2-r2 | -3.410 |
|---|
| x2-y2 | 6.247 |
|---|
| xy | -0.642 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.725 |
0.612 |
0.000 |
| y |
0.612 |
6.525 |
0.000 |
| z |
0.000 |
0.000 |
4.080 |
<r2> (average value of r
2) Å
2
| <r2> |
74.766 |
| (<r2>)1/2 |
8.647 |
