Jump to
S1C2
Energy calculated at CCSD(T)=FULL/6-311+G(3df,2p)
| | hartrees |
|---|
| Energy at 0K | -191.686308 |
| Energy at 298.15K | |
| HF Energy | -190.831145 |
| Nuclear repulsion energy | 103.078718 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD(T)=FULL/6-311+G(3df,2p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3242 |
3242 |
|
|
|
|
| 2 |
A' |
3197 |
3197 |
|
|
|
|
| 3 |
A' |
3140 |
3140 |
|
|
|
|
| 4 |
A' |
2919 |
2919 |
|
|
|
|
| 5 |
A' |
1767 |
1767 |
|
|
|
|
| 6 |
A' |
1660 |
1660 |
|
|
|
|
| 7 |
A' |
1448 |
1448 |
|
|
|
|
| 8 |
A' |
1398 |
1398 |
|
|
|
|
| 9 |
A' |
1291 |
1291 |
|
|
|
|
| 10 |
A' |
1175 |
1175 |
|
|
|
|
| 11 |
A' |
923 |
923 |
|
|
|
|
| 12 |
A' |
566 |
566 |
|
|
|
|
| 13 |
A' |
288 |
288 |
|
|
|
|
| 14 |
A" |
1026 |
1026 |
|
|
|
|
| 15 |
A" |
1016 |
1016 |
|
|
|
|
| 16 |
A" |
977 |
977 |
|
|
|
|
| 17 |
A" |
609 |
609 |
|
|
|
|
| 18 |
A" |
164 |
164 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13402.0 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 13402.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCSD(T)=FULL/6-311+G(3df,2p)
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.153 |
-0.744 |
0.000 |
| C2 |
0.000 |
0.722 |
0.000 |
| C3 |
1.219 |
1.276 |
0.000 |
| O4 |
-1.220 |
-1.316 |
0.000 |
| H5 |
0.798 |
-1.312 |
0.000 |
| H6 |
-0.910 |
1.311 |
0.000 |
| H7 |
1.365 |
2.348 |
0.000 |
| H8 |
2.108 |
0.655 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.4733 | 2.4417 | 1.2115 | 1.1073 | 2.1892 | 3.4438 | 2.6582 |
C2 | 1.4733 | | 1.3394 | 2.3753 | 2.1847 | 1.0836 | 2.1230 | 2.1091 | C3 | 2.4417 | 1.3394 | | 3.5597 | 2.6225 | 2.1292 | 1.0814 | 1.0844 | O4 | 1.2115 | 2.3753 | 3.5597 | | 2.0178 | 2.6449 | 4.4842 | 3.8681 | H5 | 1.1073 | 2.1847 | 2.6225 | 2.0178 | | 3.1293 | 3.7037 | 2.3635 | H6 | 2.1892 | 1.0836 | 2.1292 | 2.6449 | 3.1293 | | 2.5000 | 3.0881 | H7 | 3.4438 | 2.1230 | 1.0814 | 4.4842 | 3.7037 | 2.5000 | | 1.8488 | H8 | 2.6582 | 2.1091 | 1.0844 | 3.8681 | 2.3635 | 3.0881 | 1.8488 | |
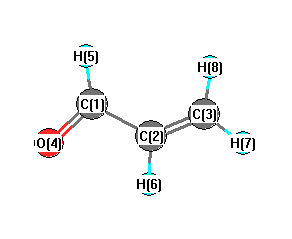 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
120.398 |
|
C1 |
C2 |
H6 |
116.964 |
| C2 |
C1 |
O4 |
124.149 |
|
C2 |
C1 |
H5 |
114.949 |
| C2 |
C3 |
H7 |
122.197 |
|
C2 |
C3 |
H8 |
120.586 |
| C3 |
C2 |
H6 |
122.638 |
|
O4 |
C1 |
H5 |
120.903 |
| H7 |
C3 |
H8 |
117.217 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CCSD(T)=FULL/6-311+G(3df,2p)
| | hartrees |
|---|
| Energy at 0K | -191.683004 |
| Energy at 298.15K | |
| HF Energy | -190.827155 |
| Nuclear repulsion energy | 104.627177 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD(T)=FULL/6-311+G(3df,2p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3258 |
3258 |
|
|
|
|
| 2 |
A' |
3185 |
3185 |
|
|
|
|
| 3 |
A' |
3158 |
3158 |
|
|
|
|
| 4 |
A' |
2957 |
2957 |
|
|
|
|
| 5 |
A' |
1767 |
1767 |
|
|
|
|
| 6 |
A' |
1661 |
1661 |
|
|
|
|
| 7 |
A' |
1440 |
1440 |
|
|
|
|
| 8 |
A' |
1430 |
1430 |
|
|
|
|
| 9 |
A' |
1307 |
1307 |
|
|
|
|
| 10 |
A' |
1067 |
1067 |
|
|
|
|
| 11 |
A' |
938 |
938 |
|
|
|
|
| 12 |
A' |
677 |
677 |
|
|
|
|
| 13 |
A' |
285 |
285 |
|
|
|
|
| 14 |
A" |
1023 |
1023 |
|
|
|
|
| 15 |
A" |
1011 |
1011 |
|
|
|
|
| 16 |
A" |
988 |
988 |
|
|
|
|
| 17 |
A" |
556 |
556 |
|
|
|
|
| 18 |
A" |
137 |
137 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13421.5 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 13421.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCSD(T)=FULL/6-311+G(3df,2p)
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.891 |
-0.289 |
0.000 |
| C2 |
0.000 |
0.898 |
0.000 |
| C3 |
1.333 |
0.769 |
0.000 |
| O4 |
-0.496 |
-1.435 |
0.000 |
| H5 |
-1.975 |
-0.076 |
0.000 |
| H6 |
-0.474 |
1.873 |
0.000 |
| H7 |
1.987 |
1.630 |
0.000 |
| H8 |
1.782 |
-0.217 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.4845 | 2.4627 | 1.2118 | 1.1046 | 2.2024 | 3.4595 | 2.6738 |
C2 | 1.4845 | | 1.3388 | 2.3847 | 2.2021 | 1.0844 | 2.1175 | 2.1017 | C3 | 2.4627 | 1.3388 | | 2.8633 | 3.4138 | 2.1173 | 1.0819 | 1.0832 | O4 | 1.2118 | 2.3847 | 2.8633 | | 2.0086 | 3.3080 | 3.9444 | 2.5826 | H5 | 1.1046 | 2.2021 | 3.4138 | 2.0086 | | 2.4604 | 4.3137 | 3.7593 | H6 | 2.2024 | 1.0844 | 2.1173 | 3.3080 | 2.4604 | | 2.4725 | 3.0750 | H7 | 3.4595 | 2.1175 | 1.0819 | 3.9444 | 4.3137 | 2.4725 | | 1.8587 | H8 | 2.6738 | 2.1017 | 1.0832 | 2.5826 | 3.7593 | 3.0750 | 1.8587 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
121.365 |
|
C1 |
C2 |
H6 |
117.196 |
| C2 |
C1 |
O4 |
124.055 |
|
C2 |
C1 |
H5 |
115.764 |
| C2 |
C3 |
H7 |
121.673 |
|
C2 |
C3 |
H8 |
120.024 |
| C3 |
C2 |
H6 |
121.440 |
|
O4 |
C1 |
H5 |
120.180 |
| H7 |
C3 |
H8 |
118.304 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
