Jump to
S1C2
Vibrational Frequencies calculated at HSEh1PBE/6-311+G(3df,2p)
Geometric Data calculated at HSEh1PBE/6-311+G(3df,2p)
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at HSEh1PBE/6-311+G(3df,2p)
| | hartrees |
|---|
| Energy at 0K | -3112.441438 |
| Energy at 298.15K | |
| HF Energy | -3112.441438 |
| Nuclear repulsion energy | 305.928542 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-311+G(3df,2p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3170 |
3170 |
0.56 |
37.95 |
0.75 |
0.86 |
| 2 |
A |
3144 |
3144 |
0.82 |
71.28 |
0.75 |
0.86 |
| 3 |
A |
3099 |
3099 |
10.00 |
163.64 |
0.03 |
0.06 |
| 4 |
A |
3077 |
3077 |
8.64 |
111.50 |
0.15 |
0.26 |
| 5 |
A |
1465 |
1465 |
5.77 |
4.76 |
0.75 |
0.86 |
| 6 |
A |
1459 |
1459 |
10.49 |
6.33 |
0.73 |
0.84 |
| 7 |
A |
1329 |
1329 |
16.40 |
1.50 |
0.34 |
0.51 |
| 8 |
A |
1289 |
1289 |
43.45 |
3.91 |
0.31 |
0.48 |
| 9 |
A |
1212 |
1212 |
1.59 |
5.19 |
0.71 |
0.83 |
| 10 |
A |
1146 |
1146 |
0.74 |
1.29 |
0.67 |
0.81 |
| 11 |
A |
1053 |
1053 |
2.36 |
1.11 |
0.70 |
0.82 |
| 12 |
A |
936 |
936 |
8.52 |
3.80 |
0.16 |
0.28 |
| 13 |
A |
874 |
874 |
20.62 |
1.26 |
0.75 |
0.86 |
| 14 |
A |
694 |
694 |
18.02 |
7.76 |
0.26 |
0.42 |
| 15 |
A |
590 |
590 |
10.47 |
16.87 |
0.13 |
0.23 |
| 16 |
A |
389 |
389 |
6.86 |
1.54 |
0.68 |
0.81 |
| 17 |
A |
243 |
243 |
1.30 |
1.20 |
0.21 |
0.35 |
| 18 |
A |
98 |
98 |
0.50 |
0.85 |
0.65 |
0.79 |
Unscaled Zero Point Vibrational Energy (zpe) 12633.2 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 12633.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-311+G(3df,2p)
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.012 |
1.097 |
-0.380 |
| C2 |
1.281 |
0.892 |
0.400 |
| Br3 |
-1.351 |
-0.223 |
0.036 |
| Cl4 |
2.160 |
-0.574 |
-0.086 |
| H5 |
-0.421 |
2.064 |
-0.131 |
| H6 |
0.183 |
1.037 |
-1.452 |
| H7 |
1.950 |
1.734 |
0.219 |
| H8 |
1.086 |
0.814 |
1.467 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Br3 |
Cl4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5029 | 1.9424 | 2.7364 | 1.0887 | 1.0867 | 2.1258 | 2.1552 |
C2 | 1.5029 | | 2.8809 | 1.7769 | 2.1334 | 2.1573 | 1.0908 | 1.0879 | Br3 | 1.9424 | 2.8809 | | 3.5296 | 2.4741 | 2.4804 | 3.8416 | 3.0103 | Cl4 | 2.7364 | 1.7769 | 3.5296 | | 3.6906 | 2.8926 | 2.3374 | 2.3437 | H5 | 1.0887 | 2.1334 | 2.4741 | 3.6906 | | 1.7788 | 2.4196 | 2.5271 | H6 | 1.0867 | 2.1573 | 2.4804 | 2.8926 | 1.7788 | | 2.5296 | 3.0635 | H7 | 2.1258 | 1.0908 | 3.8416 | 2.3374 | 2.4196 | 2.5296 | | 1.7751 | H8 | 2.1552 | 1.0879 | 3.0103 | 2.3437 | 2.5271 | 3.0635 | 1.7751 | |
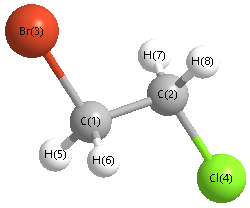 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Cl4 |
112.827 |
|
C1 |
C2 |
H7 |
109.060 |
| C1 |
C2 |
H8 |
111.580 |
|
C2 |
C1 |
Br3 |
112.853 |
| C2 |
C1 |
Cl4 |
36.762 |
|
C2 |
C1 |
H6 |
111.823 |
| Br3 |
C1 |
H5 |
105.960 |
|
Br3 |
C1 |
H6 |
106.491 |
| Cl4 |
C2 |
H7 |
106.739 |
|
Cl4 |
C2 |
H8 |
107.340 |
| H5 |
C1 |
H6 |
109.709 |
|
H7 |
C2 |
H8 |
109.127 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-311+G(3df,2p)
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.081 |
|
|
|
| 2 |
C |
-0.232 |
|
|
|
| 3 |
Br |
-0.117 |
|
|
|
| 4 |
Cl |
-0.247 |
|
|
|
| 5 |
H |
0.160 |
|
|
|
| 6 |
H |
0.172 |
|
|
|
| 7 |
H |
0.166 |
|
|
|
| 8 |
H |
0.179 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.393 |
2.615 |
0.092 |
2.646 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-47.756 |
2.595 |
0.479 |
| y |
2.595 |
-40.665 |
-0.096 |
| z |
0.479 |
-0.096 |
-43.355 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-5.746 |
2.595 |
0.479 |
| y |
2.595 |
4.891 |
-0.096 |
| z |
0.479 |
-0.096 |
0.855 |
|
| Polar |
|---|
| 3z2-r2 | 1.710 |
|---|
| x2-y2 | -7.091 |
|---|
| xy | 2.595 |
|---|
| xz | 0.479 |
|---|
| yz | -0.096 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.538 |
0.183 |
-0.156 |
| y |
0.183 |
8.962 |
0.032 |
| z |
-0.156 |
0.032 |
7.568 |
<r2> (average value of r
2) Å
2
| <r2> |
220.248 |
| (<r2>)1/2 |
14.841 |
