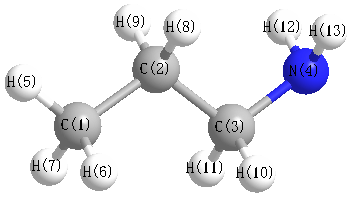
Vibrational Frequencies calculated at PBEPBE/6-311+G(3df,2p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3405 |
3377 |
2.84 |
|
|
|
| 2 |
A' |
3031 |
3006 |
30.77 |
|
|
|
| 3 |
A' |
2957 |
2933 |
42.41 |
|
|
|
| 4 |
A' |
2944 |
2919 |
55.98 |
|
|
|
| 5 |
A' |
2933 |
2909 |
3.36 |
|
|
|
| 6 |
A' |
1605 |
1592 |
21.58 |
|
|
|
| 7 |
A' |
1457 |
1445 |
6.68 |
|
|
|
| 8 |
A' |
1442 |
1430 |
0.75 |
|
|
|
| 9 |
A' |
1430 |
1418 |
0.01 |
|
|
|
| 10 |
A' |
1356 |
1345 |
5.18 |
|
|
|
| 11 |
A' |
1339 |
1328 |
3.80 |
|
|
|
| 12 |
A' |
1276 |
1265 |
4.10 |
|
|
|
| 13 |
A' |
1102 |
1093 |
2.23 |
|
|
|
| 14 |
A' |
1064 |
1055 |
14.73 |
|
|
|
| 15 |
A' |
1011 |
1003 |
0.15 |
|
|
|
| 16 |
A' |
877 |
870 |
63.89 |
|
|
|
| 17 |
A' |
793 |
787 |
117.21 |
|
|
|
| 18 |
A' |
431 |
428 |
3.61 |
|
|
|
| 19 |
A' |
253 |
251 |
4.15 |
|
|
|
| 20 |
A" |
3483 |
3455 |
0.28 |
|
|
|
| 21 |
A" |
3021 |
2997 |
50.39 |
|
|
|
| 22 |
A" |
2986 |
2962 |
36.41 |
|
|
|
| 23 |
A" |
2958 |
2934 |
0.34 |
|
|
|
| 24 |
A" |
1448 |
1436 |
8.17 |
|
|
|
| 25 |
A" |
1341 |
1330 |
0.24 |
|
|
|
| 26 |
A" |
1277 |
1266 |
0.59 |
|
|
|
| 27 |
A" |
1196 |
1186 |
0.01 |
|
|
|
| 28 |
A" |
999 |
991 |
0.00 |
|
|
|
| 29 |
A" |
829 |
822 |
0.89 |
|
|
|
| 30 |
A" |
722 |
716 |
2.63 |
|
|
|
| 31 |
A" |
266 |
264 |
33.80 |
|
|
|
| 32 |
A" |
213 |
211 |
8.97 |
|
|
|
| 33 |
A" |
126 |
125 |
0.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25785.5 cm
-1
Scaled (by 0.9918) Zero Point Vibrational Energy (zpe) 25574.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/6-311+G(3df,2p)
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.444 |
|
|
|
| 2 |
C |
-0.035 |
|
|
|
| 3 |
C |
0.082 |
|
|
|
| 4 |
N |
-0.769 |
|
|
|
| 5 |
H |
0.112 |
|
|
|
| 6 |
H |
0.126 |
|
|
|
| 7 |
H |
0.126 |
|
|
|
| 8 |
H |
0.113 |
|
|
|
| 9 |
H |
0.113 |
|
|
|
| 10 |
H |
0.111 |
|
|
|
| 11 |
H |
0.111 |
|
|
|
| 12 |
H |
0.177 |
|
|
|
| 13 |
H |
0.177 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.118 |
1.352 |
0.000 |
1.357 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-26.872 |
-1.833 |
0.000 |
| y |
-1.833 |
-33.148 |
0.000 |
| z |
0.000 |
0.000 |
-26.136 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.769 |
-1.833 |
0.000 |
| y |
-1.833 |
-6.644 |
0.000 |
| z |
0.000 |
0.000 |
3.874 |
|
| Polar |
|---|
| 3z2-r2 | 7.749 |
|---|
| x2-y2 | 6.275 |
|---|
| xy | -1.833 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.318 |
0.810 |
0.000 |
| y |
0.810 |
8.145 |
0.000 |
| z |
0.000 |
0.000 |
6.833 |
<r2> (average value of r
2) Å
2
| <r2> |
113.956 |
| (<r2>)1/2 |
10.675 |