Jump to
S1C2
S1C3
Energy calculated at CCSD(T)=FULL/6-311+G(3df,2p)
| | hartrees |
|---|
| Energy at 0K | -692.295565 |
| Energy at 298.15K | -692.294977 |
| HF Energy | -691.510655 |
| Nuclear repulsion energy | 60.674315 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at CCSD(T)=FULL/6-311+G(3df,2p)
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| K1 |
0.000 |
0.000 |
1.316 |
| C2 |
0.000 |
0.000 |
-1.292 |
| N3 |
0.000 |
0.000 |
-2.465 |
Atom - Atom Distances (Å)
| |
K1 |
C2 |
N3 |
| K1 | | 2.6078 | 3.7817 |
C2 | 2.6078 | | 1.1739 | N3 | 3.7817 | 1.1739 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| K1 |
C2 |
N3 |
180.000 |
|
K1 |
N3 |
C2 |
0.000 |
| C2 |
K1 |
N3 |
0.000 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Energy calculated at CCSD(T)=FULL/6-311+G(3df,2p)
| | hartrees |
|---|
| Energy at 0K | -692.297826 |
| Energy at 298.15K | |
| HF Energy | -691.518107 |
| Nuclear repulsion energy | 64.162038 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at CCSD(T)=FULL/6-311+G(3df,2p)
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| K1 |
0.000 |
0.000 |
1.217 |
| C2 |
0.000 |
0.000 |
-2.414 |
| N3 |
0.000 |
0.000 |
-1.234 |
Atom - Atom Distances (Å)
| |
K1 |
C2 |
N3 |
| K1 | | 3.6310 | 2.4503 |
C2 | 3.6310 | | 1.1807 | N3 | 2.4503 | 1.1807 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| K1 |
C2 |
N3 |
0.000 |
|
K1 |
N3 |
C2 |
180.000 |
| C2 |
K1 |
N3 |
0.000 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at CCSD(T)=FULL/6-311+G(3df,2p)
| | hartrees |
|---|
| Energy at 0K | -692.301927 |
| Energy at 298.15K | -692.301791 |
| HF Energy | -691.518646 |
| Nuclear repulsion energy | 68.365631 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at CCSD(T)=FULL/6-311+G(3df,2p)
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| K1 |
0.000 |
1.047 |
0.000 |
| C2 |
0.629 |
-1.620 |
0.000 |
| N3 |
-0.539 |
-1.452 |
0.000 |
Atom - Atom Distances (Å)
| |
K1 |
C2 |
N3 |
| K1 | | 2.7403 | 2.5561 |
C2 | 2.7403 | | 1.1809 | N3 | 2.5561 | 1.1809 | |
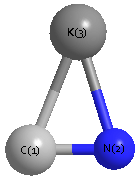 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| K1 |
C2 |
N3 |
68.520 |
|
K1 |
N3 |
C2 |
86.018 |
| C2 |
K1 |
N3 |
25.462 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
