Jump to
S1C2
S1C3
Energy calculated at CCSD(T)/6-311+G(3df,2p)
| | hartrees |
|---|
| Energy at 0K | -1590.887856 |
| Energy at 298.15K | |
| HF Energy | -1590.127480 |
| Nuclear repulsion energy | 349.836796 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD(T)/6-311+G(3df,2p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
513 |
506 |
|
|
|
|
| 2 |
A1 |
208 |
205 |
|
|
|
|
| 3 |
B1 |
454 |
448 |
|
|
|
|
| 4 |
B2 |
308 |
304 |
|
|
|
|
| 5 |
E |
427 |
422 |
|
|
|
|
| 5 |
E |
427 |
422 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 1168.8 cm
-1
Scaled (by 0.9869) Zero Point Vibrational Energy (zpe) 1153.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCSD(T)/6-311+G(3df,2p)
Point Group is D2d
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.000 |
1.432 |
0.319 |
| S2 |
0.000 |
-1.432 |
0.319 |
| S3 |
-1.432 |
0.000 |
-0.319 |
| S4 |
1.432 |
0.000 |
-0.319 |
Atom - Atom Distances (Å)
| |
S1 |
S2 |
S3 |
S4 |
| S1 | | 2.8639 | 2.1231 | 2.1231 |
S2 | 2.8639 | | 2.1231 | 2.1231 | S3 | 2.1231 | 2.1231 | | 2.8639 | S4 | 2.1231 | 2.1231 | 2.8639 | |
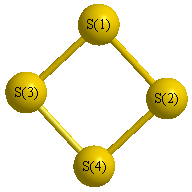 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
S2 |
S4 |
47.589 |
|
S1 |
S3 |
S4 |
47.589 |
| S2 |
S1 |
S3 |
47.589 |
|
S2 |
S4 |
S3 |
47.589 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Energy calculated at CCSD(T)/6-311+G(3df,2p)
| | hartrees |
|---|
| Energy at 0K | -1590.915165 |
| Energy at 298.15K | |
| HF Energy | -1590.099285 |
| Nuclear repulsion energy | 331.397941 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD(T)/6-311+G(3df,2p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
683 |
675 |
|
|
|
|
| 2 |
A1 |
329 |
325 |
|
|
|
|
| 3 |
A1 |
116 |
115 |
|
|
|
|
| 4 |
A2 |
207 |
204 |
|
|
|
|
| 5 |
B2 |
643 |
635 |
|
|
|
|
| 6 |
B2 |
325 |
321 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 1151.9 cm
-1
Scaled (by 0.9869) Zero Point Vibrational Energy (zpe) 1136.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCSD(T)/6-311+G(3df,2p)
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.000 |
1.077 |
0.924 |
| S2 |
0.000 |
-1.077 |
0.924 |
| S3 |
0.000 |
1.573 |
-0.924 |
| S4 |
0.000 |
-1.573 |
-0.924 |
Atom - Atom Distances (Å)
| |
S1 |
S2 |
S3 |
S4 |
| S1 | | 2.1533 | 1.9139 | 3.2305 |
S2 | 2.1533 | | 3.2305 | 1.9139 | S3 | 1.9139 | 3.2305 | | 3.1454 | S4 | 3.2305 | 1.9139 | 3.1454 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
S2 |
S4 |
105.022 |
|
S1 |
S3 |
S4 |
74.978 |
| S2 |
S1 |
S3 |
105.022 |
|
S2 |
S4 |
S3 |
74.978 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at CCSD(T)/6-311+G(3df,2p)
| | hartrees |
|---|
| Energy at 0K | -1590.911938 |
| Energy at 298.15K | |
| HF Energy | -1590.067081 |
| Nuclear repulsion energy | 332.603107 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD(T)/6-311+G(3df,2p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
655 |
647 |
|
|
|
|
| 2 |
Ag |
258 |
254 |
|
|
|
|
| 3 |
Au |
229 |
226 |
|
|
|
|
| 4 |
B1u |
689i |
680i |
|
|
|
|
| 5 |
B2u |
110i |
108i |
|
|
|
|
| 6 |
B3g |
172 |
170 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 257.5 cm
-1
Scaled (by 0.9869) Zero Point Vibrational Energy (zpe) 254.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCSD(T)/6-311+G(3df,2p)
Point Group is D2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.000 |
0.950 |
1.286 |
| S2 |
0.000 |
0.950 |
-1.286 |
| S3 |
0.000 |
-0.950 |
1.286 |
| S4 |
0.000 |
-0.950 |
-1.286 |
Atom - Atom Distances (Å)
| |
S1 |
S2 |
S3 |
S4 |
| S1 | | 2.5725 | 1.9005 | 3.1984 |
S2 | 2.5725 | | 3.1984 | 1.9005 | S3 | 1.9005 | 3.1984 | | 2.5725 | S4 | 3.1984 | 1.9005 | 2.5725 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
S2 |
S4 |
90.000 |
|
S1 |
S3 |
S4 |
90.000 |
| S2 |
S1 |
S3 |
90.000 |
|
S2 |
S4 |
S3 |
90.000 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
