Jump to
S1C2
S1C3
Energy calculated at MP2/6-311+G(3df,2p)
| | hartrees |
|---|
| Energy at 0K | -1590.797799 |
| Energy at 298.15K | |
| HF Energy | -1590.128165 |
| Nuclear repulsion energy | 353.339045 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-311+G(3df,2p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
535 |
505 |
0.00 |
35.63 |
0.04 |
0.08 |
| 2 |
A1 |
218 |
206 |
0.00 |
1.00 |
0.67 |
0.80 |
| 3 |
B1 |
477 |
450 |
0.00 |
13.40 |
0.75 |
0.86 |
| 4 |
B2 |
308 |
290 |
0.02 |
5.41 |
0.75 |
0.86 |
| 5 |
E |
446 |
421 |
0.04 |
2.95 |
0.75 |
0.86 |
| 5 |
E |
446 |
421 |
0.04 |
2.95 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 1215.0 cm
-1
Scaled (by 0.9434) Zero Point Vibrational Energy (zpe) 1146.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
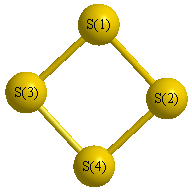
Geometric Data calculated at MP2/6-311+G(3df,2p)
Point Group is D2d
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.000 |
1.415 |
0.325 |
| S2 |
0.000 |
-1.415 |
0.325 |
| S3 |
-1.415 |
0.000 |
-0.325 |
| S4 |
1.415 |
0.000 |
-0.325 |
Atom - Atom Distances (Å)
| |
S1 |
S2 |
S3 |
S4 |
| S1 | | 2.8296 | 2.1037 | 2.1037 |
S2 | 2.8296 | | 2.1037 | 2.1037 | S3 | 2.1037 | 2.1037 | | 2.8296 | S4 | 2.1037 | 2.1037 | 2.8296 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
S2 |
S4 |
47.737 |
|
S1 |
S3 |
S4 |
47.737 |
| S2 |
S1 |
S3 |
47.737 |
|
S2 |
S4 |
S3 |
47.737 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Energy calculated at MP2/6-311+G(3df,2p)
| | hartrees |
|---|
| Energy at 0K | -1590.869108 |
| Energy at 298.15K | |
| HF Energy | -1590.062051 |
| Nuclear repulsion energy | 328.209399 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-311+G(3df,2p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
663 |
626 |
0.00 |
3568601.00 |
0.34 |
0.50 |
| 2 |
A1 |
423 |
399 |
0.25 |
1900.59 |
0.33 |
0.50 |
| 3 |
A1 |
217 |
205 |
0.00 |
0.91 |
0.33 |
0.50 |
| 4 |
A2 |
226 |
214 |
0.00 |
754.85 |
0.75 |
0.86 |
| 5 |
B2 |
2765 |
2609 |
1640564.00 |
8.64 |
0.75 |
0.86 |
| 6 |
B2 |
309 |
292 |
0.00 |
0.00 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 2302.1 cm
-1
Scaled (by 0.9434) Zero Point Vibrational Energy (zpe) 2171.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/6-311+G(3df,2p)
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.000 |
1.307 |
0.960 |
| S2 |
0.000 |
-1.307 |
0.960 |
| S3 |
0.000 |
1.308 |
-0.960 |
| S4 |
0.000 |
-1.308 |
-0.960 |
Atom - Atom Distances (Å)
| |
S1 |
S2 |
S3 |
S4 |
| S1 | | 2.6150 | 1.9203 | 3.2444 |
S2 | 2.6150 | | 3.2444 | 1.9203 | S3 | 1.9203 | 3.2444 | | 2.6152 | S4 | 3.2444 | 1.9203 | 2.6152 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
S2 |
S4 |
90.003 |
|
S1 |
S3 |
S4 |
89.997 |
| S2 |
S1 |
S3 |
90.003 |
|
S2 |
S4 |
S3 |
89.997 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at MP2/6-311+G(3df,2p)
| | hartrees |
|---|
| Energy at 0K | -1590.869108 |
| Energy at 298.15K | |
| HF Energy | -1590.062118 |
| Nuclear repulsion energy | 328.280207 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-311+G(3df,2p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
663 |
626 |
0.00 |
5025448.00 |
0.33 |
0.50 |
| 2 |
Ag |
218 |
206 |
0.00 |
0.00 |
0.33 |
0.50 |
| 3 |
Au |
227 |
214 |
0.00 |
947.56 |
0.75 |
0.86 |
| 4 |
B1u |
2895 |
2731 |
1989103.00 |
0.00 |
0.75 |
0.86 |
| 5 |
B2u |
423 |
399 |
0.25 |
0.00 |
0.75 |
0.86 |
| 6 |
B3g |
309 |
292 |
0.00 |
0.00 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 2367.7 cm
-1
Scaled (by 0.9434) Zero Point Vibrational Energy (zpe) 2233.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/6-311+G(3df,2p)
Point Group is D2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.000 |
0.960 |
1.307 |
| S2 |
0.000 |
0.960 |
-1.307 |
| S3 |
0.000 |
-0.960 |
1.307 |
| S4 |
0.000 |
-0.960 |
-1.307 |
Atom - Atom Distances (Å)
| |
S1 |
S2 |
S3 |
S4 |
| S1 | | 2.6139 | 1.9203 | 3.2435 |
S2 | 2.6139 | | 3.2435 | 1.9203 | S3 | 1.9203 | 3.2435 | | 2.6139 | S4 | 3.2435 | 1.9203 | 2.6139 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
S2 |
S4 |
90.000 |
|
S1 |
S3 |
S4 |
90.000 |
| S2 |
S1 |
S3 |
90.000 |
|
S2 |
S4 |
S3 |
90.000 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
