Jump to
S1C2
Energy calculated at HSEh1PBE/6-311+G(3df,2p)
| | hartrees |
|---|
| Energy at 0K | -218.224573 |
| Energy at 298.15K | -218.232399 |
| HF Energy | -218.224573 |
| Nuclear repulsion energy | 128.469612 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-311+G(3df,2p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3125 |
3125 |
24.04 |
|
|
|
| 2 |
A' |
3052 |
3052 |
42.03 |
|
|
|
| 3 |
A' |
3042 |
3042 |
20.30 |
|
|
|
| 4 |
A' |
3034 |
3034 |
18.87 |
|
|
|
| 5 |
A' |
1514 |
1514 |
7.26 |
|
|
|
| 6 |
A' |
1502 |
1502 |
3.36 |
|
|
|
| 7 |
A' |
1489 |
1489 |
0.62 |
|
|
|
| 8 |
A' |
1422 |
1422 |
14.15 |
|
|
|
| 9 |
A' |
1410 |
1410 |
0.70 |
|
|
|
| 10 |
A' |
1328 |
1328 |
0.78 |
|
|
|
| 11 |
A' |
1144 |
1144 |
1.45 |
|
|
|
| 12 |
A' |
1079 |
1079 |
106.44 |
|
|
|
| 13 |
A' |
1051 |
1051 |
20.38 |
|
|
|
| 14 |
A' |
907 |
907 |
8.15 |
|
|
|
| 15 |
A' |
454 |
454 |
6.54 |
|
|
|
| 16 |
A' |
267 |
267 |
3.19 |
|
|
|
| 17 |
A" |
3116 |
3116 |
62.81 |
|
|
|
| 18 |
A" |
3094 |
3094 |
5.98 |
|
|
|
| 19 |
A" |
3072 |
3072 |
6.71 |
|
|
|
| 20 |
A" |
1497 |
1497 |
8.12 |
|
|
|
| 21 |
A" |
1315 |
1315 |
0.03 |
|
|
|
| 22 |
A" |
1269 |
1269 |
0.58 |
|
|
|
| 23 |
A" |
1190 |
1190 |
0.92 |
|
|
|
| 24 |
A" |
892 |
892 |
1.19 |
|
|
|
| 25 |
A" |
761 |
761 |
2.40 |
|
|
|
| 26 |
A" |
231 |
231 |
0.00 |
|
|
|
| 27 |
A" |
125 |
125 |
3.21 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21189.9 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 21189.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-311+G(3df,2p)
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.127 |
-0.774 |
0.000 |
| C2 |
0.000 |
0.728 |
0.000 |
| C3 |
-1.458 |
1.162 |
0.000 |
| F4 |
1.460 |
-1.150 |
0.000 |
| H5 |
-0.338 |
-1.212 |
0.888 |
| H6 |
-0.338 |
-1.212 |
-0.888 |
| H7 |
0.516 |
1.125 |
-0.878 |
| H8 |
0.516 |
1.125 |
0.878 |
| H9 |
-1.544 |
2.249 |
0.000 |
| H10 |
-1.984 |
0.790 |
-0.882 |
| H11 |
-1.984 |
0.790 |
0.882 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
F4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| C1 | | 1.5076 | 2.5017 | 1.3848 | 1.0943 | 1.0943 | 2.1281 | 2.1281 | 3.4535 | 2.7714 | 2.7714 |
C2 | 1.5076 | | 1.5207 | 2.3789 | 2.1608 | 2.1608 | 1.0930 | 1.0930 | 2.1665 | 2.1723 | 2.1723 | C3 | 2.5017 | 1.5207 | | 3.7224 | 2.7711 | 2.7711 | 2.1605 | 2.1605 | 1.0902 | 1.0924 | 1.0924 | F4 | 1.3848 | 2.3789 | 3.7224 | | 2.0062 | 2.0062 | 2.6149 | 2.6149 | 4.5355 | 4.0500 | 4.0500 | H5 | 1.0943 | 2.1608 | 2.7711 | 2.0062 | | 1.7768 | 3.0517 | 2.4885 | 3.7710 | 3.1388 | 2.5921 | H6 | 1.0943 | 2.1608 | 2.7711 | 2.0062 | 1.7768 | | 2.4885 | 3.0517 | 3.7710 | 2.5921 | 3.1388 | H7 | 2.1281 | 1.0930 | 2.1605 | 2.6149 | 3.0517 | 2.4885 | | 1.7561 | 2.5051 | 2.5229 | 3.0760 | H8 | 2.1281 | 1.0930 | 2.1605 | 2.6149 | 2.4885 | 3.0517 | 1.7561 | | 2.5051 | 3.0760 | 2.5229 | H9 | 3.4535 | 2.1665 | 1.0902 | 4.5355 | 3.7710 | 3.7710 | 2.5051 | 2.5051 | | 1.7607 | 1.7607 | H10 | 2.7714 | 2.1723 | 1.0924 | 4.0500 | 3.1388 | 2.5921 | 2.5229 | 3.0760 | 1.7607 | | 1.7635 | H11 | 2.7714 | 2.1723 | 1.0924 | 4.0500 | 2.5921 | 3.1388 | 3.0760 | 2.5229 | 1.7607 | 1.7635 | |
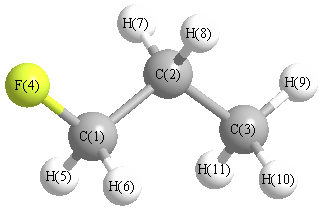 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
111.397 |
|
C1 |
C2 |
H7 |
108.777 |
| C1 |
C2 |
H8 |
108.777 |
|
C2 |
C1 |
F4 |
110.593 |
| C2 |
C1 |
H5 |
111.301 |
|
C2 |
C1 |
H6 |
111.301 |
| C2 |
C3 |
H9 |
111.082 |
|
C2 |
C3 |
H10 |
111.413 |
| C2 |
C3 |
H11 |
111.413 |
|
C3 |
C2 |
H7 |
110.432 |
| C3 |
C2 |
H8 |
110.432 |
|
F4 |
C1 |
H5 |
107.466 |
| F4 |
C1 |
H6 |
107.466 |
|
H5 |
C1 |
H6 |
108.551 |
| H7 |
C2 |
H8 |
106.895 |
|
H9 |
C3 |
H10 |
107.551 |
| H9 |
C3 |
H11 |
107.551 |
|
H10 |
C3 |
H11 |
107.635 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-311+G(3df,2p)
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.168 |
|
|
|
| 2 |
C |
-0.207 |
|
|
|
| 3 |
C |
-0.417 |
|
|
|
| 4 |
F |
-0.460 |
|
|
|
| 5 |
H |
0.129 |
|
|
|
| 6 |
H |
0.129 |
|
|
|
| 7 |
H |
0.127 |
|
|
|
| 8 |
H |
0.127 |
|
|
|
| 9 |
H |
0.125 |
|
|
|
| 10 |
H |
0.139 |
|
|
|
| 11 |
H |
0.139 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.805 |
0.850 |
0.000 |
1.995 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-28.675 |
2.254 |
0.000 |
| y |
2.254 |
-25.972 |
0.000 |
| z |
0.000 |
0.000 |
-24.709 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.334 |
2.254 |
0.000 |
| y |
2.254 |
0.721 |
0.000 |
| z |
0.000 |
0.000 |
2.614 |
|
| Polar |
|---|
| 3z2-r2 | 5.228 |
|---|
| x2-y2 | -2.703 |
|---|
| xy | 2.254 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.305 |
-0.416 |
0.000 |
| y |
-0.416 |
6.124 |
0.000 |
| z |
0.000 |
0.000 |
5.435 |
<r2> (average value of r
2) Å
2
| <r2> |
102.794 |
| (<r2>)1/2 |
10.139 |
Jump to
S1C1
Energy calculated at HSEh1PBE/6-311+G(3df,2p)
| | hartrees |
|---|
| Energy at 0K | -218.224811 |
| Energy at 298.15K | |
| HF Energy | -218.224811 |
| Nuclear repulsion energy | 130.720730 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-311+G(3df,2p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3130 |
3130 |
20.29 |
|
|
|
| 2 |
A |
3116 |
3116 |
39.91 |
|
|
|
| 3 |
A |
3094 |
3094 |
38.82 |
|
|
|
| 4 |
A |
3077 |
3077 |
15.53 |
|
|
|
| 5 |
A |
3044 |
3044 |
19.99 |
|
|
|
| 6 |
A |
3041 |
3041 |
31.58 |
|
|
|
| 7 |
A |
3036 |
3036 |
18.97 |
|
|
|
| 8 |
A |
1507 |
1507 |
2.96 |
|
|
|
| 9 |
A |
1502 |
1502 |
9.73 |
|
|
|
| 10 |
A |
1490 |
1490 |
7.02 |
|
|
|
| 11 |
A |
1469 |
1469 |
3.66 |
|
|
|
| 12 |
A |
1420 |
1420 |
9.00 |
|
|
|
| 13 |
A |
1406 |
1406 |
4.76 |
|
|
|
| 14 |
A |
1376 |
1376 |
0.08 |
|
|
|
| 15 |
A |
1301 |
1301 |
1.82 |
|
|
|
| 16 |
A |
1275 |
1275 |
1.50 |
|
|
|
| 17 |
A |
1175 |
1175 |
1.10 |
|
|
|
| 18 |
A |
1127 |
1127 |
12.23 |
|
|
|
| 19 |
A |
1098 |
1098 |
54.60 |
|
|
|
| 20 |
A |
986 |
986 |
50.37 |
|
|
|
| 21 |
A |
920 |
920 |
2.93 |
|
|
|
| 22 |
A |
886 |
886 |
4.05 |
|
|
|
| 23 |
A |
766 |
766 |
0.76 |
|
|
|
| 24 |
A |
479 |
479 |
4.02 |
|
|
|
| 25 |
A |
315 |
315 |
1.22 |
|
|
|
| 26 |
A |
213 |
213 |
1.46 |
|
|
|
| 27 |
A |
142 |
142 |
2.27 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21194.5 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 21194.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-311+G(3df,2p)
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.790 |
0.506 |
0.300 |
| C2 |
-0.591 |
0.648 |
-0.285 |
| C3 |
-1.528 |
-0.479 |
0.117 |
| F4 |
1.399 |
-0.649 |
-0.172 |
| H5 |
1.432 |
1.347 |
0.024 |
| H6 |
0.755 |
0.429 |
1.391 |
| H7 |
-0.506 |
0.705 |
-1.374 |
| H8 |
-0.989 |
1.613 |
0.048 |
| H9 |
-2.516 |
-0.348 |
-0.326 |
| H10 |
-1.139 |
-1.444 |
-0.209 |
| H11 |
-1.652 |
-0.518 |
1.202 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
F4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| C1 | | 1.5073 | 2.5257 | 1.3881 | 1.0930 | 1.0945 | 2.1266 | 2.1109 | 3.4720 | 2.7896 | 2.7978 |
C2 | 1.5073 | | 1.5200 | 2.3783 | 2.1630 | 2.1615 | 1.0936 | 1.0950 | 2.1679 | 2.1639 | 2.1674 | C3 | 2.5257 | 1.5200 | | 2.9461 | 3.4793 | 2.7680 | 2.1610 | 2.1610 | 1.0907 | 1.0904 | 1.0927 | F4 | 1.3881 | 2.3783 | 2.9461 | | 2.0051 | 2.0048 | 2.6279 | 3.2964 | 3.9296 | 2.6593 | 3.3485 | H5 | 1.0930 | 2.1630 | 3.4793 | 2.0051 | | 1.7802 | 2.4747 | 2.4365 | 4.3111 | 3.8013 | 3.7917 | H6 | 1.0945 | 2.1615 | 2.7680 | 2.0048 | 1.7802 | | 3.0518 | 2.5002 | 3.7756 | 3.1072 | 2.5937 | H7 | 2.1266 | 1.0936 | 2.1610 | 2.6279 | 2.4747 | 3.0518 | | 1.7545 | 2.4996 | 2.5247 | 3.0734 | H8 | 2.1109 | 1.0950 | 2.1610 | 3.2964 | 2.4365 | 2.5002 | 1.7545 | | 2.5130 | 3.0708 | 2.5126 | H9 | 3.4720 | 2.1679 | 1.0907 | 3.9296 | 4.3111 | 3.7756 | 2.4996 | 2.5130 | | 1.7641 | 1.7638 | H10 | 2.7896 | 2.1639 | 1.0904 | 2.6593 | 3.8013 | 3.1072 | 2.5247 | 3.0708 | 1.7641 | | 1.7640 | H11 | 2.7978 | 2.1674 | 1.0927 | 3.3485 | 3.7917 | 2.5937 | 3.0734 | 2.5126 | 1.7638 | 1.7640 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
113.090 |
|
C1 |
C2 |
H7 |
108.656 |
| C1 |
C2 |
H8 |
107.364 |
|
C2 |
C1 |
F4 |
110.387 |
| C2 |
C1 |
H5 |
111.587 |
|
C2 |
C1 |
H6 |
111.371 |
| C2 |
C3 |
H9 |
111.210 |
|
C2 |
C3 |
H10 |
110.906 |
| C2 |
C3 |
H11 |
111.054 |
|
C3 |
C2 |
H7 |
110.484 |
| C3 |
C2 |
H8 |
110.410 |
|
F4 |
C1 |
H5 |
107.240 |
| F4 |
C1 |
H6 |
107.122 |
|
H5 |
C1 |
H6 |
108.945 |
| H7 |
C2 |
H8 |
106.576 |
|
H9 |
C3 |
H10 |
107.955 |
| H9 |
C3 |
H11 |
107.763 |
|
H10 |
C3 |
H11 |
107.802 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-311+G(3df,2p)
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.187 |
|
|
|
| 2 |
C |
-0.212 |
|
|
|
| 3 |
C |
-0.439 |
|
|
|
| 4 |
F |
-0.442 |
|
|
|
| 5 |
H |
0.115 |
|
|
|
| 6 |
H |
0.127 |
|
|
|
| 7 |
H |
0.136 |
|
|
|
| 8 |
H |
0.125 |
|
|
|
| 9 |
H |
0.120 |
|
|
|
| 10 |
H |
0.146 |
|
|
|
| 11 |
H |
0.136 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.054 |
1.477 |
0.475 |
1.876 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-27.159 |
1.785 |
0.730 |
| y |
1.785 |
-25.619 |
-0.060 |
| z |
0.730 |
-0.060 |
-24.907 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.896 |
1.785 |
0.730 |
| y |
1.785 |
0.414 |
-0.060 |
| z |
0.730 |
-0.060 |
1.482 |
|
| Polar |
|---|
| 3z2-r2 | 2.964 |
|---|
| x2-y2 | -1.540 |
|---|
| xy | 1.785 |
|---|
| xz | 0.730 |
|---|
| yz | -0.060 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.374 |
0.241 |
0.057 |
| y |
0.241 |
5.901 |
-0.034 |
| z |
0.057 |
-0.034 |
5.600 |
<r2> (average value of r
2) Å
2
| <r2> |
91.446 |
| (<r2>)1/2 |
9.563 |
