Jump to
S1C2
Energy calculated at CCSD(T)=FULL/6-311+G(3df,2p)
| | hartrees |
|---|
| Energy at 0K | -190.430926 |
| Energy at 298.15K | |
| HF Energy | -189.603388 |
| Nuclear repulsion energy | 88.272120 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD(T)=FULL/6-311+G(3df,2p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3104 |
3104 |
|
|
|
|
| 2 |
A1 |
2291 |
2291 |
|
|
|
|
| 3 |
A1 |
1761 |
1761 |
|
|
|
|
| 4 |
A1 |
1487 |
1487 |
|
|
|
|
| 5 |
A1 |
909 |
909 |
|
|
|
|
| 6 |
B1 |
992 |
992 |
|
|
|
|
| 7 |
B1 |
638 |
638 |
|
|
|
|
| 8 |
B1 |
212 |
212 |
|
|
|
|
| 9 |
B2 |
3183 |
3183 |
|
|
|
|
| 10 |
B2 |
1066 |
1066 |
|
|
|
|
| 11 |
B2 |
455 |
455 |
|
|
|
|
| 12 |
B2 |
143i |
143i |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 7977.1 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 7977.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCSD(T)=FULL/6-311+G(3df,2p)
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-1.879 |
| C2 |
0.000 |
0.000 |
-0.556 |
| C3 |
0.000 |
0.000 |
0.722 |
| O4 |
0.000 |
0.000 |
1.898 |
| H5 |
0.000 |
0.924 |
-2.451 |
| H6 |
0.000 |
-0.924 |
-2.451 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
| C1 | | 1.3230 | 2.6018 | 3.7771 | 1.0867 | 1.0867 |
C2 | 1.3230 | | 1.2788 | 2.4541 | 2.1081 | 2.1081 | C3 | 2.6018 | 1.2788 | | 1.1753 | 3.3055 | 3.3055 | O4 | 3.7771 | 2.4541 | 1.1753 | | 4.4461 | 4.4461 | H5 | 1.0867 | 2.1081 | 3.3055 | 4.4461 | | 1.8480 | H6 | 1.0867 | 2.1081 | 3.3055 | 4.4461 | 1.8480 | |
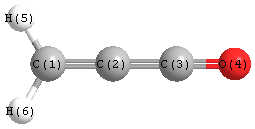 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
180.000 |
|
C2 |
C1 |
H5 |
121.752 |
| C2 |
C1 |
H6 |
121.752 |
|
C2 |
C3 |
O4 |
180.000 |
| H5 |
C1 |
H6 |
116.495 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CCSD(T)=FULL/6-311+G(3df,2p)
| | hartrees |
|---|
| Energy at 0K | -190.432648 |
| Energy at 298.15K | |
| HF Energy | -189.602792 |
| Nuclear repulsion energy | 88.579316 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD(T)=FULL/6-311+G(3df,2p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3200 |
3200 |
|
|
|
|
| 2 |
A' |
3114 |
3114 |
|
|
|
|
| 3 |
A' |
2186 |
2186 |
|
|
|
|
| 4 |
A' |
1720 |
1720 |
|
|
|
|
| 5 |
A' |
1485 |
1485 |
|
|
|
|
| 6 |
A' |
1074 |
1074 |
|
|
|
|
| 7 |
A' |
935 |
935 |
|
|
|
|
| 8 |
A' |
499 |
499 |
|
|
|
|
| 9 |
A' |
173 |
173 |
|
|
|
|
| 10 |
A" |
1004 |
1004 |
|
|
|
|
| 11 |
A" |
702 |
702 |
|
|
|
|
| 12 |
A" |
276 |
276 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 8183.6 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 8183.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCSD(T)=FULL/6-311+G(3df,2p)
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.071 |
-1.465 |
0.000 |
| C2 |
0.000 |
-0.672 |
0.000 |
| C3 |
-0.364 |
0.586 |
0.000 |
| O4 |
-0.912 |
1.616 |
0.000 |
| H5 |
2.087 |
-1.080 |
0.000 |
| H6 |
0.963 |
-2.545 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
| C1 | | 1.3333 | 2.5031 | 3.6645 | 1.0864 | 1.0849 |
C2 | 1.3333 | | 1.3088 | 2.4628 | 2.1270 | 2.1063 | C3 | 2.5031 | 1.3088 | | 1.1672 | 2.9638 | 3.4001 | O4 | 3.6645 | 2.4628 | 1.1672 | | 4.0333 | 4.5640 | H5 | 1.0864 | 2.1270 | 2.9638 | 4.0333 | | 1.8458 | H6 | 1.0849 | 2.1063 | 3.4001 | 4.5640 | 1.8458 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
142.667 |
|
C2 |
C1 |
H5 |
122.734 |
| C2 |
C1 |
H6 |
120.819 |
|
C2 |
C3 |
O4 |
168.127 |
| H5 |
C1 |
H6 |
116.447 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
