Jump to
S1C2
Energy calculated at HSEh1PBE/6-311+G(3df,2p)
| | hartrees |
|---|
| Energy at 0K | -278.129629 |
| Energy at 298.15K | |
| HF Energy | -278.129629 |
| Nuclear repulsion energy | 126.113933 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-311+G(3df,2p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3058 |
3058 |
0.00 |
235.93 |
0.06 |
0.11 |
| 2 |
Ag |
1511 |
1511 |
0.00 |
7.56 |
0.74 |
0.85 |
| 3 |
Ag |
1445 |
1445 |
0.00 |
2.05 |
0.37 |
0.54 |
| 4 |
Ag |
1100 |
1100 |
0.00 |
6.44 |
0.61 |
0.76 |
| 5 |
Ag |
1084 |
1084 |
0.00 |
8.38 |
0.29 |
0.44 |
| 6 |
Ag |
463 |
463 |
0.00 |
2.43 |
0.37 |
0.54 |
| 7 |
Au |
3130 |
3130 |
40.77 |
0.00 |
0.00 |
0.00 |
| 8 |
Au |
1237 |
1237 |
4.20 |
0.00 |
0.00 |
0.00 |
| 9 |
Au |
812 |
812 |
0.11 |
0.00 |
0.00 |
0.00 |
| 10 |
Au |
113 |
113 |
12.68 |
0.00 |
0.00 |
0.00 |
| 11 |
Bg |
3103 |
3103 |
0.00 |
110.57 |
0.75 |
0.86 |
| 12 |
Bg |
1299 |
1299 |
0.00 |
7.08 |
0.75 |
0.86 |
| 13 |
Bg |
1176 |
1176 |
0.00 |
1.30 |
0.75 |
0.86 |
| 14 |
Bu |
3065 |
3065 |
56.23 |
0.00 |
0.00 |
0.00 |
| 15 |
Bu |
1520 |
1520 |
5.46 |
0.00 |
0.00 |
0.00 |
| 16 |
Bu |
1356 |
1356 |
5.37 |
0.00 |
0.00 |
0.00 |
| 17 |
Bu |
1096 |
1096 |
218.44 |
0.00 |
0.00 |
0.00 |
| 18 |
Bu |
279 |
279 |
18.73 |
0.00 |
0.00 |
0.00 |
Unscaled Zero Point Vibrational Energy (zpe) 13423.8 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 13423.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-311+G(3df,2p)
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.422 |
0.627 |
0.000 |
| C2 |
-0.422 |
-0.627 |
0.000 |
| F3 |
-0.422 |
1.718 |
0.000 |
| F4 |
0.422 |
-1.718 |
0.000 |
| H5 |
1.052 |
0.673 |
0.891 |
| H6 |
1.052 |
0.673 |
-0.891 |
| H7 |
-1.052 |
-0.673 |
0.891 |
| H8 |
-1.052 |
-0.673 |
-0.891 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5110 | 1.3794 | 2.3449 | 1.0920 | 1.0920 | 2.1572 | 2.1572 |
C2 | 1.5110 | | 2.3449 | 1.3794 | 2.1572 | 2.1572 | 1.0920 | 1.0920 | F3 | 1.3794 | 2.3449 | | 3.5383 | 2.0142 | 2.0142 | 2.6281 | 2.6281 | F4 | 2.3449 | 1.3794 | 3.5383 | | 2.6281 | 2.6281 | 2.0142 | 2.0142 | H5 | 1.0920 | 2.1572 | 2.0142 | 2.6281 | | 1.7819 | 2.4968 | 3.0674 | H6 | 1.0920 | 2.1572 | 2.0142 | 2.6281 | 1.7819 | | 3.0674 | 2.4968 | H7 | 2.1572 | 1.0920 | 2.6281 | 2.0142 | 2.4968 | 3.0674 | | 1.7819 | H8 | 2.1572 | 1.0920 | 2.6281 | 2.0142 | 3.0674 | 2.4968 | 1.7819 | |
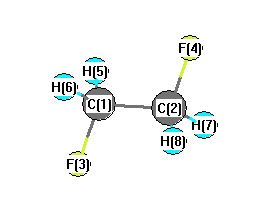 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
108.359 |
|
C1 |
C2 |
H7 |
110.913 |
| C1 |
C2 |
H8 |
110.913 |
|
C2 |
C1 |
F3 |
108.359 |
| C2 |
C1 |
H5 |
110.913 |
|
C2 |
C1 |
H6 |
110.913 |
| F3 |
C1 |
H5 |
108.623 |
|
F3 |
C1 |
H6 |
108.623 |
| F4 |
C2 |
H7 |
108.623 |
|
F4 |
C2 |
H8 |
108.623 |
| H5 |
C1 |
H6 |
109.345 |
|
H7 |
C2 |
H8 |
109.345 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-311+G(3df,2p)
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.173 |
|
|
|
| 2 |
C |
0.173 |
|
|
|
| 3 |
F |
-0.434 |
|
|
|
| 4 |
F |
-0.434 |
|
|
|
| 5 |
H |
0.130 |
|
|
|
| 6 |
H |
0.130 |
|
|
|
| 7 |
H |
0.130 |
|
|
|
| 8 |
H |
0.130 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-22.230 |
3.166 |
0.000 |
| y |
3.166 |
-29.294 |
0.000 |
| z |
0.000 |
0.000 |
-21.573 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.204 |
3.166 |
0.000 |
| y |
3.166 |
-7.393 |
0.000 |
| z |
0.000 |
0.000 |
4.189 |
|
| Polar |
|---|
| 3z2-r2 | 8.378 |
|---|
| x2-y2 | 7.064 |
|---|
| xy | 3.166 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.283 |
-0.061 |
0.000 |
| y |
-0.061 |
4.510 |
0.000 |
| z |
0.000 |
0.000 |
3.921 |
<r2> (average value of r
2) Å
2
| <r2> |
87.815 |
| (<r2>)1/2 |
9.371 |
Jump to
S1C1
Energy calculated at HSEh1PBE/6-311+G(3df,2p)
| | hartrees |
|---|
| Energy at 0K | -278.131467 |
| Energy at 298.15K | |
| HF Energy | -278.131467 |
| Nuclear repulsion energy | 128.145973 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-311+G(3df,2p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3099 |
3099 |
13.49 |
85.85 |
0.75 |
0.85 |
| 2 |
A |
3054 |
3054 |
35.87 |
221.81 |
0.02 |
0.03 |
| 3 |
A |
1483 |
1483 |
0.90 |
1.38 |
0.71 |
0.83 |
| 4 |
A |
1435 |
1435 |
11.47 |
1.49 |
0.22 |
0.36 |
| 5 |
A |
1307 |
1307 |
3.44 |
8.52 |
0.66 |
0.79 |
| 6 |
A |
1135 |
1135 |
84.02 |
3.29 |
0.35 |
0.52 |
| 7 |
A |
1130 |
1130 |
14.33 |
1.28 |
0.46 |
0.63 |
| 8 |
A |
880 |
880 |
20.75 |
5.29 |
0.17 |
0.29 |
| 9 |
A |
325 |
325 |
0.49 |
0.39 |
0.29 |
0.45 |
| 10 |
A |
151 |
151 |
3.65 |
0.13 |
0.75 |
0.86 |
| 11 |
B |
3113 |
3113 |
31.02 |
12.12 |
0.75 |
0.86 |
| 12 |
B |
3044 |
3044 |
8.35 |
33.13 |
0.75 |
0.86 |
| 13 |
B |
1483 |
1483 |
10.90 |
6.71 |
0.75 |
0.86 |
| 14 |
B |
1395 |
1395 |
5.44 |
0.01 |
0.75 |
0.86 |
| 15 |
B |
1266 |
1266 |
7.00 |
2.24 |
0.75 |
0.86 |
| 16 |
B |
1103 |
1103 |
62.11 |
3.83 |
0.75 |
0.86 |
| 17 |
B |
910 |
910 |
40.52 |
2.29 |
0.75 |
0.86 |
| 18 |
B |
499 |
499 |
15.80 |
0.44 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 13405.2 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 13405.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-311+G(3df,2p)
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.264 |
0.701 |
0.505 |
| C2 |
-0.264 |
-0.701 |
0.505 |
| F3 |
-0.264 |
1.425 |
-0.541 |
| F4 |
0.264 |
-1.425 |
-0.541 |
| H5 |
-0.014 |
1.197 |
1.440 |
| H6 |
1.352 |
0.702 |
0.402 |
| H7 |
0.014 |
-1.197 |
1.440 |
| H8 |
-1.352 |
-0.702 |
0.402 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.4978 | 1.3780 | 2.3693 | 1.0942 | 1.0927 | 2.1299 | 2.1428 |
C2 | 1.4978 | | 2.3693 | 1.3780 | 2.1299 | 2.1428 | 1.0942 | 1.0927 | F3 | 1.3780 | 2.3693 | | 2.8988 | 2.0098 | 2.0067 | 3.2978 | 2.5688 | F4 | 2.3693 | 1.3780 | 2.8988 | | 3.2978 | 2.5688 | 2.0098 | 2.0067 | H5 | 1.0942 | 2.1299 | 2.0098 | 3.2978 | | 1.7853 | 2.3935 | 2.5440 | H6 | 1.0927 | 2.1428 | 2.0067 | 2.5688 | 1.7853 | | 2.5440 | 3.0472 | H7 | 2.1299 | 1.0942 | 3.2978 | 2.0098 | 2.3935 | 2.5440 | | 1.7853 | H8 | 2.1428 | 1.0927 | 2.5688 | 2.0067 | 2.5440 | 3.0472 | 1.7853 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
110.878 |
|
C1 |
C2 |
H7 |
109.529 |
| C1 |
C2 |
H8 |
110.651 |
|
C2 |
C1 |
F3 |
110.878 |
| C2 |
C1 |
H5 |
109.529 |
|
C2 |
C1 |
H6 |
110.651 |
| F3 |
C1 |
H5 |
108.220 |
|
F3 |
C1 |
H6 |
108.065 |
| F4 |
C2 |
H7 |
108.220 |
|
F4 |
C2 |
H8 |
108.065 |
| H5 |
C1 |
H6 |
109.445 |
|
H7 |
C2 |
H8 |
109.445 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-311+G(3df,2p)
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.178 |
|
|
|
| 2 |
C |
0.178 |
|
|
|
| 3 |
F |
-0.438 |
|
|
|
| 4 |
F |
-0.438 |
|
|
|
| 5 |
H |
0.128 |
|
|
|
| 6 |
H |
0.133 |
|
|
|
| 7 |
H |
0.128 |
|
|
|
| 8 |
H |
0.133 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
2.700 |
2.700 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-21.798 |
1.719 |
0.000 |
| y |
1.719 |
-26.259 |
0.000 |
| z |
0.000 |
0.000 |
-22.528 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.596 |
1.719 |
0.000 |
| y |
1.719 |
-4.096 |
0.000 |
| z |
0.000 |
0.000 |
1.500 |
|
| Polar |
|---|
| 3z2-r2 | 3.000 |
|---|
| x2-y2 | 4.461 |
|---|
| xy | 1.719 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.081 |
0.038 |
0.000 |
| y |
0.038 |
4.476 |
0.000 |
| z |
0.000 |
0.000 |
4.265 |
<r2> (average value of r
2) Å
2
| <r2> |
79.549 |
| (<r2>)1/2 |
8.919 |
