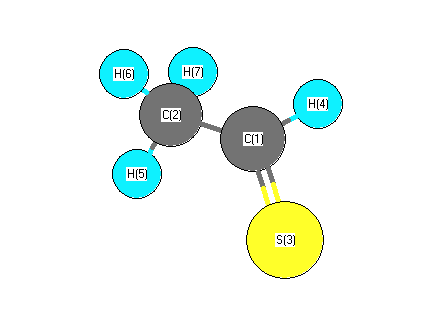
Vibrational Frequencies calculated at HSEh1PBE/6-311+G(3df,2p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3156 |
3156 |
5.67 |
|
|
|
| 2 |
A' |
3079 |
3079 |
20.49 |
|
|
|
| 3 |
A' |
3025 |
3025 |
2.43 |
|
|
|
| 4 |
A' |
1476 |
1476 |
8.13 |
|
|
|
| 5 |
A' |
1380 |
1380 |
62.93 |
|
|
|
| 6 |
A' |
1376 |
1376 |
30.02 |
|
|
|
| 7 |
A' |
1183 |
1183 |
41.48 |
|
|
|
| 8 |
A' |
1085 |
1085 |
7.43 |
|
|
|
| 9 |
A' |
845 |
845 |
1.27 |
|
|
|
| 10 |
A' |
396 |
396 |
2.05 |
|
|
|
| 11 |
A" |
3078 |
3078 |
3.73 |
|
|
|
| 12 |
A" |
1462 |
1462 |
9.98 |
|
|
|
| 13 |
A" |
1043 |
1043 |
4.78 |
|
|
|
| 14 |
A" |
769 |
769 |
13.76 |
|
|
|
| 15 |
A" |
173 |
173 |
0.14 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 11761.5 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 11761.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-311+G(3df,2p)
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.183 |
|
|
|
| 2 |
C |
-0.185 |
|
|
|
| 3 |
S |
-0.202 |
|
|
|
| 4 |
H |
0.141 |
|
|
|
| 5 |
H |
0.143 |
|
|
|
| 6 |
H |
0.143 |
|
|
|
| 7 |
H |
0.143 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.712 |
1.897 |
0.000 |
2.555 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-26.534 |
-0.670 |
0.000 |
| y |
-0.670 |
-25.333 |
0.000 |
| z |
0.000 |
0.000 |
-26.168 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.784 |
-0.670 |
0.000 |
| y |
-0.670 |
1.018 |
0.000 |
| z |
0.000 |
0.000 |
-0.235 |
|
| Polar |
|---|
| 3z2-r2 | -0.469 |
|---|
| x2-y2 | -1.201 |
|---|
| xy | -0.670 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.066 |
-1.666 |
0.000 |
| y |
-1.666 |
7.823 |
0.000 |
| z |
0.000 |
0.000 |
5.226 |
<r2> (average value of r
2) Å
2
| <r2> |
73.479 |
| (<r2>)1/2 |
8.572 |