return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for Si3H8 (trisilane)

using model chemistry: MP2/6-311+G(3df,2p)

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2/6-311+G(3df,2p)
 hartrees
Energy at 0K-871.846216
Energy at 298.15K-871.853938
HF Energy-871.475315
Nuclear repulsion energy193.081557
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-311+G(3df,2p)
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 2276 2148 128.28      
2 A1 2266 2138 24.20      
3 A1 2246 2119 67.40      
4 A1 979 923 73.95      
5 A1 952 898 1.68      
6 A1 912 861 207.55      
7 A1 579 546 6.14      
8 A1 398 376 0.54      
9 A1 94 88 1.38      
10 A2 2272 2144 0.00      
11 A2 971 916 0.00      
12 A2 721 680 0.00      
13 A2 421 397 0.00      
14 A2 86 81 0.00      
15 B1 2277 2148 236.98      
16 B1 2255 2127 22.21      
17 B1 975 920 81.12      
18 B1 608 574 9.80      
19 B1 319 301 21.28      
20 B1 104 98 0.09      
21 B2 2275 2146 90.47      
22 B2 2262 2134 86.68      
23 B2 971 916 41.35      
24 B2 900 849 336.20      
25 B2 728 687 286.67      
26 B2 474 447 8.49      
27 B2 439 415 20.38      

Unscaled Zero Point Vibrational Energy (zpe) 14879.3 cm-1
Scaled (by 0.9434) Zero Point Vibrational Energy (zpe) 14037.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/6-311+G(3df,2p)
ABC
0.30264 0.06724 0.05834

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/6-311+G(3df,2p)

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
Si1 0.000 0.000 0.913
Si2 0.000 1.923 -0.428
Si3 0.000 -1.923 -0.428
H4 1.201 0.000 1.782
H5 -1.201 0.000 1.782
H6 0.000 3.152 0.397
H7 0.000 -3.152 0.397
H8 1.202 1.928 -1.291
H9 -1.202 1.928 -1.291
H10 -1.202 -1.928 -1.291
H11 1.202 -1.928 -1.291

Atom - Atom Distances (Å)
  Si1 Si2 Si3 H4 H5 H6 H7 H8 H9 H10 H11
Si12.34432.34431.48271.48273.19353.19353.16543.16543.16543.1654
Si22.34433.84653.16623.16621.47925.14131.47971.47974.12614.1261
Si32.34433.84653.16623.16625.14131.47924.12614.12611.47971.4797
H41.48273.16623.16622.40203.64623.64623.62824.35164.35163.6282
H51.48273.16623.16622.40203.64623.64624.35163.62823.62824.3516
H63.19351.47925.14133.64623.64626.30302.40582.40585.48615.4861
H73.19355.14131.47923.64623.64626.30305.48615.48612.40582.4058
H83.16541.47974.12613.62824.35162.40585.48612.40334.54443.8568
H93.16541.47974.12614.35163.62822.40585.48612.40333.85684.5444
H103.16544.12611.47974.35163.62825.48612.40584.54443.85682.4033
H113.16544.12611.47973.62824.35165.48612.40583.85684.54442.4033

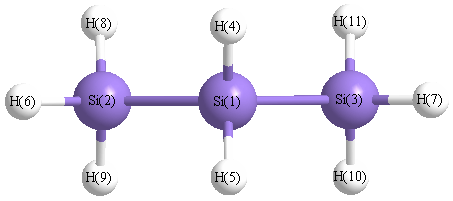
picture of trisilane state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
S1 S2 H6 111.263 S1 S2 H8 109.663
S1 S2 H9 109.663 S1 S3 H7 111.263
S1 S3 H10 109.663 S1 S3 H11 109.663
S2 S1 S3 110.252 S2 S1 H4 109.591
S2 S1 H5 109.591 S3 S1 H4 109.591
S3 S1 H5 109.591 H4 S1 H5 108.194
H6 S2 H8 108.798 H6 S2 H9 108.798
H7 S3 H10 108.798 H7 S3 H11 108.798
H8 S2 H9 108.604 H10 S3 H11 108.604
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability